Other Sites:


Bibliography Options Menu

Robert J. Robbins is a biologist, an educator, a science administrator, a publisher, an information technologist, and an IT leader and manager who specializes in advancing biomedical knowledge and supporting education through the application of information technology. More About: RJR | OUR TEAM | OUR SERVICES | THIS WEBSITE
RJR: Recommended Bibliography 17 Nov 2026 at 01:31 Created:
Metagenomics
While genomics is the study of DNA extracted from individuals — individual cells, tissues, or organisms — metagenomics is a more recent refinement that analyzes samples of pooled DNA taken from the environment, not from an individual. Like genomics, metagenomic methods have great potential in many areas of biology, but none so much as in providing access to the hitherto invisible world of unculturable microbes, often estimated to comprise 90% or more of bacterial species and, in some ecosystems, the bulk of the biomass. A recent describes how this new science of metagenomics is beginning to reveal the secrets of our microbial world: The opportunity that stands before microbiologists today is akin to a reinvention of the microscope in the expanse of research questions it opens to investigation. Metagenomics provides a new way of examining the microbial world that not only will transform modern microbiology but has the potential to revolutionize understanding of the entire living world. In metagenomics, the power of genomic analysis is applied to entire communities of microbes, bypassing the need to isolate and culture individual bacterial community members.
Created with PubMed® Query: ( metagenomic OR metagenomics OR metagenome ) NOT pmcbook NOT ispreviousversion
Citations The Papers (from PubMed®)
RevDate: 2026-11-14
CmpDate: 2026-11-14
Cell-associated viral community composition and its functional potential in a dimictic lake on the Canadian shield.
Frontiers in microbiology, 17:1941165.
The Turkey Lake Watershed (TLW), located in Northern Ontario, forms a cascading lake system that drains into Lake Superior. Historically, the TLW has served as a key research site for studies on acid rain and climate change. Despite its long-standing ecological importance, investigations into its microbial communities remain limited, and to date, no studies have explicitly examined the viral component of this watershed. Viruses have been frequently implicated in nutrient cycling in aquatic environments and strongly influence their microbial hosts. Therefore, viral ecology offers a powerful way to extend the environmental legacy of the TLW by identifying the biological mechanisms through which long-term changes in acid deposition, climate, hydrology, carbon chemistry, and forest disturbance propagate through microbial food webs. First steps can be a preliminary view into the type of viruses that are present, and information on which microbial hosts they may be infecting. We conducted a seasonal metagenomic survey of Big Turkey Lake (BTL)-one of the primary lakes within the TLW-from summer 2018 to winter 2020. Our study characterized the diversity and composition of cell-associated viral communities, predicted potential host relationships, and identified auxiliary metabolic genes (AMGs) to better understand the functional roles of these viruses. The results revealed a dynamic viral community that varied across seasons. Most viral contigs were classified as Caudoviricetes, while members of Phycodnaviridae emerged as important non-bacteriophage contributors to the viral assemblage. We reconstructed six draft viral metagenome-assembled genomes (vMAGs), ranging in size from 11 to 45 kbp, all identified as Caudoviricetes, with two being potential temperate phages and a third containing ribosomal genes. Notably, AMGs associated with lipid metabolism were detected exclusively in winter, suggesting a potential season- or niche-specific role for these genes in BTL. AMGs related to phosphorus acquisition were present in all samples, suggesting a potential coupling between transport dynamics of dissolved organic matter and phosphorus previously described in the TLW, and the host-virus dynamics described here. Together, these findings provide the first insight into the viral ecology of the TLW and establish a foundation for future studies investigating the roles of viruses in shaping the microbial dynamics in this important habitat.
Additional Links: PMID-43032552
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032552,
year = {2026},
author = {Collis, CC and Cameron, ES and Müller, KM and Emelko, MB and Tromas, N and Nissimov, JI},
title = {Cell-associated viral community composition and its functional potential in a dimictic lake on the Canadian shield.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1941165},
pmid = {43032552},
issn = {1664-302X},
abstract = {The Turkey Lake Watershed (TLW), located in Northern Ontario, forms a cascading lake system that drains into Lake Superior. Historically, the TLW has served as a key research site for studies on acid rain and climate change. Despite its long-standing ecological importance, investigations into its microbial communities remain limited, and to date, no studies have explicitly examined the viral component of this watershed. Viruses have been frequently implicated in nutrient cycling in aquatic environments and strongly influence their microbial hosts. Therefore, viral ecology offers a powerful way to extend the environmental legacy of the TLW by identifying the biological mechanisms through which long-term changes in acid deposition, climate, hydrology, carbon chemistry, and forest disturbance propagate through microbial food webs. First steps can be a preliminary view into the type of viruses that are present, and information on which microbial hosts they may be infecting. We conducted a seasonal metagenomic survey of Big Turkey Lake (BTL)-one of the primary lakes within the TLW-from summer 2018 to winter 2020. Our study characterized the diversity and composition of cell-associated viral communities, predicted potential host relationships, and identified auxiliary metabolic genes (AMGs) to better understand the functional roles of these viruses. The results revealed a dynamic viral community that varied across seasons. Most viral contigs were classified as Caudoviricetes, while members of Phycodnaviridae emerged as important non-bacteriophage contributors to the viral assemblage. We reconstructed six draft viral metagenome-assembled genomes (vMAGs), ranging in size from 11 to 45 kbp, all identified as Caudoviricetes, with two being potential temperate phages and a third containing ribosomal genes. Notably, AMGs associated with lipid metabolism were detected exclusively in winter, suggesting a potential season- or niche-specific role for these genes in BTL. AMGs related to phosphorus acquisition were present in all samples, suggesting a potential coupling between transport dynamics of dissolved organic matter and phosphorus previously described in the TLW, and the host-virus dynamics described here. Together, these findings provide the first insight into the viral ecology of the TLW and establish a foundation for future studies investigating the roles of viruses in shaping the microbial dynamics in this important habitat.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-14
Multi-scale hybrid correction of noisy long reads.
iScience, 29(11):118150.
Long-read sequencing offers superior genome resolution but is hindered by error rates of 1-15%. Existing correction methods often struggle to suppress sequencing errors without removing genuine biological variation, leading to over-correction. DADEC addresses this trade-off through a confidence-ordered framework comprising stringent de Bruijn graph (DBG) correction, haplotype-aware short-read alignment refinement, and permissive DBG recovery. Across seven simulated and three real datasets, DADEC achieved the lowest mismatch and indel rates in most benchmark settings while maintaining high haplotype coverage. DADEC also improved assembly contiguity and strain-level metagenomic classification. On an independently simulated Arabidopsis thaliana dataset with ground truth, DADEC reduced correction-event false discovery and false negative rates by 38.9% and 40.2%, respectively, compared with the next-best method. DADEC therefore provides an accurate hybrid correction strategy for noisy long reads while reducing over-correction and maintaining haplotype representation.
Additional Links: PMID-43032624
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032624,
year = {2026},
author = {Zhang, W and Zhang, Y and Li, F and Li, Y and Gong, H and Luo, J and Yuan, S and Luo, X and Liu, Y},
title = {Multi-scale hybrid correction of noisy long reads.},
journal = {iScience},
volume = {29},
number = {11},
pages = {118150},
pmid = {43032624},
issn = {2589-0042},
abstract = {Long-read sequencing offers superior genome resolution but is hindered by error rates of 1-15%. Existing correction methods often struggle to suppress sequencing errors without removing genuine biological variation, leading to over-correction. DADEC addresses this trade-off through a confidence-ordered framework comprising stringent de Bruijn graph (DBG) correction, haplotype-aware short-read alignment refinement, and permissive DBG recovery. Across seven simulated and three real datasets, DADEC achieved the lowest mismatch and indel rates in most benchmark settings while maintaining high haplotype coverage. DADEC also improved assembly contiguity and strain-level metagenomic classification. On an independently simulated Arabidopsis thaliana dataset with ground truth, DADEC reduced correction-event false discovery and false negative rates by 38.9% and 40.2%, respectively, compared with the next-best method. DADEC therefore provides an accurate hybrid correction strategy for noisy long reads while reducing over-correction and maintaining haplotype representation.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-14
Temporal enrichment of bacterial monoamine oxidase gene aboard the international space station and neuromodulator impacts of dopamine-driven ammonium formed by space isolate Leifsonia virtsii on Caenorhabditis elegans.
Frontiers in microbiology, 17:1943403.
Dopamine regulates neurological functions, whereas its deficiency is linked with cognitive impairment and neurodegenerative diseases. The spatiotemporal distribution of dopamine-degrading microbes and the implications of dopamine catabolites on eukaryotes remain unexplored. Here, we demonstrate dopamine metabolism in Leifsonia virtsii F6_8S_P_1A[T], an Actinomycetes isolated from the crew quarters aboard the International Space Station (ISS), and reveal dopamine catabolite-driven modulation of cognition and chemotaxis in Caenorhabditis elegans. L. virtsii shared a high 16S rRNA gene sequence similarity (>97.9%) and OrthoANI values (79.44-84.96%) with the Leifsonia species, isolated from terrestrial and aquatic habitats. A key gene encoding monoamine oxidase (MAO) involved in dopamine catabolism was found in 61.5% of validly established Leifsonia species, including L. virtsii, but not in ISS co-isolate L. williamsii F6_8S_P_1B[T]. Analysis of the ISS metagenome revealed temporal enrichment of MAO-associated signatures across Microbial Tracking studies (MT1 to MT3, 2015-2022), driven predominantly by candidate MAO homologs taxonomically associated with Actinomycetota in MT3, whereas strong F6-8S-P-1A-like MAO signals were restricted to two MT2 libraries. Dopamine input stimulated MAO gene expression in L. virtsii at early growth phase, and was found to complement cell division, biofilm establishment and extracellular NH4 [+] formation in L. virtsii. MAO expression decreased with increasing time/NH4 [+] concentration, whereas expression of marker genes representing DNA synthesis and transcription (gyrB), translation (tuf) and biofilm formation (cslA) exhibited concurrent upregulation. Dopamine-driven NH4 [+] formation further altered cognition and chemotaxis in C. elegans. Our analysis suggests that the dopamine metabolic pathway detected in L. virtsii is not an ISS-specific trait and rather represents a phenotype retrieved from Earth. The present study provides direct evidence of spatiotemporal flexibility and heterogeneous dopamine-metabolizing capacity among Actinomycetes. The findings further demonstrate that a dopamine catabolite can modulate chemosensory interactions in a eukaryotic model and highlight the succession and persistence of MAO-bearing taxa aboard ISS.
Additional Links: PMID-43032692
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032692,
year = {2026},
author = {Mayuri, N and Dhanyashree, HR and Shastry, RP and Venkateswaran, K and Miliotis, G and Singh, NK and Stothard, P and Arun, AB and Hameed, A},
title = {Temporal enrichment of bacterial monoamine oxidase gene aboard the international space station and neuromodulator impacts of dopamine-driven ammonium formed by space isolate Leifsonia virtsii on Caenorhabditis elegans.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1943403},
pmid = {43032692},
issn = {1664-302X},
abstract = {Dopamine regulates neurological functions, whereas its deficiency is linked with cognitive impairment and neurodegenerative diseases. The spatiotemporal distribution of dopamine-degrading microbes and the implications of dopamine catabolites on eukaryotes remain unexplored. Here, we demonstrate dopamine metabolism in Leifsonia virtsii F6_8S_P_1A[T], an Actinomycetes isolated from the crew quarters aboard the International Space Station (ISS), and reveal dopamine catabolite-driven modulation of cognition and chemotaxis in Caenorhabditis elegans. L. virtsii shared a high 16S rRNA gene sequence similarity (>97.9%) and OrthoANI values (79.44-84.96%) with the Leifsonia species, isolated from terrestrial and aquatic habitats. A key gene encoding monoamine oxidase (MAO) involved in dopamine catabolism was found in 61.5% of validly established Leifsonia species, including L. virtsii, but not in ISS co-isolate L. williamsii F6_8S_P_1B[T]. Analysis of the ISS metagenome revealed temporal enrichment of MAO-associated signatures across Microbial Tracking studies (MT1 to MT3, 2015-2022), driven predominantly by candidate MAO homologs taxonomically associated with Actinomycetota in MT3, whereas strong F6-8S-P-1A-like MAO signals were restricted to two MT2 libraries. Dopamine input stimulated MAO gene expression in L. virtsii at early growth phase, and was found to complement cell division, biofilm establishment and extracellular NH4 [+] formation in L. virtsii. MAO expression decreased with increasing time/NH4 [+] concentration, whereas expression of marker genes representing DNA synthesis and transcription (gyrB), translation (tuf) and biofilm formation (cslA) exhibited concurrent upregulation. Dopamine-driven NH4 [+] formation further altered cognition and chemotaxis in C. elegans. Our analysis suggests that the dopamine metabolic pathway detected in L. virtsii is not an ISS-specific trait and rather represents a phenotype retrieved from Earth. The present study provides direct evidence of spatiotemporal flexibility and heterogeneous dopamine-metabolizing capacity among Actinomycetes. The findings further demonstrate that a dopamine catabolite can modulate chemosensory interactions in a eukaryotic model and highlight the succession and persistence of MAO-bearing taxa aboard ISS.},
}
RevDate: 2026-11-14
Intratumoral microbiota: novel perspectives on pathogenesis, immune microenvironment and clinical translation of nasopharyngeal carcinoma.
Clinical & translational oncology : official publication of the Federation of Spanish Oncology Societies and of the National Cancer Institute of Mexico [Epub ahead of print].
Nasopharyngeal carcinoma (NPC) is an epithelial malignancy characterized by marked geographic heterogeneity, with a high endemic prevalence in East and Southeast Asia. Despite advances in radiotherapy, chemotherapy, and immunotherapy, the prognosis of locally advanced and metastatic NPC remains suboptimal, creating an urgent need for novel pathogenic insights and therapeutic targets. Recent metagenomic and multi-omics studies have revealed that intratumoral microbial communities, including viruses, bacteria, and fungi, are profoundly implicated in tumor initiation, treatment response, and clinical outcomes. NPC has become a focus of research into tumor-resident microbiota and host-microbe interactions. This review summarizes the compositional characteristics, colonization pathways, and spatial distribution of the microbial community in NPC, with a focus on the molecular mechanisms of Epstein-Barr virus (EBV) and other intratumoral microorganisms in regulating inflammation, genomic stability, and anti-tumor immunity. We further summarized the clinical application of intratumoral microbiota in the development of diagnostic biomarkers, prognostic stratification, and precise treatment. This includes non-invasive screening that combines microbial characteristics with EBV DNA, microbiota-mediated chemoradiotherapy, enhanced immune checkpoint inhibitors, and emerging models, such as engineered bacteria and phage therapy. Finally, we analyzed key challenges, such as low-biomass detection, standardization, and interindividual heterogeneity, while looking ahead to future directions in spatial multi-omics, AI-driven data integration, and interdisciplinary collaboration to advance precision medicine for NPC.
Additional Links: PMID-43033309
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43033309,
year = {2026},
author = {Xu, Y and Gong, L and Lu, J and Wang, M},
title = {Intratumoral microbiota: novel perspectives on pathogenesis, immune microenvironment and clinical translation of nasopharyngeal carcinoma.},
journal = {Clinical & translational oncology : official publication of the Federation of Spanish Oncology Societies and of the National Cancer Institute of Mexico},
volume = {},
number = {},
pages = {},
pmid = {43033309},
issn = {1699-3055},
abstract = {Nasopharyngeal carcinoma (NPC) is an epithelial malignancy characterized by marked geographic heterogeneity, with a high endemic prevalence in East and Southeast Asia. Despite advances in radiotherapy, chemotherapy, and immunotherapy, the prognosis of locally advanced and metastatic NPC remains suboptimal, creating an urgent need for novel pathogenic insights and therapeutic targets. Recent metagenomic and multi-omics studies have revealed that intratumoral microbial communities, including viruses, bacteria, and fungi, are profoundly implicated in tumor initiation, treatment response, and clinical outcomes. NPC has become a focus of research into tumor-resident microbiota and host-microbe interactions. This review summarizes the compositional characteristics, colonization pathways, and spatial distribution of the microbial community in NPC, with a focus on the molecular mechanisms of Epstein-Barr virus (EBV) and other intratumoral microorganisms in regulating inflammation, genomic stability, and anti-tumor immunity. We further summarized the clinical application of intratumoral microbiota in the development of diagnostic biomarkers, prognostic stratification, and precise treatment. This includes non-invasive screening that combines microbial characteristics with EBV DNA, microbiota-mediated chemoradiotherapy, enhanced immune checkpoint inhibitors, and emerging models, such as engineered bacteria and phage therapy. Finally, we analyzed key challenges, such as low-biomass detection, standardization, and interindividual heterogeneity, while looking ahead to future directions in spatial multi-omics, AI-driven data integration, and interdisciplinary collaboration to advance precision medicine for NPC.},
}
RevDate: 2026-11-14
Removal performance and putative degradation mechanisms of bisphenol A in bioretention cells.
Journal of environmental management, 420:131383 pii:S0301-4797(26)02843-4 [Epub ahead of print].
Bisphenol A (BPA) is a typical endocrine-disrupting pollutant. It can enter receiving waters through urban stormwater runoff and pose potential risks to aquatic ecosystems. Bioretention cells are widely used stormwater control measures for urban runoff and can retain and transform organic pollutants. However, the removal performance and mechanisms of BPA in bioretention cells remain unclear. In this study, two bioretention cells were constructed. BPA removal under different influent concentrations was investigated. The main removal zone and transformation process of BPA were also analyzed. The results showed that BPA removal by the bioretention cells exceeded 70%, with the main removal occurring in the upper filter media layer. Both filter media adsorption and microbial processes contributed to BPA removal. Several transformation products in the bioretention cells were putatively identified by liquid chromatography-mass spectrometry (LC-MS), including monohydroxylated BPA, 4-hydroxyacetophenone, 3-oxohexanedioic acid, and other intermediates. Together with the metagenomic results, these products suggest that BPA may undergo initial oxidation, side-chain cleavage, intermediate transformation, and ring-opening processes in bioretention cells. These findings provide a scientific basis for BPA control in urban stormwater runoff.
Additional Links: PMID-43033655
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43033655,
year = {2026},
author = {Huang, M and Zhou, J and Xiong, J and Wan, Q and Ma, J},
title = {Removal performance and putative degradation mechanisms of bisphenol A in bioretention cells.},
journal = {Journal of environmental management},
volume = {420},
number = {},
pages = {131383},
doi = {10.1016/j.jenvman.2026.131383},
pmid = {43033655},
issn = {1095-8630},
abstract = {Bisphenol A (BPA) is a typical endocrine-disrupting pollutant. It can enter receiving waters through urban stormwater runoff and pose potential risks to aquatic ecosystems. Bioretention cells are widely used stormwater control measures for urban runoff and can retain and transform organic pollutants. However, the removal performance and mechanisms of BPA in bioretention cells remain unclear. In this study, two bioretention cells were constructed. BPA removal under different influent concentrations was investigated. The main removal zone and transformation process of BPA were also analyzed. The results showed that BPA removal by the bioretention cells exceeded 70%, with the main removal occurring in the upper filter media layer. Both filter media adsorption and microbial processes contributed to BPA removal. Several transformation products in the bioretention cells were putatively identified by liquid chromatography-mass spectrometry (LC-MS), including monohydroxylated BPA, 4-hydroxyacetophenone, 3-oxohexanedioic acid, and other intermediates. Together with the metagenomic results, these products suggest that BPA may undergo initial oxidation, side-chain cleavage, intermediate transformation, and ring-opening processes in bioretention cells. These findings provide a scientific basis for BPA control in urban stormwater runoff.},
}
RevDate: 2026-11-14
Spatial heterogeneity of sediment reducing capacity and its control on internal phosphorus cycling from a multi-ecotype eutrophic lake (Lake Taihu, China).
Water research, 309(Pt B):127321 pii:S0043-1354(26)01992-5 [Epub ahead of print].
Internal phosphorus (P) release from sediments constrains the ecological restoration of eutrophic lakes. However, how sediment reducing capacity (RC) is associated with P mobilization across contrasting lake ecotypes remains poorly understood. Here, we investigated the spatial heterogeneity of sediment RC, P mobility, microbial community composition, and metagenomic functional potential across algae- and macrophyte-dominated regions of Lake Taihu, China. The results indicate that algae-dominated sediments were characterized by higher RCpH7/RCpH2 ratios, elevated sulfide (S[n]) levels, and greater P mobilization, whereas macrophyte-dominated sediments generally exhibited lower RCpH7/RCpH2 ratios, higher Fe[2+] availability, and stronger P retention. Accordingly, the two ecotypes were classified as ROM-S[n] and ROM-Fe[2+] types, respectively. The maximum P adsorption capacity (Qm) in algae-dominated zones was approximately 0.6 times that in macrophyte-dominated zones, while the equilibrium P concentration (EPC0) and soluble reactive P (SRP) flux were 16.5 and 6.92 times higher, respectively. Microbial community patterns and metagenomic functional profiles further revealed greater potential for carbon metabolism, Fe/S reduction, and other anaerobic processes in algae-dominated sediments, favoring the activation of Fe-P. In contrast, the high Fe (II):P molar ratios, low S[n] levels, and greater Ca-P content in macrophyte-dominated sediments may favor Fe(Ⅱ)- and Ca-mediated P immobilization, thereby strengthening sediment P retention. Collectively, these findings suggest that RCpH7/RCpH2 ratio can serve as an operational indicator of sediment RC composition and associated redox characteristics, helping characterize contrasting P mobilization and retention patterns across lake ecotypes.
Additional Links: PMID-43033768
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43033768,
year = {2026},
author = {Lv, Y and Li, F and Zhang, L and Li, Y and Zhao, X and Lei, Q and Yun, W and Yin, H},
title = {Spatial heterogeneity of sediment reducing capacity and its control on internal phosphorus cycling from a multi-ecotype eutrophic lake (Lake Taihu, China).},
journal = {Water research},
volume = {309},
number = {Pt B},
pages = {127321},
doi = {10.1016/j.watres.2026.127321},
pmid = {43033768},
issn = {1879-2448},
abstract = {Internal phosphorus (P) release from sediments constrains the ecological restoration of eutrophic lakes. However, how sediment reducing capacity (RC) is associated with P mobilization across contrasting lake ecotypes remains poorly understood. Here, we investigated the spatial heterogeneity of sediment RC, P mobility, microbial community composition, and metagenomic functional potential across algae- and macrophyte-dominated regions of Lake Taihu, China. The results indicate that algae-dominated sediments were characterized by higher RCpH7/RCpH2 ratios, elevated sulfide (S[n]) levels, and greater P mobilization, whereas macrophyte-dominated sediments generally exhibited lower RCpH7/RCpH2 ratios, higher Fe[2+] availability, and stronger P retention. Accordingly, the two ecotypes were classified as ROM-S[n] and ROM-Fe[2+] types, respectively. The maximum P adsorption capacity (Qm) in algae-dominated zones was approximately 0.6 times that in macrophyte-dominated zones, while the equilibrium P concentration (EPC0) and soluble reactive P (SRP) flux were 16.5 and 6.92 times higher, respectively. Microbial community patterns and metagenomic functional profiles further revealed greater potential for carbon metabolism, Fe/S reduction, and other anaerobic processes in algae-dominated sediments, favoring the activation of Fe-P. In contrast, the high Fe (II):P molar ratios, low S[n] levels, and greater Ca-P content in macrophyte-dominated sediments may favor Fe(Ⅱ)- and Ca-mediated P immobilization, thereby strengthening sediment P retention. Collectively, these findings suggest that RCpH7/RCpH2 ratio can serve as an operational indicator of sediment RC composition and associated redox characteristics, helping characterize contrasting P mobilization and retention patterns across lake ecotypes.},
}
RevDate: 2026-11-14
Intervertebral disc versus vertebral body as the percutaneous sample site for pyogenic spondylodiscitis: Which yields a higher microbiological diagnosis? A prospective study.
The spine journal : official journal of the North American Spine Society pii:S1529-9430(26)00751-5 [Epub ahead of print].
BACKGROUND CONTEXT: Pyogenic spondylodiscitis requires microbiological confirmation, yet percutaneous sample collection yield remains modest. The optimal sampling site in patients without drainable abscess is controversial. Retrospective studies suggest the disc may outperform the vertebral body, but evidence is limited by design biases, leaving the question unresolved.
PURPOSE: To determine whether the intervertebral disc or the vertebral body provides a better percutaneous biopsy site for microbiological diagnosis in patients with pyogenic spondylodiscitis.
STUDY DESIGN: Prospective cohort study conducted at a single spine center.
PATIENT SAMPLE: A total of 68 patients with high clinical suspicion of pyogenic spondylodiscitis who underwent C-arm guided percutaneous biopsy between January 2024 and April 2025.
OUTCOME MEASURES: Positive rate of bacterial culture (including aerobic and anaerobic) from biopsy specimens; factors associated with positive culture using generalized estimating equations (GEE).
METHODS: All enrolled patients underwent simultaneous percutaneous sample collections of both the vertebral body and the intervertebral disc under C-arm guidance. Specimens were sent for bacterial culture, anaerobic culture, metagenomic next-generation sequencing (mNGS), and histopathological examination. The McNemar test was used to compare paired culture positivity rates between the two sites. Multivariable analysis was performed using GEE to identify factors associated with positive cultures.
RESULTS: Of 279 screened patients with suspected spinal infections, 68 were included (39 males, 29 females; median age 64.0 years). The intervertebral disc yielded a significantly higher microbial identification rate than the vertebral body (35/68 [51.5%] vs. 22/68 [32.4%]; P=0.002). On multivariable analysis, factors significantly associated with positive cultures included male sex, elevated preoperative dynamic erythrocyte sedimentation rate (ESR), increased procalcitonin (PCT), elevated interleukin-6 (IL-6), histopathological evidence of acute inflammation, and biopsy site (all P<0.05). Notably, the intervertebral disc site remained independently associated with higher culture positivity.
CONCLUSIONS: In patients with pyogenic spondylodiscitis requiring solid-tissue sampling, percutaneous sampling of the intervertebral disc yields a significantly higher bacterial culture positivity rate than sampling of the vertebral body. Male sex, elevated ESR, PCT, IL-6, and acute inflammation on histopathology are also associated with positive cultures. These findings support prioritizing the intervertebral disc as the preferred percutaneous sampling target-reserving direct aspiration for accessible fluid collections-to optimize microbiological diagnosis in pyogenic spondylodiscitis.
Additional Links: PMID-43033832
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43033832,
year = {2026},
author = {Tang, K and Li, Y and He, Y and Fan, J and Yan, G and Li, Y and Wang, H and Zhu, Q and Lan, T and Dong, W},
title = {Intervertebral disc versus vertebral body as the percutaneous sample site for pyogenic spondylodiscitis: Which yields a higher microbiological diagnosis? A prospective study.},
journal = {The spine journal : official journal of the North American Spine Society},
volume = {},
number = {},
pages = {},
doi = {10.1016/j.spinee.2026.11.004},
pmid = {43033832},
issn = {1878-1632},
abstract = {BACKGROUND CONTEXT: Pyogenic spondylodiscitis requires microbiological confirmation, yet percutaneous sample collection yield remains modest. The optimal sampling site in patients without drainable abscess is controversial. Retrospective studies suggest the disc may outperform the vertebral body, but evidence is limited by design biases, leaving the question unresolved.
PURPOSE: To determine whether the intervertebral disc or the vertebral body provides a better percutaneous biopsy site for microbiological diagnosis in patients with pyogenic spondylodiscitis.
STUDY DESIGN: Prospective cohort study conducted at a single spine center.
PATIENT SAMPLE: A total of 68 patients with high clinical suspicion of pyogenic spondylodiscitis who underwent C-arm guided percutaneous biopsy between January 2024 and April 2025.
OUTCOME MEASURES: Positive rate of bacterial culture (including aerobic and anaerobic) from biopsy specimens; factors associated with positive culture using generalized estimating equations (GEE).
METHODS: All enrolled patients underwent simultaneous percutaneous sample collections of both the vertebral body and the intervertebral disc under C-arm guidance. Specimens were sent for bacterial culture, anaerobic culture, metagenomic next-generation sequencing (mNGS), and histopathological examination. The McNemar test was used to compare paired culture positivity rates between the two sites. Multivariable analysis was performed using GEE to identify factors associated with positive cultures.
RESULTS: Of 279 screened patients with suspected spinal infections, 68 were included (39 males, 29 females; median age 64.0 years). The intervertebral disc yielded a significantly higher microbial identification rate than the vertebral body (35/68 [51.5%] vs. 22/68 [32.4%]; P=0.002). On multivariable analysis, factors significantly associated with positive cultures included male sex, elevated preoperative dynamic erythrocyte sedimentation rate (ESR), increased procalcitonin (PCT), elevated interleukin-6 (IL-6), histopathological evidence of acute inflammation, and biopsy site (all P<0.05). Notably, the intervertebral disc site remained independently associated with higher culture positivity.
CONCLUSIONS: In patients with pyogenic spondylodiscitis requiring solid-tissue sampling, percutaneous sampling of the intervertebral disc yields a significantly higher bacterial culture positivity rate than sampling of the vertebral body. Male sex, elevated ESR, PCT, IL-6, and acute inflammation on histopathology are also associated with positive cultures. These findings support prioritizing the intervertebral disc as the preferred percutaneous sampling target-reserving direct aspiration for accessible fluid collections-to optimize microbiological diagnosis in pyogenic spondylodiscitis.},
}
RevDate: 2026-11-14
Metagenomic insights into regulatory effects of micro-aerobic conditions on microbial metabolism during hydrolytic acidification of printing and dyeing wastewater.
Bioresource technology pii:S0960-8524(26)02323-0 [Epub ahead of print].
Micro-aerobic hydrolysis and acidification (MAHA) is a promising pretreatment for printing and dyeing wastewater (PDW), yet the role of dissolved oxygen (DO) in regulating hydrolysis-acidification performance and microbial functions remains poorly understood. In this study, laboratory-scale reactors treating synthetic PDW were operated under four DO ranges to investigate changes in reactor performance, microbial communities, and metagenomic functional potential. A DO of 0.1-0.3 mg/L achieved the best overall performance, with an apparent decolorization efficiency of 77.0 ± 7.5 %, a BOD5/COD ratio of 0.58 ± 0.029, and a COD removal efficiency of 41.4 ± 5.5 %. Acetate was the predominant volatile fatty acid (VFA), whereas VFA accumulation decreased with increasing DO. Microbial community analysis revealed distinct community structures under different DO conditions, with several potential hydrolytic and organic matter-transforming genera enriched at 0.1-0.3 mg/L DO. Metagenomic analysis further indicated enhanced functional potential for central carbon metabolism, electron transfer, redox cofactor metabolism, and aromatic compound transformation under this condition. Notably, azo-reductase-related genes did not increase concomitantly, suggesting that the improved apparent decolorization was likely associated with broader community-level functional potential rather than enrichment of a single azo-reduction gene. Overall, maintaining DO at 0.1-0.3 mg/L was associated with enhanced apparent decolorization, providing operational insights for further optimization of MAHA under the investigated conditions.
Additional Links: PMID-43033992
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43033992,
year = {2026},
author = {Ren, J and Peng, Q and Du, Z and Xie, Q and Hui, J and Cheng, W},
title = {Metagenomic insights into regulatory effects of micro-aerobic conditions on microbial metabolism during hydrolytic acidification of printing and dyeing wastewater.},
journal = {Bioresource technology},
volume = {},
number = {},
pages = {136241},
doi = {10.1016/j.biortech.2026.136241},
pmid = {43033992},
issn = {1873-2976},
abstract = {Micro-aerobic hydrolysis and acidification (MAHA) is a promising pretreatment for printing and dyeing wastewater (PDW), yet the role of dissolved oxygen (DO) in regulating hydrolysis-acidification performance and microbial functions remains poorly understood. In this study, laboratory-scale reactors treating synthetic PDW were operated under four DO ranges to investigate changes in reactor performance, microbial communities, and metagenomic functional potential. A DO of 0.1-0.3 mg/L achieved the best overall performance, with an apparent decolorization efficiency of 77.0 ± 7.5 %, a BOD5/COD ratio of 0.58 ± 0.029, and a COD removal efficiency of 41.4 ± 5.5 %. Acetate was the predominant volatile fatty acid (VFA), whereas VFA accumulation decreased with increasing DO. Microbial community analysis revealed distinct community structures under different DO conditions, with several potential hydrolytic and organic matter-transforming genera enriched at 0.1-0.3 mg/L DO. Metagenomic analysis further indicated enhanced functional potential for central carbon metabolism, electron transfer, redox cofactor metabolism, and aromatic compound transformation under this condition. Notably, azo-reductase-related genes did not increase concomitantly, suggesting that the improved apparent decolorization was likely associated with broader community-level functional potential rather than enrichment of a single azo-reduction gene. Overall, maintaining DO at 0.1-0.3 mg/L was associated with enhanced apparent decolorization, providing operational insights for further optimization of MAHA under the investigated conditions.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-15
Microbial community multi-omic analysis of marsh sediment following enrichment with crustacean shell compost.
ISME communications, 6(1):ycag307.
Nutrient availability influences microbial processes in natural environments, notably biogeochemical cycling and community structure. Short-term, functional responses of microbial communities to nutrient pulses in coastal communities are not fully understood, especially in temperate environments undergoing climate driven changes. A 48 hour microcosm experiment was carried out in which paired metagenomic and metatranscriptomic sequencing was employed to elucidate how decomposition of chitin rich substrates, namely, crab and lobster shell compost, alter salt marsh microbiome structure and function. Within 48 hours of amendment, a notable shift in community function was observed, including increased chitin degradation, altered stress-response, and enhanced sporulation. These responses coincided with marked decreases in genes regulating key biogeochemical processes, including carbon fixation, sulfur oxidation and reduction, and other metabolic pathways. After addition of shell compost, changes in microbial community structure and increased incidence of virulence-associated genes were also observed, accompanied by measurable transcriptional activation, notably aerolysin A (aerA), which encodes the pore-forming exotoxin aerolysin. These results support shifts in both community composition and function in temperate salt marsh sediment microbiomes following chitin-rich nutrient addition. It is concluded that temperate coastal systems are sensitive to organic nutrient enrichment, with the potential for ecological and public-health relevant consequences. These effects may be particularly significant given chitin is among the most abundant and readily available bionutrients in aquatic ecosystems.
Additional Links: PMID-43034448
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43034448,
year = {2026},
author = {Yancey, CE and Brumfield, KD and Ettwiller, L and Colwell, RR},
title = {Microbial community multi-omic analysis of marsh sediment following enrichment with crustacean shell compost.},
journal = {ISME communications},
volume = {6},
number = {1},
pages = {ycag307},
pmid = {43034448},
issn = {2730-6151},
abstract = {Nutrient availability influences microbial processes in natural environments, notably biogeochemical cycling and community structure. Short-term, functional responses of microbial communities to nutrient pulses in coastal communities are not fully understood, especially in temperate environments undergoing climate driven changes. A 48 hour microcosm experiment was carried out in which paired metagenomic and metatranscriptomic sequencing was employed to elucidate how decomposition of chitin rich substrates, namely, crab and lobster shell compost, alter salt marsh microbiome structure and function. Within 48 hours of amendment, a notable shift in community function was observed, including increased chitin degradation, altered stress-response, and enhanced sporulation. These responses coincided with marked decreases in genes regulating key biogeochemical processes, including carbon fixation, sulfur oxidation and reduction, and other metabolic pathways. After addition of shell compost, changes in microbial community structure and increased incidence of virulence-associated genes were also observed, accompanied by measurable transcriptional activation, notably aerolysin A (aerA), which encodes the pore-forming exotoxin aerolysin. These results support shifts in both community composition and function in temperate salt marsh sediment microbiomes following chitin-rich nutrient addition. It is concluded that temperate coastal systems are sensitive to organic nutrient enrichment, with the potential for ecological and public-health relevant consequences. These effects may be particularly significant given chitin is among the most abundant and readily available bionutrients in aquatic ecosystems.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-15
Probiotics and Prebiotics in Restoring Gastrointestinal Microbiota Dysbiosis: Emerging Alternatives to Antibiotic Therapy.
International journal of food science, 2026:6956918.
The human gastrointestinal (GI) microbiota is a complex and dynamic microbial ecosystem that plays a fundamental role in maintaining host metabolism, immune homeostasis, intestinal barrier integrity, and protection against pathogenic microorganisms. Disruption of this microbial community, known as gut dysbiosis, has been implicated in the pathogenesis of numerous GI and systemic diseases, including inflammatory bowel disease (IBD), irritable bowel syndrome (IBS), Clostridioides difficile infection, obesity, type 2 diabetes mellitus, colorectal cancer, and nonalcoholic fatty liver disease. Although antibiotics remain indispensable for the treatment of bacterial infections, their indiscriminate use frequently disrupts gut microbial composition, reduces microbial diversity, promotes opportunistic pathogen overgrowth, and accelerates the emergence of antimicrobial resistance. These limitations have stimulated increasing interest in microbiota-targeted interventions as safer and more sustainable therapeutic strategies. Probiotics and prebiotics have emerged as promising approaches for restoring gut microbial homeostasis through complementary mechanisms. Probiotics inhibit pathogen colonization, enhance intestinal barrier function, produce antimicrobial metabolites, and modulate host immune responses. Prebiotics selectively stimulate the growth and metabolic activity of beneficial microorganisms, promoting the production of short-chain fatty acids (SCFAs) that support intestinal and systemic health. Increasing evidence from experimental studies, randomized clinical trials, and meta-analyses demonstrates that these microbiota-directed interventions can reduce antibiotic-associated diarrhea, improve symptoms of IBD and IBS, enhance Helicobacter pylori eradication, decrease recurrence of C. difficile infection, and contribute to the management of several metabolic disorders. Despite these promising findings, important challenges remain regarding strain specificity, optimal dosage, treatment duration, product standardization, regulatory oversight, and interindividual variability in treatment response. Emerging advances in metagenomics, metabolomics, artificial intelligence, and precision medicine are facilitating the development of personalized microbiome-based therapies that may further enhance clinical outcomes. This review critically summarizes the current understanding of GI microbiota dysbiosis, discusses the mechanisms through which probiotics and prebiotics restore microbial balance, evaluates their clinical applications as alternatives or adjuncts to antibiotic therapy, and highlights existing challenges and future research directions for microbiome-based therapeutics.
Additional Links: PMID-43034624
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43034624,
year = {2026},
author = {Goa, T and Wolde, T and Sahilu, E},
title = {Probiotics and Prebiotics in Restoring Gastrointestinal Microbiota Dysbiosis: Emerging Alternatives to Antibiotic Therapy.},
journal = {International journal of food science},
volume = {2026},
number = {},
pages = {6956918},
pmid = {43034624},
issn = {2314-5765},
abstract = {The human gastrointestinal (GI) microbiota is a complex and dynamic microbial ecosystem that plays a fundamental role in maintaining host metabolism, immune homeostasis, intestinal barrier integrity, and protection against pathogenic microorganisms. Disruption of this microbial community, known as gut dysbiosis, has been implicated in the pathogenesis of numerous GI and systemic diseases, including inflammatory bowel disease (IBD), irritable bowel syndrome (IBS), Clostridioides difficile infection, obesity, type 2 diabetes mellitus, colorectal cancer, and nonalcoholic fatty liver disease. Although antibiotics remain indispensable for the treatment of bacterial infections, their indiscriminate use frequently disrupts gut microbial composition, reduces microbial diversity, promotes opportunistic pathogen overgrowth, and accelerates the emergence of antimicrobial resistance. These limitations have stimulated increasing interest in microbiota-targeted interventions as safer and more sustainable therapeutic strategies. Probiotics and prebiotics have emerged as promising approaches for restoring gut microbial homeostasis through complementary mechanisms. Probiotics inhibit pathogen colonization, enhance intestinal barrier function, produce antimicrobial metabolites, and modulate host immune responses. Prebiotics selectively stimulate the growth and metabolic activity of beneficial microorganisms, promoting the production of short-chain fatty acids (SCFAs) that support intestinal and systemic health. Increasing evidence from experimental studies, randomized clinical trials, and meta-analyses demonstrates that these microbiota-directed interventions can reduce antibiotic-associated diarrhea, improve symptoms of IBD and IBS, enhance Helicobacter pylori eradication, decrease recurrence of C. difficile infection, and contribute to the management of several metabolic disorders. Despite these promising findings, important challenges remain regarding strain specificity, optimal dosage, treatment duration, product standardization, regulatory oversight, and interindividual variability in treatment response. Emerging advances in metagenomics, metabolomics, artificial intelligence, and precision medicine are facilitating the development of personalized microbiome-based therapies that may further enhance clinical outcomes. This review critically summarizes the current understanding of GI microbiota dysbiosis, discusses the mechanisms through which probiotics and prebiotics restore microbial balance, evaluates their clinical applications as alternatives or adjuncts to antibiotic therapy, and highlights existing challenges and future research directions for microbiome-based therapeutics.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-15
Genome skimming of Poecilosclerida (Porifera) reveals conserved and hyperrearranged mitochondrial genomes.
PeerJ, 14:e21781.
Genome skimming is a cost-effective approach that efficiently recovers multi-copy genomic regions, making it particularly valuable for phylogenetic research and biodiversity assessments. In this study, we applied shallow shotgun metagenome sequencing to 15 poecilosclerid sponges, successfully assembling complete mitochondrial genomes for four species and full rDNA clusters for the majority of samples, despite considerable variation in the retrieval of genomic regions and exogenous, non-poriferan sequences. Phylogenetic reconstructions using both rDNA and mtDNA sequences uncovered extensive polyphyly within several key families, such as Coelosphaeridae and Mycalidae, underscoring significant systematic inconsistencies at generic, familial, and ordinal levels, and emphasizing the need for comprehensive taxonomic revision throughout Poecilosclerida to reflect natural groups. The mitogenomes obtained from Clathria (Microciona) sp. (18,725 bp), Desmapsamma anchorata (19,099 bp), Latrunculia biformis (19,630 bp), and Neopodospongia tupecomareni (19,983 bp) exhibited conserved architectural features typical of the order, alongside notable variations such as gene overlaps, non-canonical start codons, and an unusual fused nd3::nd4L gene. Our findings highlight the applicability of genome skimming for phylogenetics and metagenomic insights, though success can vary due to biological and methodological factors. This approach has proven to be a cost-effective and powerful tool for advancing sponge systematics, despite challenges in assembling hyperrearranged mitogenomes in certain lineages, paving the way for broader genomic surveys in non-model organisms.
Additional Links: PMID-43034769
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43034769,
year = {2026},
author = {de Paula, TS and Leite, DMB and Thompson, F and Hajdu, E},
title = {Genome skimming of Poecilosclerida (Porifera) reveals conserved and hyperrearranged mitochondrial genomes.},
journal = {PeerJ},
volume = {14},
number = {},
pages = {e21781},
pmid = {43034769},
issn = {2167-8359},
mesh = {Animals ; *Genome, Mitochondrial ; *Porifera/genetics/classification ; Phylogeny ; Shotgun Sequencing ; DNA, Mitochondrial/genetics ; },
abstract = {Genome skimming is a cost-effective approach that efficiently recovers multi-copy genomic regions, making it particularly valuable for phylogenetic research and biodiversity assessments. In this study, we applied shallow shotgun metagenome sequencing to 15 poecilosclerid sponges, successfully assembling complete mitochondrial genomes for four species and full rDNA clusters for the majority of samples, despite considerable variation in the retrieval of genomic regions and exogenous, non-poriferan sequences. Phylogenetic reconstructions using both rDNA and mtDNA sequences uncovered extensive polyphyly within several key families, such as Coelosphaeridae and Mycalidae, underscoring significant systematic inconsistencies at generic, familial, and ordinal levels, and emphasizing the need for comprehensive taxonomic revision throughout Poecilosclerida to reflect natural groups. The mitogenomes obtained from Clathria (Microciona) sp. (18,725 bp), Desmapsamma anchorata (19,099 bp), Latrunculia biformis (19,630 bp), and Neopodospongia tupecomareni (19,983 bp) exhibited conserved architectural features typical of the order, alongside notable variations such as gene overlaps, non-canonical start codons, and an unusual fused nd3::nd4L gene. Our findings highlight the applicability of genome skimming for phylogenetics and metagenomic insights, though success can vary due to biological and methodological factors. This approach has proven to be a cost-effective and powerful tool for advancing sponge systematics, despite challenges in assembling hyperrearranged mitogenomes in certain lineages, paving the way for broader genomic surveys in non-model organisms.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Genome, Mitochondrial
*Porifera/genetics/classification
Phylogeny
Shotgun Sequencing
DNA, Mitochondrial/genetics
RevDate: 2026-11-15
Thermophilic intensification governs abiotic fragmentation and microbiome restructuring during anaerobic co-digestion of bioplastics with OFMSW and waste activated sludge.
Journal of environmental management, 420:131382 pii:S0301-4797(26)02842-2 [Epub ahead of print].
Anaerobic co-digestion (Co-AD) of the organic fraction of municipal solid waste (OFMSW) and waste-activated sludge (WAS) is a promising strategy for waste valorisation and renewable energy recovery; however, the increasing presence of compostable bioplastics in organic waste streams raises concerns regarding their degradation behaviour and impact on anaerobic digestion (AD) performance. In this study, semi-continuous mesophilic-to-thermophilic Co-AD of OFMSW and WAS containing Mater-Bi, crystallised polylactic acid (CPLA), and polyhydroxyalkanoate (PHA) was investigated under progressively intensified operating conditions, with and without graphite supplementation. Reactor performance was monitored through methane production, volatile fatty acids (VFAs), pH, and FOS/TAC ratio, while complementary abiotic assays and ATR-FTIR analyses were used to assess physicochemical polymer transformation in the absence and presence of microbial activity. Long-read metagenomics was further applied to evaluate microbial community restructuring during thermophilic operation. Thermophilic intensification strongly affected polymer transformation pathways and microbial composition. Among the tested materials, CPLA showed the highest abiotic solubilisation at 55 °C, reaching 1280 mg COD L[-1] and exhibiting clear fragmentation under thermophilic conditions. In contrast, P(3HB-co-4HB) showed limited solubilisation and structural transformation. Graphite supplementation produced limited and condition-dependent effects on reactor performance. Metagenomic analysis revealed a marked transition from mesophilic Bacillota-dominated communities toward thermophilic enrichment of Patescibacteria, Thermotogota, and syntrophy-associated Pseudomonadota taxa. Overall, thermophilic intensification was associated with material-specific physicochemical transformation and marked microbial community restructuring, highlighting parallel responses of bioplastics and the anaerobic microbiome to changing operating conditions.
Additional Links: PMID-43035085
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43035085,
year = {2026},
author = {Pedullà, A and Pavlović, J and Ferreri, M and Pangallo, D and Galová, D and Pangallo, D and Kraková, L and Calabrò, PS},
title = {Thermophilic intensification governs abiotic fragmentation and microbiome restructuring during anaerobic co-digestion of bioplastics with OFMSW and waste activated sludge.},
journal = {Journal of environmental management},
volume = {420},
number = {},
pages = {131382},
doi = {10.1016/j.jenvman.2026.131382},
pmid = {43035085},
issn = {1095-8630},
abstract = {Anaerobic co-digestion (Co-AD) of the organic fraction of municipal solid waste (OFMSW) and waste-activated sludge (WAS) is a promising strategy for waste valorisation and renewable energy recovery; however, the increasing presence of compostable bioplastics in organic waste streams raises concerns regarding their degradation behaviour and impact on anaerobic digestion (AD) performance. In this study, semi-continuous mesophilic-to-thermophilic Co-AD of OFMSW and WAS containing Mater-Bi, crystallised polylactic acid (CPLA), and polyhydroxyalkanoate (PHA) was investigated under progressively intensified operating conditions, with and without graphite supplementation. Reactor performance was monitored through methane production, volatile fatty acids (VFAs), pH, and FOS/TAC ratio, while complementary abiotic assays and ATR-FTIR analyses were used to assess physicochemical polymer transformation in the absence and presence of microbial activity. Long-read metagenomics was further applied to evaluate microbial community restructuring during thermophilic operation. Thermophilic intensification strongly affected polymer transformation pathways and microbial composition. Among the tested materials, CPLA showed the highest abiotic solubilisation at 55 °C, reaching 1280 mg COD L[-1] and exhibiting clear fragmentation under thermophilic conditions. In contrast, P(3HB-co-4HB) showed limited solubilisation and structural transformation. Graphite supplementation produced limited and condition-dependent effects on reactor performance. Metagenomic analysis revealed a marked transition from mesophilic Bacillota-dominated communities toward thermophilic enrichment of Patescibacteria, Thermotogota, and syntrophy-associated Pseudomonadota taxa. Overall, thermophilic intensification was associated with material-specific physicochemical transformation and marked microbial community restructuring, highlighting parallel responses of bioplastics and the anaerobic microbiome to changing operating conditions.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-15
[The 522nd case: fever of unknown origin and low back pain].
Zhonghua nei ke za zhi, 65(11):1266-1270.
A 72-year-old man presented with acute low back pain accompanied by high fever and chills. During the first hospitalization, one anaerobic blood culture bottle became positive after 120 hours of incubation and was identified as Parvimonas micra. The isolate was initially considered a contaminant. The patient received empirical antimicrobial therapy with meropenem for one week, after which his symptoms improved and he was discharged. However, the patient subsequently developed progressively worsening back pain. Lumbar magnetic resonance imaging revealed L2-L3 vertebral osteomyelitis with paravertebral abscess formation. After readmission, a vertebral biopsy was performed, and both tissue culture and metagenomic next-generation sequencing (mNGS) identified P. micra. The final diagnosis was lumbar vertebral osteomyelitis with psoas abscess caused by P. micra. Infection caused by P. micra is often insidious in onset, and positive blood culture results may be misinterpreted as contamination. This case highlights that anaerobic infection should be considered in patients with unexplained fever and focal bone pain, and that clinical manifestations, imaging findings, and microbiological evidence should be integrated to establish the diagnosis. This rare case provides useful insights for clinical practice.
Additional Links: PMID-43035275
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43035275,
year = {2026},
author = {Gong, CC and Huang, ZX and Guo, FP and Zhou, BT and Li, J and Li, TS and Liu, ZY},
title = {[The 522nd case: fever of unknown origin and low back pain].},
journal = {Zhonghua nei ke za zhi},
volume = {65},
number = {11},
pages = {1266-1270},
doi = {10.3760/cma.j.cn112138-20260527-00318},
pmid = {43035275},
issn = {0578-1426},
support = {2022-PUMCH-B-043//Peking Union Medical College Hospital Central High Level Hospital Clinical Research Project/ ; },
mesh = {Humans ; Male ; Aged ; *Fever of Unknown Origin ; *Low Back Pain/microbiology ; Osteomyelitis ; Bacillota ; *Gram-Positive Bacterial Infections ; },
abstract = {A 72-year-old man presented with acute low back pain accompanied by high fever and chills. During the first hospitalization, one anaerobic blood culture bottle became positive after 120 hours of incubation and was identified as Parvimonas micra. The isolate was initially considered a contaminant. The patient received empirical antimicrobial therapy with meropenem for one week, after which his symptoms improved and he was discharged. However, the patient subsequently developed progressively worsening back pain. Lumbar magnetic resonance imaging revealed L2-L3 vertebral osteomyelitis with paravertebral abscess formation. After readmission, a vertebral biopsy was performed, and both tissue culture and metagenomic next-generation sequencing (mNGS) identified P. micra. The final diagnosis was lumbar vertebral osteomyelitis with psoas abscess caused by P. micra. Infection caused by P. micra is often insidious in onset, and positive blood culture results may be misinterpreted as contamination. This case highlights that anaerobic infection should be considered in patients with unexplained fever and focal bone pain, and that clinical manifestations, imaging findings, and microbiological evidence should be integrated to establish the diagnosis. This rare case provides useful insights for clinical practice.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
Male
Aged
*Fever of Unknown Origin
*Low Back Pain/microbiology
Osteomyelitis
Bacillota
*Gram-Positive Bacterial Infections
RevDate: 2026-11-15
CmpDate: 2026-11-15
Precision Prediction of Microbial Ecosystem Impact on Host Metabolism Using Genome-Resolved Metagenomics.
NPJ biofilms and microbiomes, 12(1):.
Mammalian gut dysbiosis is recognized to influence host metabolism, yet key microbiota and mechanisms governing their effects remain poorly understood. Here we developed a genome-resolved ecology-aware, systems-level workflow to predict how gut microbial metabolism affects mammalian health, and we apply it to a "spinal cord-gut axis" dataset. By scaling and integrating temporally resolved network analytics and consensus statistical approaches, we identified 19 microbial species that best predict host physiology following neurological impairment. In silico validation through pathway-centric and comparative genomic analyses revealed that among these species, the biggest encoded microbial metabolic changes were in pathways linked to host nitrogen balance, varying by host sex and microbial ecotype/species. Further inference identified the specific bacteria (and their draft genomes) potentially driving urease-dependent versus amino acid-dependent nitrogen metabolism-findings that can explain previously mechanistically-ambiguous, but clinically relevant, ammonia-driven host nitrogen imbalance. This provides a generalizable data-driven hypothesis generation workflow for longitudinal microbiome data.
Additional Links: PMID-43035523
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43035523,
year = {2026},
author = {Mohssen, M and Zayed, AA and Kigerl, KA and Du, J and Smith, GJ and Schwab, JM and Sullivan, MB and Popovich, PG},
title = {Precision Prediction of Microbial Ecosystem Impact on Host Metabolism Using Genome-Resolved Metagenomics.},
journal = {NPJ biofilms and microbiomes},
volume = {12},
number = {1},
pages = {},
pmid = {43035523},
issn = {2055-5008},
support = {890085//Craig H. Nielsen Foundation/ ; ABI#2149505 and DBI#2022070//National Science Foundation/ ; 1R35NS111582//National Institutes of Neurological Disorders and Stroke R35/ ; },
mesh = {*Metagenomics/methods ; Animals ; Nitrogen/metabolism ; *Bacteria/genetics/classification/metabolism ; *Microbiota ; Genome, Bacterial ; Ecosystem ; Humans ; Metagenome ; Host Microbial Interactions ; },
abstract = {Mammalian gut dysbiosis is recognized to influence host metabolism, yet key microbiota and mechanisms governing their effects remain poorly understood. Here we developed a genome-resolved ecology-aware, systems-level workflow to predict how gut microbial metabolism affects mammalian health, and we apply it to a "spinal cord-gut axis" dataset. By scaling and integrating temporally resolved network analytics and consensus statistical approaches, we identified 19 microbial species that best predict host physiology following neurological impairment. In silico validation through pathway-centric and comparative genomic analyses revealed that among these species, the biggest encoded microbial metabolic changes were in pathways linked to host nitrogen balance, varying by host sex and microbial ecotype/species. Further inference identified the specific bacteria (and their draft genomes) potentially driving urease-dependent versus amino acid-dependent nitrogen metabolism-findings that can explain previously mechanistically-ambiguous, but clinically relevant, ammonia-driven host nitrogen imbalance. This provides a generalizable data-driven hypothesis generation workflow for longitudinal microbiome data.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Metagenomics/methods
Animals
Nitrogen/metabolism
*Bacteria/genetics/classification/metabolism
*Microbiota
Genome, Bacterial
Ecosystem
Humans
Metagenome
Host Microbial Interactions
RevDate: 2026-11-15
CmpDate: 2026-11-16
Population genomics of the endangered Peruvian algarrobo (Neltuma pallida) reveals low genetic diversity and a recent bottleneck in a protected dry-forest population.
BMC genomics, 27(1):.
The Pomac Forest Historic Sanctuary (SHBP), a protected area on the northern coast of Peru, conserves biodiversity representative of Equatorial dry forests. SHBP is mainly composed of algarrobo (Neltuma pallida), a species adapted to the variable climate conditions and known for its multiple ecological functions and ecosystem services. Algarrobos face an unprecedented population decline despite their remarkable natural adaptation, worsened by their ongoing habitat degradation and overexploitation. In this study, we aim to characterize the genomic diversity of the protected forest in the SHBP to establish a baseline for future monitoring and research. Results show that the algarrobo population in the SHBP is unstructured, has low inbreeding indices, an average autosomal heterozygosity of 0.19%, and a per-site nucleotide diversity of 0.0023. Patterns of low genetic diversity are associated with a recent bottleneck decline starting around four generations ago. Potentially deleterious variants with varying effects, possibly playing a role in population adaptability were predicted. In conclusion, our results provide reference genetic indices for future monitoring of algarrobo in its native range and raises questions about the impact of recent habitat degradation on its genome.
Additional Links: PMID-43035642
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43035642,
year = {2026},
author = {La Torre Ramirez, RD and Caycho, E and Saucedo-Bazalar, M and Saldaña, CL and Bieker, VC and Arbizu, CI and Orjeda, G},
title = {Population genomics of the endangered Peruvian algarrobo (Neltuma pallida) reveals low genetic diversity and a recent bottleneck in a protected dry-forest population.},
journal = {BMC genomics},
volume = {27},
number = {1},
pages = {},
pmid = {43035642},
issn = {1471-2164},
support = {Beca de investigación a corto plazo, "Genómica poblacional de Prosopis pallida mediante la técnica de genotipado por secuenciación", Convenio N°034-2022-PRFNP//Servicio Nacional de Áreas Naturales Protegidas por el Estado (SERNANP) y PROFONANPE/ ; Proyecto CIF, "Mejoramiento del Servicio de Promoción de la Ciencia, Tecnología e Innovación Tecnológica en el Centro de Investigación en Fruticultura "CIF" de la UNTRM - distrito Magdalena, de la provincia de Chachapoyas, del departamento de Amazonas" con CUI N°2590699 INDES-CES UNTRM//Universidad Nacional Toribio Rodríguez de Mendoza de Amazonas/ ; Proyecto B2110006i - Programa de Proyectos Interdisciplinarios para el Fomento de la Cooperación Interinstitucional en Investigación e Innovación (PNESTP-MINEDU-2021), R.R. N°009412-2021-R/UNMSM//Universidad Nacional Mayor de San Marcos/ ; },
mesh = {*Genetic Variation ; Peru ; *Endangered Species ; *Genetics, Population ; *Forests ; *Genomics ; Conservation of Natural Resources ; *Metagenomics ; },
abstract = {The Pomac Forest Historic Sanctuary (SHBP), a protected area on the northern coast of Peru, conserves biodiversity representative of Equatorial dry forests. SHBP is mainly composed of algarrobo (Neltuma pallida), a species adapted to the variable climate conditions and known for its multiple ecological functions and ecosystem services. Algarrobos face an unprecedented population decline despite their remarkable natural adaptation, worsened by their ongoing habitat degradation and overexploitation. In this study, we aim to characterize the genomic diversity of the protected forest in the SHBP to establish a baseline for future monitoring and research. Results show that the algarrobo population in the SHBP is unstructured, has low inbreeding indices, an average autosomal heterozygosity of 0.19%, and a per-site nucleotide diversity of 0.0023. Patterns of low genetic diversity are associated with a recent bottleneck decline starting around four generations ago. Potentially deleterious variants with varying effects, possibly playing a role in population adaptability were predicted. In conclusion, our results provide reference genetic indices for future monitoring of algarrobo in its native range and raises questions about the impact of recent habitat degradation on its genome.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Genetic Variation
Peru
*Endangered Species
*Genetics, Population
*Forests
*Genomics
Conservation of Natural Resources
*Metagenomics
RevDate: 2026-11-15
CmpDate: 2026-11-15
Seasonal dietary shifts drive gut microbiome and metabolome adaptation in wild Northern Pike (Esox lucius).
BMC microbiology, 26(1):.
The gut microbiome of fish faces challenges in maintaining stability during seasonal dietary shifts, which directly impacts host nutrition and fitness. In this study, environmental DNA metabarcoding, metagenomics, and metabolomics were employed to characterize seasonal differences in diet, gut microbiome composition, metabolites, and digestive enzyme activity of Esox lucius between spring and autumn. E. lucius exhibited broadly and opportunistically in spring but shifted to a markedly narrower in autumn. Microbiome composition exhibited marked seasonality, with spring dominated by Mycoplasma and Yersinia, whereas autumn were characterized by Aeromonas, Acinetobacter, environmental bacteria, and were accompanied by increased trypsin and lipase activities, which suggested a capacity for enhanced lipid and protein digestion for winter energy storage. Metabolomic analysis revealed enrichment of sphingolipid metabolites in autumn, which implicated in lipid metabolism and immune regulation. Furthermore, the autumn exhibited a greater diversity and abundance of antibiotic resistance genes. Our results demonstrated that seasonal dietary variation, digestive enzyme adaptation, microbiota reshaping, metabolite-mediated signaling collectively enabled E. lucius to regulate digestive physiology, as well as efficiently assimilating nutrients and maintaining energy balance in response to fluctuating resource availability in wild. Our study offers valuable insights into the ecological adaptation mechanisms of cold-water fish species.
Additional Links: PMID-43035695
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43035695,
year = {2026},
author = {Yang, Y and Wu, X and Feng, H and Zheng, Y and Wang, Z and Wu, R and Zhang, J},
title = {Seasonal dietary shifts drive gut microbiome and metabolome adaptation in wild Northern Pike (Esox lucius).},
journal = {BMC microbiology},
volume = {26},
number = {1},
pages = {},
pmid = {43035695},
issn = {1471-2180},
support = {2026D14009//the Tianshan Innovation Team Fund Project of Xinjiang Uygur Autonomous Region/ ; 2025B02020-2-9//the Key Research and Development Program in Xinjiang Uygur Autonomous Region/ ; 32560265//the National Natural Science Foundation of China/ ; },
mesh = {Animals ; Seasons ; *Metabolome ; *Bacteria/classification/genetics/isolation & purification/metabolism ; *Gastrointestinal Microbiome/physiology ; Adaptation, Physiological ; *Diet/veterinary ; Metabolomics ; Metagenomics ; },
abstract = {The gut microbiome of fish faces challenges in maintaining stability during seasonal dietary shifts, which directly impacts host nutrition and fitness. In this study, environmental DNA metabarcoding, metagenomics, and metabolomics were employed to characterize seasonal differences in diet, gut microbiome composition, metabolites, and digestive enzyme activity of Esox lucius between spring and autumn. E. lucius exhibited broadly and opportunistically in spring but shifted to a markedly narrower in autumn. Microbiome composition exhibited marked seasonality, with spring dominated by Mycoplasma and Yersinia, whereas autumn were characterized by Aeromonas, Acinetobacter, environmental bacteria, and were accompanied by increased trypsin and lipase activities, which suggested a capacity for enhanced lipid and protein digestion for winter energy storage. Metabolomic analysis revealed enrichment of sphingolipid metabolites in autumn, which implicated in lipid metabolism and immune regulation. Furthermore, the autumn exhibited a greater diversity and abundance of antibiotic resistance genes. Our results demonstrated that seasonal dietary variation, digestive enzyme adaptation, microbiota reshaping, metabolite-mediated signaling collectively enabled E. lucius to regulate digestive physiology, as well as efficiently assimilating nutrients and maintaining energy balance in response to fluctuating resource availability in wild. Our study offers valuable insights into the ecological adaptation mechanisms of cold-water fish species.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
Seasons
*Metabolome
*Bacteria/classification/genetics/isolation & purification/metabolism
*Gastrointestinal Microbiome/physiology
Adaptation, Physiological
*Diet/veterinary
Metabolomics
Metagenomics
RevDate: 2026-11-16
CmpDate: 2026-11-16
Gastro16: A Novel eDNA Metabarcoding Marker for Detecting Gastropod Diversity in Marine Environments.
Molecular ecology resources, 26(8):.
The diversity of gastropod assemblages can be used to assess changes in environmental conditions because of their close associations with other organisms, short lifespans and sensitivity to ecological shifts. Their hyper-diversity in marine environments can be partly attributed to the specialisation of the often overlooked and under-sampled cryptic and smaller gastropods, leading to an underestimation of biodiversity. This underscores the need for molecular approaches to enable more efficient detection. However, current universal metabarcoding markers may lack the specificity and resolution to evaluate exceptionally diverse groups such as gastropods. Here, we develop a short gastropod-focused metabarcoding marker 'Gastro16', a novel primer pair that targets an 84-100-bp segment within the mitochondrial 16S rRNA gene, and construct a comprehensive reference library comprising 7335 sequences representing 312 or 55% of known gastropod families. To test the marker's utility, we characterised spatial and temporal patterns of gastropod diversity through metabarcoding Gastro16 and Leray-XT (a marker based on the cytochrome c oxidase subunit I gene; COI) of environmental samples collected from 25 sites along the Norwegian coast and for 15 months from an experimental aquaculture site. Metabarcoding with Gastro16 retrieved more gastropod sequence reads and up to 11 times more gastropod molecular operational taxonomic units compared to COI. Accordingly, the new marker detected greater numbers of gastropod subclasses, orders and families than the COI marker. We present the overall taxonomic diversity of the identified gastropods and their relative occurrences and demonstrate its value for detecting contemporary gastropod diversity, with potential applications to past ecosystems remaining to be tested.
Additional Links: PMID-43036001
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036001,
year = {2026},
author = {Torres, AF and Moles, J and Puillandre, N and Buge, B and Peijnenburg, KTCA and Præbel, K and Renema, W and Todd, JA and Wangensteen, OS},
title = {Gastro16: A Novel eDNA Metabarcoding Marker for Detecting Gastropod Diversity in Marine Environments.},
journal = {Molecular ecology resources},
volume = {26},
number = {8},
pages = {},
doi = {10.1111/1755-0998.70233},
pmid = {43036001},
issn = {1755-0998},
support = {//University of Amsterdam/ ; ECR Grant 2024//Malacological Society of London/ ; 813360//H2020 Marie Skłodowska-Curie Actions/ ; },
mesh = {Animals ; *DNA Barcoding, Taxonomic/methods ; *Gastropoda/classification/genetics ; RNA, Ribosomal, 16S/genetics ; *Biodiversity ; Norway ; Sequence Analysis, DNA ; Extrachromosomal DNA/genetics ; Electron Transport Complex IV/genetics ; DNA Primers/genetics ; DNA, Mitochondrial/genetics/chemistry ; *Metagenomics/methods ; },
abstract = {The diversity of gastropod assemblages can be used to assess changes in environmental conditions because of their close associations with other organisms, short lifespans and sensitivity to ecological shifts. Their hyper-diversity in marine environments can be partly attributed to the specialisation of the often overlooked and under-sampled cryptic and smaller gastropods, leading to an underestimation of biodiversity. This underscores the need for molecular approaches to enable more efficient detection. However, current universal metabarcoding markers may lack the specificity and resolution to evaluate exceptionally diverse groups such as gastropods. Here, we develop a short gastropod-focused metabarcoding marker 'Gastro16', a novel primer pair that targets an 84-100-bp segment within the mitochondrial 16S rRNA gene, and construct a comprehensive reference library comprising 7335 sequences representing 312 or 55% of known gastropod families. To test the marker's utility, we characterised spatial and temporal patterns of gastropod diversity through metabarcoding Gastro16 and Leray-XT (a marker based on the cytochrome c oxidase subunit I gene; COI) of environmental samples collected from 25 sites along the Norwegian coast and for 15 months from an experimental aquaculture site. Metabarcoding with Gastro16 retrieved more gastropod sequence reads and up to 11 times more gastropod molecular operational taxonomic units compared to COI. Accordingly, the new marker detected greater numbers of gastropod subclasses, orders and families than the COI marker. We present the overall taxonomic diversity of the identified gastropods and their relative occurrences and demonstrate its value for detecting contemporary gastropod diversity, with potential applications to past ecosystems remaining to be tested.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*DNA Barcoding, Taxonomic/methods
*Gastropoda/classification/genetics
RNA, Ribosomal, 16S/genetics
*Biodiversity
Norway
Sequence Analysis, DNA
Extrachromosomal DNA/genetics
Electron Transport Complex IV/genetics
DNA Primers/genetics
DNA, Mitochondrial/genetics/chemistry
*Metagenomics/methods
RevDate: 2026-11-16
Genome-Resolved Metagenomics Uncovers Microbial Dark Matter and Complementary Modular Strategies for Lignocellulose-Degradation in Forest Humus Soils.
Advanced science (Weinheim, Baden-Wurttemberg, Germany) pii:e78766 [Epub ahead of print].
Forest humus soil (FHS) harbors highly diverse microbial communities driving carbon transformation, yet genomic resources and strategies underlying lignocellulolysis remain poorly understood. A cross‑climatic FHS metagenome-assembled genome catalog (HSMAG) was constructed from 204 metagenomic FHS samples. HSMAG included 3338 species-level genome bins (96.1% undescribed species) out of 9776 MAGs. These taxa harbored > 669 000 CAZyme sequences. To decipher how they facilitate lignocellulolysis through functional organization, we performed multidomain identification. Results showed that CAZymes were extensively organized into multi-tiered modular degradation strategies, including free CAZymes (FCs), cellulosomes, and polysaccharide utilization loci (PULs). Cellulosome scaffoldins were enriched in nonclassical taxa (Blastocatellia and Gammaproteobacteria), whereas > 44.5% of PULs occurred in Mucilaginibacter and Puia dinghuensis. FCs were enriched in endo- and exo-glucanases, while PULs harbored clustered xylan-degrading glycoside hydrolases (GHs). Thus, these modular strategies might cooperatively deconstruct lignocellulose through complementary GHs. Metatranscriptomics revealed PULs responsive to high lignocellulose levels and identified their core GHs. Recombinant expression and biochemical characterization confirmed the catalytic potential of these GHs. Notably, GH5 exhibited endoglucanase activity of 38.2 U/mg, and GH3 displayed xylanase activity of 20.3 U/mg. Taken together, HSMAG expanded FHS genomic resources and revealed synergistic modular strategies for lignocellulolysis, prioritizing targets for mechanistic investigation and lignocellulose bioconversion.
Additional Links: PMID-43036006
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036006,
year = {2026},
author = {Bai, H and Li, K and Wang, X and Yao, B and Wang, X and Wang, G and Zeng, X and Guo, W and Deng, A and Qiao, S and Chen, Y and Yu, H},
title = {Genome-Resolved Metagenomics Uncovers Microbial Dark Matter and Complementary Modular Strategies for Lignocellulose-Degradation in Forest Humus Soils.},
journal = {Advanced science (Weinheim, Baden-Wurttemberg, Germany)},
volume = {},
number = {},
pages = {},
doi = {10.1002/advs.78766},
pmid = {43036006},
issn = {2198-3844},
support = {//Agricultural Science and Technology Major Project/ ; 2023YFD1302100//Young Scientists Program of the National Key Research and Development Program/ ; NASC2024KY25//Local Financial Funds of National Agricultural Science and Technology Center, Chengdu/ ; NCTIP-XD/B15//National Center of Technology Innovation for Pigs/ ; AKS-2024-AGTI-F14//Project of Joint Research Institute of China Agricultural University in Aksu/ ; 2026TC110//Chinese Universities Scientific Fund/ ; //China Agricultural University Graduate Student Independent Innovation Research Fund/ ; },
abstract = {Forest humus soil (FHS) harbors highly diverse microbial communities driving carbon transformation, yet genomic resources and strategies underlying lignocellulolysis remain poorly understood. A cross‑climatic FHS metagenome-assembled genome catalog (HSMAG) was constructed from 204 metagenomic FHS samples. HSMAG included 3338 species-level genome bins (96.1% undescribed species) out of 9776 MAGs. These taxa harbored > 669 000 CAZyme sequences. To decipher how they facilitate lignocellulolysis through functional organization, we performed multidomain identification. Results showed that CAZymes were extensively organized into multi-tiered modular degradation strategies, including free CAZymes (FCs), cellulosomes, and polysaccharide utilization loci (PULs). Cellulosome scaffoldins were enriched in nonclassical taxa (Blastocatellia and Gammaproteobacteria), whereas > 44.5% of PULs occurred in Mucilaginibacter and Puia dinghuensis. FCs were enriched in endo- and exo-glucanases, while PULs harbored clustered xylan-degrading glycoside hydrolases (GHs). Thus, these modular strategies might cooperatively deconstruct lignocellulose through complementary GHs. Metatranscriptomics revealed PULs responsive to high lignocellulose levels and identified their core GHs. Recombinant expression and biochemical characterization confirmed the catalytic potential of these GHs. Notably, GH5 exhibited endoglucanase activity of 38.2 U/mg, and GH3 displayed xylanase activity of 20.3 U/mg. Taken together, HSMAG expanded FHS genomic resources and revealed synergistic modular strategies for lignocellulolysis, prioritizing targets for mechanistic investigation and lignocellulose bioconversion.},
}
RevDate: 2026-11-16
CmpDate: 2026-11-16
The chromosomal genome sequence of a peltospirid gastropod, Rhynchopelta concentrica McLean, 1989 (Neomphalida: Peltospiridae) and its associated microbial metagenome sequences.
Wellcome open research, 11:519.
We present a genome assembly from a specimen of Rhynchopelta concentrica (peltospirid gastropod; Mollusca; Gastropoda; Neomphalida; Peltospiridae). The genome sequence has a total length of 716.07 megabases. Most of the assembly (92.08%) is scaffolded into 15 chromosomal pseudomolecules. The mitochondrial genome has also been assembled, with a length of 15.75 kilobases. From the metagenome data, we recovered nine bins, of which four were high-quality MAGs.
Additional Links: PMID-43036310
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036310,
year = {2026},
author = {Hughes, IV and Hourdez, S and Beinart, RA and Girguis, P and Oatley, G and Sinclair, E and Aunin, E and Gettle, N and Santos, C and Paulini, M and Niu, H and McKenna, V and O'Brien, R and , and , and , and , and , },
title = {The chromosomal genome sequence of a peltospirid gastropod, Rhynchopelta concentrica McLean, 1989 (Neomphalida: Peltospiridae) and its associated microbial metagenome sequences.},
journal = {Wellcome open research},
volume = {11},
number = {},
pages = {519},
pmid = {43036310},
issn = {2398-502X},
mesh = {Animals ; *Gastropoda/genetics/microbiology ; *Metagenome ; *Chromosomes/genetics ; *Genome ; Genome, Mitochondrial ; Sequence Analysis, DNA ; },
abstract = {We present a genome assembly from a specimen of Rhynchopelta concentrica (peltospirid gastropod; Mollusca; Gastropoda; Neomphalida; Peltospiridae). The genome sequence has a total length of 716.07 megabases. Most of the assembly (92.08%) is scaffolded into 15 chromosomal pseudomolecules. The mitochondrial genome has also been assembled, with a length of 15.75 kilobases. From the metagenome data, we recovered nine bins, of which four were high-quality MAGs.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Gastropoda/genetics/microbiology
*Metagenome
*Chromosomes/genetics
*Genome
Genome, Mitochondrial
Sequence Analysis, DNA
RevDate: 2026-11-16
CmpDate: 2026-11-16
Lung microbiota in COPD: a double-edged sword.
Current research in microbial sciences, 11:100679.
The respiratory microbiota constitutes a dynamic, low-biomass microbial ecosystem whose composition and function are shaped by continuous immigration, elimination, local growth conditions and host-environment interactions. Chronic obstructive pulmonary disease (COPD) provides a clinically important model in which this ecological equilibrium is profoundly altered. This review examines the lung microbiota through a "double-edged sword" framework, proposing that its biological effects depend less on the presence of individual beneficial or pathogenic taxa than on community structure, ecological stability, microbial function and host context. We examine the organization of the lung microbial ecosystem in health and its transition toward dysbiotic, unstable and pathobiont-enriched states in COPD, together with the host, environmental and therapeutic factors that shape these transitions. Particular attention is given to microbial community dynamics, functional alterations, epithelial and immune interactions, antimicrobial defense, and the contribution of the gut-lung microbial axis. We further address the unresolved directionality of the dysbiosis-COPD relationship, considering evidence that COPD-associated changes in the airway environment can select for altered microbial communities while established dysbiosis may subsequently reinforce disease-associated conditions, generating a self-perpetuating ecological cycle. Longitudinal profiling, metagenomics, metatranscriptomics, metabolomics and integrated host-microbiome approaches are discussed as means of moving beyond taxonomic associations toward microbial function and mechanism. Finally, we critically evaluate emerging microbial signatures with potential prognostic or treatment-response relevance and microbiota-targeted interventions, while emphasizing current limitations in causality, methodological standardization, strain-level resolution and clinical validation. A microbiota-centered perspective may therefore help clarify how changes in the structure and function of the lung microbiota contribute to the transition from pulmonary homeostasis to COPD-associated dysbiosis and disease progression.
Additional Links: PMID-43036457
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036457,
year = {2026},
author = {Bentaher, A},
title = {Lung microbiota in COPD: a double-edged sword.},
journal = {Current research in microbial sciences},
volume = {11},
number = {},
pages = {100679},
pmid = {43036457},
issn = {2666-5174},
abstract = {The respiratory microbiota constitutes a dynamic, low-biomass microbial ecosystem whose composition and function are shaped by continuous immigration, elimination, local growth conditions and host-environment interactions. Chronic obstructive pulmonary disease (COPD) provides a clinically important model in which this ecological equilibrium is profoundly altered. This review examines the lung microbiota through a "double-edged sword" framework, proposing that its biological effects depend less on the presence of individual beneficial or pathogenic taxa than on community structure, ecological stability, microbial function and host context. We examine the organization of the lung microbial ecosystem in health and its transition toward dysbiotic, unstable and pathobiont-enriched states in COPD, together with the host, environmental and therapeutic factors that shape these transitions. Particular attention is given to microbial community dynamics, functional alterations, epithelial and immune interactions, antimicrobial defense, and the contribution of the gut-lung microbial axis. We further address the unresolved directionality of the dysbiosis-COPD relationship, considering evidence that COPD-associated changes in the airway environment can select for altered microbial communities while established dysbiosis may subsequently reinforce disease-associated conditions, generating a self-perpetuating ecological cycle. Longitudinal profiling, metagenomics, metatranscriptomics, metabolomics and integrated host-microbiome approaches are discussed as means of moving beyond taxonomic associations toward microbial function and mechanism. Finally, we critically evaluate emerging microbial signatures with potential prognostic or treatment-response relevance and microbiota-targeted interventions, while emphasizing current limitations in causality, methodological standardization, strain-level resolution and clinical validation. A microbiota-centered perspective may therefore help clarify how changes in the structure and function of the lung microbiota contribute to the transition from pulmonary homeostasis to COPD-associated dysbiosis and disease progression.},
}
RevDate: 2026-11-16
CmpDate: 2026-11-16
Suspected Schizophyllum commune Pulmonary Disease in an Apparently Immunocompetent Host: A Case Report and Literature Review.
Infection and drug resistance, 19:642917.
BACKGROUND: Schizophyllum commune causes allergic bronchopulmonary mycosis (ABPM) and other respiratory disorders. Its detection in a respiratory specimen, however, does not establish whether it is responsible for the illness.
CASE PRESENTATION: An apparently immunocompetent 45-year-old man developed cough and wheezing two weeks after cleaning a moldy basement without respiratory protection. Symptoms persisted despite moxifloxacin, with marked blood eosinophilia and elevated total IgE. Metagenomic next-generation sequencing (mNGS) of bronchoalveolar lavage fluid (BALF) detected S. commune and Haemophilus influenzae. BALF fungal culture was not performed, and an endobronchial biopsy showed no fungal elements. Symptoms resolved during combined corticosteroid, antibacterial and antifungal treatment. The patient continued oral voriconazole after discharge but stopped it without medical advice during the fourth month. Cough and wheezing recurred approximately two weeks later, with an absolute eosinophil count of 17.83 × 10[9]/L and no interval change on chest computed tomography (CT). Voriconazole was resumed together with a 10-week course of oral prednisone acetate, and symptoms resolved. The interval from starting voriconazole to final discontinuation was six months, including the interruption. The patient remained symptom-free during 12 months of follow-up after treatment ended. This interval reflects the individual clinical course and should not be interpreted as a recommended treatment duration.
CONCLUSION: This case illustrates the difficulty of interpreting BALF mNGS detection of S. commune in a patient with recurrent wheezing and eosinophilia. Neither ABPM nor invasive infection was established. Concurrent corticosteroid and antifungal treatment limited assessment of the contribution of each drug.
Additional Links: PMID-43036548
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036548,
year = {2026},
author = {Liu, J and Jiang, W and Duan, J and Zhang, L and Yang, X and Yang, W},
title = {Suspected Schizophyllum commune Pulmonary Disease in an Apparently Immunocompetent Host: A Case Report and Literature Review.},
journal = {Infection and drug resistance},
volume = {19},
number = {},
pages = {642917},
pmid = {43036548},
issn = {1178-6973},
abstract = {BACKGROUND: Schizophyllum commune causes allergic bronchopulmonary mycosis (ABPM) and other respiratory disorders. Its detection in a respiratory specimen, however, does not establish whether it is responsible for the illness.
CASE PRESENTATION: An apparently immunocompetent 45-year-old man developed cough and wheezing two weeks after cleaning a moldy basement without respiratory protection. Symptoms persisted despite moxifloxacin, with marked blood eosinophilia and elevated total IgE. Metagenomic next-generation sequencing (mNGS) of bronchoalveolar lavage fluid (BALF) detected S. commune and Haemophilus influenzae. BALF fungal culture was not performed, and an endobronchial biopsy showed no fungal elements. Symptoms resolved during combined corticosteroid, antibacterial and antifungal treatment. The patient continued oral voriconazole after discharge but stopped it without medical advice during the fourth month. Cough and wheezing recurred approximately two weeks later, with an absolute eosinophil count of 17.83 × 10[9]/L and no interval change on chest computed tomography (CT). Voriconazole was resumed together with a 10-week course of oral prednisone acetate, and symptoms resolved. The interval from starting voriconazole to final discontinuation was six months, including the interruption. The patient remained symptom-free during 12 months of follow-up after treatment ended. This interval reflects the individual clinical course and should not be interpreted as a recommended treatment duration.
CONCLUSION: This case illustrates the difficulty of interpreting BALF mNGS detection of S. commune in a patient with recurrent wheezing and eosinophilia. Neither ABPM nor invasive infection was established. Concurrent corticosteroid and antifungal treatment limited assessment of the contribution of each drug.},
}
RevDate: 2026-11-16
CmpDate: 2026-11-16
Ovarian function via gut microbiota-driven serotonin dysregulation in zebrafish.
iScience, 29(11):117879.
Ciprofloxacin (CIP) is frequently detected in aquatic environments, yet its effects on reproduction through gut microbiota-mediated pathways remain unclear. Adult female zebrafish were exposed to 5,000 ng/L CIP for 30 days and evaluated using histology, hormone assays, metagenomics, metabolomics, transcriptomics, and fecal microbiota transplantation (FMT). CIP impaired folliculogenesis, reduced follicle-stimulating hormone, luteinizing hormone, and estradiol levels, and disrupted intestinal barrier integrity. Multi-omics analyses revealed gut dysbiosis and altered microbial metabolism, including increased serotonin (5-HT) and indole-3-lactic acid associated with Cetobacterium and Plesiomonas shigelloides. Brain transcriptomics showed suppressed hormone signaling and elevated 5-HT, consistent with hypothalamic-pituitary-gonadal axis disruption. FMT from CIP-exposed donors reproduced hormonal and ovarian abnormalities in microbiota-depleted recipients. This study underscores the need to integrate microbiota-host interactions into the risk assessment of emerging aquatic contaminants and highlights the broader implications of environmental exposures for reproductive and endocrine health in both aquatic organisms and potentially humans.
Additional Links: PMID-43036710
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036710,
year = {2026},
author = {Wang, N and Dong, T and Yu, K and Qiu, Y and Lyu, D and Li, B and Yu, X and Jing, CH and Jiao, X},
title = {Ovarian function via gut microbiota-driven serotonin dysregulation in zebrafish.},
journal = {iScience},
volume = {29},
number = {11},
pages = {117879},
pmid = {43036710},
issn = {2589-0042},
abstract = {Ciprofloxacin (CIP) is frequently detected in aquatic environments, yet its effects on reproduction through gut microbiota-mediated pathways remain unclear. Adult female zebrafish were exposed to 5,000 ng/L CIP for 30 days and evaluated using histology, hormone assays, metagenomics, metabolomics, transcriptomics, and fecal microbiota transplantation (FMT). CIP impaired folliculogenesis, reduced follicle-stimulating hormone, luteinizing hormone, and estradiol levels, and disrupted intestinal barrier integrity. Multi-omics analyses revealed gut dysbiosis and altered microbial metabolism, including increased serotonin (5-HT) and indole-3-lactic acid associated with Cetobacterium and Plesiomonas shigelloides. Brain transcriptomics showed suppressed hormone signaling and elevated 5-HT, consistent with hypothalamic-pituitary-gonadal axis disruption. FMT from CIP-exposed donors reproduced hormonal and ovarian abnormalities in microbiota-depleted recipients. This study underscores the need to integrate microbiota-host interactions into the risk assessment of emerging aquatic contaminants and highlights the broader implications of environmental exposures for reproductive and endocrine health in both aquatic organisms and potentially humans.},
}
RevDate: 2026-11-16
CmpDate: 2026-11-16
[A prospective study of the microbiota present on the surface of removable orthodontic appliances].
L' Orthodontie francaise, 97(2):201-221.
INTRODUCTION: Removable orthodontic appliances (ROAs) constitute a reservoir for microorganisms. Due to their numerous retentive areas, they promote bacterial adhesion and biofilm accumulation. However, no study has yet evaluated the microbiota on the surface of ROAs using molecular techniques. The aim of this study was therefore to investigate, from a microbiological perspective, the surface of resin-based ROAs after intraoral use in children.
MATERIALS AND METHOD: This study included patients aged between 8 and 12 years wearing an ROA with a PMMA resin base (PolyMethylMethAcrylate). Samples were collected from the ROAs after 3 months (T1) and 6 months (T2) of wear. Wearing time, plaque index (PI), and bleeding on probing (BOP) were also recorded at T1 and T2. DNA extraction and sequencing were subsequently performed.
RESULTS: Thirty patients (9.6±1.4 years) were included. Forty-nine out of 60 samples were successfully analyzed, corresponding to 25 patients. Metagenomic data showed high similarity among samples, except for one patient. Fourteen phyla and 35 bacterial genera were identified, including 26 predominant genera. Six phyla belonged to the oral microbiota, with a predominance of Bacillota and Proteobacteria at both T1 and T2. Four genera were predominant at T1 and T2: Streptococcus, Haemophilus, Neisseria, and Veillonella. Between T1 and T2, a non-significant decrease in alpha diversity and a significant decrease in the genera Porphyromonas, Prevotella, and Tannerella were observed (p<0.05). No significant differences were found for PI, BOP, or wearing time between T1 and T2.
CONCLUSION: Numerous bacterial genera develop on the surface of ROAs in children aged 8 to 12 years. The use of 16S rRNA metagenomics enabled a more detailed investigation of the bacterial composition. Dynamic bacterial changes occur during the wear of these appliances, thereby impacting the oral microbiota. Further studies are now needed to investigate microbiota evolution in appliances worn beyond 6 months and the associated implications.
Additional Links: PMID-43036989
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43036989,
year = {2026},
author = {Lopez, S and Dridi, SM and Lupi, L and Charavet, C},
title = {[A prospective study of the microbiota present on the surface of removable orthodontic appliances].},
journal = {L' Orthodontie francaise},
volume = {97},
number = {2},
pages = {201-221},
doi = {10.1684/orthodfr.2026.234},
pmid = {43036989},
issn = {1954-3395},
mesh = {Humans ; Child ; *Orthodontic Appliances, Removable/microbiology ; *Microbiota/genetics ; Prospective Studies ; Female ; Male ; Surface Properties ; Biofilms ; Mouth/microbiology ; Dental Plaque Index ; },
abstract = {INTRODUCTION: Removable orthodontic appliances (ROAs) constitute a reservoir for microorganisms. Due to their numerous retentive areas, they promote bacterial adhesion and biofilm accumulation. However, no study has yet evaluated the microbiota on the surface of ROAs using molecular techniques. The aim of this study was therefore to investigate, from a microbiological perspective, the surface of resin-based ROAs after intraoral use in children.
MATERIALS AND METHOD: This study included patients aged between 8 and 12 years wearing an ROA with a PMMA resin base (PolyMethylMethAcrylate). Samples were collected from the ROAs after 3 months (T1) and 6 months (T2) of wear. Wearing time, plaque index (PI), and bleeding on probing (BOP) were also recorded at T1 and T2. DNA extraction and sequencing were subsequently performed.
RESULTS: Thirty patients (9.6±1.4 years) were included. Forty-nine out of 60 samples were successfully analyzed, corresponding to 25 patients. Metagenomic data showed high similarity among samples, except for one patient. Fourteen phyla and 35 bacterial genera were identified, including 26 predominant genera. Six phyla belonged to the oral microbiota, with a predominance of Bacillota and Proteobacteria at both T1 and T2. Four genera were predominant at T1 and T2: Streptococcus, Haemophilus, Neisseria, and Veillonella. Between T1 and T2, a non-significant decrease in alpha diversity and a significant decrease in the genera Porphyromonas, Prevotella, and Tannerella were observed (p<0.05). No significant differences were found for PI, BOP, or wearing time between T1 and T2.
CONCLUSION: Numerous bacterial genera develop on the surface of ROAs in children aged 8 to 12 years. The use of 16S rRNA metagenomics enabled a more detailed investigation of the bacterial composition. Dynamic bacterial changes occur during the wear of these appliances, thereby impacting the oral microbiota. Further studies are now needed to investigate microbiota evolution in appliances worn beyond 6 months and the associated implications.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
Child
*Orthodontic Appliances, Removable/microbiology
*Microbiota/genetics
Prospective Studies
Female
Male
Surface Properties
Biofilms
Mouth/microbiology
Dental Plaque Index
RevDate: 2026-11-16
Fragment End Motif Analysis of Microbial Cell-Free DNA Improves Detection of Pathogens over Noise from Contaminants and Commensals.
Clinical chemistry pii:9065322 [Epub ahead of print].
BACKGROUND: Circulating plasma microbial DNA (mDNA) is a promising biomarker for sepsis, but its accuracy is hindered by the low abundance of pathogen DNA and the presence of contaminants and commensals. mDNA enrichment using a selection of short fragments and single-stranded DNA (ssDNA) library preparation can improve pathogen yield but also increases background noise. Since nucleotide representation at mDNA fragment ends is biased by enzymatic digestion in vivo, we hypothesized that analysis of 5' and 3' mDNA fragment end motifs in enriched ssDNA libraries could improve pathogen detection.
METHODS: We performed deep metagenomic sequencing of size-selected ssDNA libraries from 38 plasma samples, including 20 samples from 5 sepsis patients, 18 samples from 6 patients without infection, and 6 no-template controls. Across these samples, we compared observed-to-expected ratios for end motifs in mDNA fragments from candidate pathogens and contaminants. For each candidate pathogen, we evaluated the accuracy of combining end motifs with microbial abundance for detection of infection.
RESULTS: mDNA fragments from infection-causing pathogens were biased in the nucleotide motifs at both fragment ends compared with contaminants, with the highest enrichment observed in the GG dinucleotide at the 3' end (P < 0.0001). Integrating microbial abundance with end motif features improved differentiation between clinically significant pathogens and corresponding noise from the same microbes in noninfection samples. For Staphylococcus infections, area under the receiver operating characteristic curve for the combination model was higher than than that for microbial abundance alone (0.892 vs 0.431, respectively; P = 0.018).
CONCLUSIONS: mDNA end motif analysis improves signal-to-noise ratio beyond microbial abundance alone and improves pathogen detection in plasma metagenomic sequencing.
Additional Links: PMID-43037253
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43037253,
year = {2026},
author = {Zhang, H and Dominguez, EG and Junak, M and Lauer, KV and Werner, NL and Murtaza, M and Pepperell, CS and Kisat, MT},
title = {Fragment End Motif Analysis of Microbial Cell-Free DNA Improves Detection of Pathogens over Noise from Contaminants and Commensals.},
journal = {Clinical chemistry},
volume = {},
number = {},
pages = {},
doi = {10.1093/clinchem/hvag133},
pmid = {43037253},
issn = {1530-8561},
support = {//American Association for the Surgery of Trauma/ ; //Research and Education Fund Trauma Critical Care/ ; //American College of Surgeons/ ; //NIH/ ; K08GM148858/GM/NIGMS NIH HHS/United States ; //Wisconsin Partnership Program/ ; },
abstract = {BACKGROUND: Circulating plasma microbial DNA (mDNA) is a promising biomarker for sepsis, but its accuracy is hindered by the low abundance of pathogen DNA and the presence of contaminants and commensals. mDNA enrichment using a selection of short fragments and single-stranded DNA (ssDNA) library preparation can improve pathogen yield but also increases background noise. Since nucleotide representation at mDNA fragment ends is biased by enzymatic digestion in vivo, we hypothesized that analysis of 5' and 3' mDNA fragment end motifs in enriched ssDNA libraries could improve pathogen detection.
METHODS: We performed deep metagenomic sequencing of size-selected ssDNA libraries from 38 plasma samples, including 20 samples from 5 sepsis patients, 18 samples from 6 patients without infection, and 6 no-template controls. Across these samples, we compared observed-to-expected ratios for end motifs in mDNA fragments from candidate pathogens and contaminants. For each candidate pathogen, we evaluated the accuracy of combining end motifs with microbial abundance for detection of infection.
RESULTS: mDNA fragments from infection-causing pathogens were biased in the nucleotide motifs at both fragment ends compared with contaminants, with the highest enrichment observed in the GG dinucleotide at the 3' end (P < 0.0001). Integrating microbial abundance with end motif features improved differentiation between clinically significant pathogens and corresponding noise from the same microbes in noninfection samples. For Staphylococcus infections, area under the receiver operating characteristic curve for the combination model was higher than than that for microbial abundance alone (0.892 vs 0.431, respectively; P = 0.018).
CONCLUSIONS: mDNA end motif analysis improves signal-to-noise ratio beyond microbial abundance alone and improves pathogen detection in plasma metagenomic sequencing.},
}
RevDate: 2026-11-16
CmpDate: 2026-11-16
Dynamic exchange of bacteria and carbapenem resistance genes between sewer biofilms and wastewater.
Water science and technology : a journal of the International Association on Water Pollution Research, 94(9):1524-1542.
Sewer biofilms exchange bacteria and antibiotic-resistance genes with the overlying wastewater. Using inline biofilm reactors, movement of bacteria, 16S rRNA, and carbapenemase genes (blaKPC, blaVIM, blaNDM, blaOXA-48-like, and blaIMP) between wastewater and sewer biofilms was studied. Over five experiments, biofilm establishment on polyvinyl chloride (PVC), changes in bacterial community after exposure to raw and treated wastewater, biofilm absorption of bacteria, and release of bacteria and genes were investigated. In laboratory studies, established biofilms with carbapenem-resistant bacteria contributed more bacteria to wastewater than to downstream biofilms under realistic shear. In field studies, established biofilms acquired target carbapenem-resistant bacteria and genes from wastewater within hours, and concentrations stabilized between 2 and 15 days and then varied by <0.5 most probable number/cm[2] or 0.5 log gene copies (GC)/cm[2]. Metagenomic profiles of biofilm bacteria changed over 21 days. Established biofilms with antibiotic-resistant bacteria and genes exposed to tertiary-treated wastewater without target carbapenemase genes or meropenem antibiotics retained resistant genes and bacteria over 9 days dropping < 1 log GC/cm[2]. Results show that sewer biofilms contribute to wastewater antibiotic-resistance gene signals by absorbing and later releasing bacteria and genes. Sewer biofilm dynamics evaluation is essential for accurate interpretation of bacterial concentrations in wastewater-based epidemiology studies.
Additional Links: PMID-43037632
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43037632,
year = {2026},
author = {Warren, E and Brazelton, WJ and Fusco, S and VanDerslice, J and Benson, LS and Tanner, W and Weidhaas, J},
title = {Dynamic exchange of bacteria and carbapenem resistance genes between sewer biofilms and wastewater.},
journal = {Water science and technology : a journal of the International Association on Water Pollution Research},
volume = {94},
number = {9},
pages = {1524-1542},
pmid = {43037632},
issn = {0273-1223},
support = {200-2021-12774/JG/JG CDC HHS/United States ; },
mesh = {*Biofilms ; *Wastewater/microbiology ; *Carbapenems/pharmacology ; *Bacteria/genetics/drug effects ; *Sewage/microbiology ; *Drug Resistance, Bacterial/genetics ; beta-Lactamases/genetics ; Genes, Bacterial ; RNA, Ribosomal, 16S/genetics ; Bacterial Proteins/genetics ; Anti-Bacterial Agents/pharmacology ; },
abstract = {Sewer biofilms exchange bacteria and antibiotic-resistance genes with the overlying wastewater. Using inline biofilm reactors, movement of bacteria, 16S rRNA, and carbapenemase genes (blaKPC, blaVIM, blaNDM, blaOXA-48-like, and blaIMP) between wastewater and sewer biofilms was studied. Over five experiments, biofilm establishment on polyvinyl chloride (PVC), changes in bacterial community after exposure to raw and treated wastewater, biofilm absorption of bacteria, and release of bacteria and genes were investigated. In laboratory studies, established biofilms with carbapenem-resistant bacteria contributed more bacteria to wastewater than to downstream biofilms under realistic shear. In field studies, established biofilms acquired target carbapenem-resistant bacteria and genes from wastewater within hours, and concentrations stabilized between 2 and 15 days and then varied by <0.5 most probable number/cm[2] or 0.5 log gene copies (GC)/cm[2]. Metagenomic profiles of biofilm bacteria changed over 21 days. Established biofilms with antibiotic-resistant bacteria and genes exposed to tertiary-treated wastewater without target carbapenemase genes or meropenem antibiotics retained resistant genes and bacteria over 9 days dropping < 1 log GC/cm[2]. Results show that sewer biofilms contribute to wastewater antibiotic-resistance gene signals by absorbing and later releasing bacteria and genes. Sewer biofilm dynamics evaluation is essential for accurate interpretation of bacterial concentrations in wastewater-based epidemiology studies.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Biofilms
*Wastewater/microbiology
*Carbapenems/pharmacology
*Bacteria/genetics/drug effects
*Sewage/microbiology
*Drug Resistance, Bacterial/genetics
beta-Lactamases/genetics
Genes, Bacterial
RNA, Ribosomal, 16S/genetics
Bacterial Proteins/genetics
Anti-Bacterial Agents/pharmacology
RevDate: 2026-11-16
CmpDate: 2026-11-16
Microbial succession and functional gene repertoires in Brassica rapa fermentation.
World journal of microbiology & biotechnology, 42(12):.
Traditional fermented vegetables are complex microbial ecosystems whose dynamics are shaped by processing conditions, yet many regional European fermentations remain poorly characterized. This study combined shotgun metagenomics, culture-dependent analyses, and chemical measurements to characterize microbial succession, fermentation dynamics, and metagenomic functional potential during brovada fermentation and to compare one semi-industrial and one artisanal production system over 30 days. The two systems showed distinct trajectories. In the semi-industrial system, lactic acid bacteria (LAB) increased to 8.30 ± 0.50 log CFU/g by day 30, together with rapid sugar depletion, lactic acid accumulation, and a final pH of 3.94 ± 0.09. In the artisanal system, LAB peaked at 5.63 ± 0.38 log CFU/g at day 9 and declined to 3.92 ± 0.26 log CFU/g, while the final pH remained higher (4.51 ± 0.08); yeasts became more abundant than LAB at the end of fermentation. Metagenomic profiling showed a faster transition from Proteobacteria- to Firmicutes/LAB-dominated communities in the semi-industrial system, whereas the artisanal system retained greater microbial diversity and a longer persistence of Pseudomonas, Acinetobacter, and other environmental taxa. Functional profiling identified genes associated with bile salt hydrolysis, adhesion, exopolysaccharide production, gamma-aminobutyric acid (GABA) synthesis, plantaricin production, myrosinase-like activity, nitrate/nitrite metabolism, and amino acid and folate biosynthesis. Overall, the study shows that brovada fermentation can follow markedly different microbial and metabolic trajectories under contrasting production conditions, with corresponding differences in functional gene repertoires. These findings highlight the close relationship between processing conditions, microbial succession, and the functional potential of traditional vegetable fermentations.
Additional Links: PMID-43037819
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43037819,
year = {2026},
author = {Rossi, A and Maifreni, M and Innocente, N and Trossolo, E and Marroni, F and Marino, M},
title = {Microbial succession and functional gene repertoires in Brassica rapa fermentation.},
journal = {World journal of microbiology & biotechnology},
volume = {42},
number = {12},
pages = {},
pmid = {43037819},
issn = {1573-0972},
mesh = {*Fermentation ; Metagenomics ; *Brassica rapa/microbiology/metabolism ; *Bacteria/genetics/classification/isolation & purification/metabolism ; Metagenome ; Hydrogen-Ion Concentration ; Fermented Foods/microbiology ; Microbiota/genetics ; Yeasts/genetics/classification/metabolism/isolation & purification ; Lactobacillales/genetics/metabolism ; },
abstract = {Traditional fermented vegetables are complex microbial ecosystems whose dynamics are shaped by processing conditions, yet many regional European fermentations remain poorly characterized. This study combined shotgun metagenomics, culture-dependent analyses, and chemical measurements to characterize microbial succession, fermentation dynamics, and metagenomic functional potential during brovada fermentation and to compare one semi-industrial and one artisanal production system over 30 days. The two systems showed distinct trajectories. In the semi-industrial system, lactic acid bacteria (LAB) increased to 8.30 ± 0.50 log CFU/g by day 30, together with rapid sugar depletion, lactic acid accumulation, and a final pH of 3.94 ± 0.09. In the artisanal system, LAB peaked at 5.63 ± 0.38 log CFU/g at day 9 and declined to 3.92 ± 0.26 log CFU/g, while the final pH remained higher (4.51 ± 0.08); yeasts became more abundant than LAB at the end of fermentation. Metagenomic profiling showed a faster transition from Proteobacteria- to Firmicutes/LAB-dominated communities in the semi-industrial system, whereas the artisanal system retained greater microbial diversity and a longer persistence of Pseudomonas, Acinetobacter, and other environmental taxa. Functional profiling identified genes associated with bile salt hydrolysis, adhesion, exopolysaccharide production, gamma-aminobutyric acid (GABA) synthesis, plantaricin production, myrosinase-like activity, nitrate/nitrite metabolism, and amino acid and folate biosynthesis. Overall, the study shows that brovada fermentation can follow markedly different microbial and metabolic trajectories under contrasting production conditions, with corresponding differences in functional gene repertoires. These findings highlight the close relationship between processing conditions, microbial succession, and the functional potential of traditional vegetable fermentations.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Fermentation
Metagenomics
*Brassica rapa/microbiology/metabolism
*Bacteria/genetics/classification/isolation & purification/metabolism
Metagenome
Hydrogen-Ion Concentration
Fermented Foods/microbiology
Microbiota/genetics
Yeasts/genetics/classification/metabolism/isolation & purification
Lactobacillales/genetics/metabolism
RevDate: 2026-11-16
CmpDate: 2026-11-16
Suspected partial arginine vasopressin deficiency during herpes simplex virus type 1 meningoencephalitis: a case report.
Journal of neurovirology, 32(6):.
Herpes simplex virus type 1 (HSV-1) encephalitis can be accompanied by complex disturbances of water and sodium balance. A 55-year-old woman presented with fever, recurrent seizures and profound encephalopathy. Cerebrospinal fluid DNA-based metagenomic sequencing yielded 13,426 HSV-1 reads with 94.46% genomic coverage; RNA sequencing identified 18 reads. Severe polyuria (maximum complete nursing-day output, 7,525 mL), hypernatremia (maximum 149.7 mmol/L) and increased plasma osmolality prompted suspicion of partial arginine vasopressin deficiency. Interpretation was limited by concurrent mannitol, hypokalemia, variable fluid therapy and overlapping enteral and intravenous desmopressin. Because of critical illness, acyclovir and foscarnet were empirically initiated together; acyclovir continued for 34 days. Creatinine-based renal function remained preserved on available monitoring. No standardized desmopressin challenge, paired pretreatment blood and urine osmolality, or definitive hypothalamic-neurohypophyseal lesion was documented. Desmopressin tablets were discontinued by June 2026. Serum sodium in August was 133 mmol/L, and neurological recovery was incomplete despite regained ambulation and independence in basic daily activities. The case emphasizes careful differential diagnosis of polyuria in severe viral encephalitis rather than confirmation of a direct HSV-related posterior pituitary injury.
Additional Links: PMID-43037887
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43037887,
year = {2026},
author = {Wang, R and Cao, L and Zhao, Y and Hao, H and Li, N},
title = {Suspected partial arginine vasopressin deficiency during herpes simplex virus type 1 meningoencephalitis: a case report.},
journal = {Journal of neurovirology},
volume = {32},
number = {6},
pages = {},
pmid = {43037887},
issn = {1538-2443},
mesh = {Humans ; Female ; Middle Aged ; Acyclovir/therapeutic use ; *Herpesvirus 1, Human/pathogenicity/genetics/drug effects ; *Arginine Vasopressin/deficiency ; *Encephalitis, Herpes Simplex/drug therapy/virology/complications ; Antiviral Agents/therapeutic use ; *Polyuria/virology/drug therapy ; Hypernatremia/virology/drug therapy ; *Meningoencephalitis/virology/drug therapy ; Deamino Arginine Vasopressin/therapeutic use ; Foscarnet/therapeutic use ; },
abstract = {Herpes simplex virus type 1 (HSV-1) encephalitis can be accompanied by complex disturbances of water and sodium balance. A 55-year-old woman presented with fever, recurrent seizures and profound encephalopathy. Cerebrospinal fluid DNA-based metagenomic sequencing yielded 13,426 HSV-1 reads with 94.46% genomic coverage; RNA sequencing identified 18 reads. Severe polyuria (maximum complete nursing-day output, 7,525 mL), hypernatremia (maximum 149.7 mmol/L) and increased plasma osmolality prompted suspicion of partial arginine vasopressin deficiency. Interpretation was limited by concurrent mannitol, hypokalemia, variable fluid therapy and overlapping enteral and intravenous desmopressin. Because of critical illness, acyclovir and foscarnet were empirically initiated together; acyclovir continued for 34 days. Creatinine-based renal function remained preserved on available monitoring. No standardized desmopressin challenge, paired pretreatment blood and urine osmolality, or definitive hypothalamic-neurohypophyseal lesion was documented. Desmopressin tablets were discontinued by June 2026. Serum sodium in August was 133 mmol/L, and neurological recovery was incomplete despite regained ambulation and independence in basic daily activities. The case emphasizes careful differential diagnosis of polyuria in severe viral encephalitis rather than confirmation of a direct HSV-related posterior pituitary injury.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
Female
Middle Aged
Acyclovir/therapeutic use
*Herpesvirus 1, Human/pathogenicity/genetics/drug effects
*Arginine Vasopressin/deficiency
*Encephalitis, Herpes Simplex/drug therapy/virology/complications
Antiviral Agents/therapeutic use
*Polyuria/virology/drug therapy
Hypernatremia/virology/drug therapy
*Meningoencephalitis/virology/drug therapy
Deamino Arginine Vasopressin/therapeutic use
Foscarnet/therapeutic use
RevDate: 2026-11-16
Comparative metagenomics of extremophilic microbiomes from Pakistan reveals a conserved taxonomic core and habitat-specific functional gene signatures.
Folia microbiologica [Epub ahead of print].
Microbial communities inhabiting extreme environments serve as valuable models for understanding microbial diversity under environmental stress and represent potential reservoirs of novel functional diversity with biotechnological relevance. Despite their ecological importance, the taxonomic composition and functional gene repertoires of extremophilic microbiomes from Pakistani habitats remain largely unexplored. We developed a comparative multi-database shotgun metagenomic framework to characterize conserved taxonomic features and habitat-associated functional gene signatures across three contrasting extreme environments: the Gilgit Hot Spring, the Kharan Desert, and the Kalabagh Salt Mine, using best-hit taxonomic and multi-database functional annotation (KEGG, eggNOG, GO, CAZy, Pfam, CYPED) combined with PERMANOVA-based statistical testing to evaluate habitat-associated community and functional divergence. Across all sites, Proteobacteria, Firmicutes, and Actinobacteria dominated as core phyla, while community composition diverged substantially at finer taxonomic resolutions, with beta-diversity analysis confirming statistically distinct communities across all three habitats (PERMANOVA: pseudo-F = 21.67, R[2] = 0.812, p = 0.001). Functional annotation revealed habitat-associated differences in gene representation. Hot spring microbiomes showed higher representation of genes associated with protein synthesis, folding, and redox-related processes. Desert communities exhibited a relatively broader representation of carbohydrate and amino acid metabolic pathways, including a higher relative abundance of carbohydrate-active enzymes. Salt mine communities showed a higher representation of genes associated with genome maintenance, membrane transport, and osmoregulation. These habitat-associated differences in functional gene representation were independently identified across multiple annotation frameworks, supporting the consistency of the observed trends. Notably, eggNOG revealed a substantial fraction of uncharacterized genes (16-19%), and CYPED analysis identified cytochrome P450 homologs with only 22-33% sequence identity to known homologs, highlighting the unexplored genetic potential of these microbiomes. Together, these findings suggest that environmental differences among habitats are associated with habitat-specific structuring of both microbial community composition and functional gene repertoires.
Additional Links: PMID-43037924
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43037924,
year = {2026},
author = {Nughman, M and Khan, H and Muhammad, N and Almehmadi, M and Bakhuraysah, MM and Tayeb, FJ and Dawar, FU and Khan, FA and Khan, Z and Ali, N},
title = {Comparative metagenomics of extremophilic microbiomes from Pakistan reveals a conserved taxonomic core and habitat-specific functional gene signatures.},
journal = {Folia microbiologica},
volume = {},
number = {},
pages = {},
pmid = {43037924},
issn = {1874-9356},
abstract = {Microbial communities inhabiting extreme environments serve as valuable models for understanding microbial diversity under environmental stress and represent potential reservoirs of novel functional diversity with biotechnological relevance. Despite their ecological importance, the taxonomic composition and functional gene repertoires of extremophilic microbiomes from Pakistani habitats remain largely unexplored. We developed a comparative multi-database shotgun metagenomic framework to characterize conserved taxonomic features and habitat-associated functional gene signatures across three contrasting extreme environments: the Gilgit Hot Spring, the Kharan Desert, and the Kalabagh Salt Mine, using best-hit taxonomic and multi-database functional annotation (KEGG, eggNOG, GO, CAZy, Pfam, CYPED) combined with PERMANOVA-based statistical testing to evaluate habitat-associated community and functional divergence. Across all sites, Proteobacteria, Firmicutes, and Actinobacteria dominated as core phyla, while community composition diverged substantially at finer taxonomic resolutions, with beta-diversity analysis confirming statistically distinct communities across all three habitats (PERMANOVA: pseudo-F = 21.67, R[2] = 0.812, p = 0.001). Functional annotation revealed habitat-associated differences in gene representation. Hot spring microbiomes showed higher representation of genes associated with protein synthesis, folding, and redox-related processes. Desert communities exhibited a relatively broader representation of carbohydrate and amino acid metabolic pathways, including a higher relative abundance of carbohydrate-active enzymes. Salt mine communities showed a higher representation of genes associated with genome maintenance, membrane transport, and osmoregulation. These habitat-associated differences in functional gene representation were independently identified across multiple annotation frameworks, supporting the consistency of the observed trends. Notably, eggNOG revealed a substantial fraction of uncharacterized genes (16-19%), and CYPED analysis identified cytochrome P450 homologs with only 22-33% sequence identity to known homologs, highlighting the unexplored genetic potential of these microbiomes. Together, these findings suggest that environmental differences among habitats are associated with habitat-specific structuring of both microbial community composition and functional gene repertoires.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Nutritional intervention trial of yeast mannans derived from baker's yeast enriches Bacteroides and Parabacteroides with concurrent reduction of Klebsiella pneumoniae.
Frontiers in microbiology, 17:1958679.
INTRODUCTION: Supplementation with a yeast cell wall fraction rich in mannans is well tolerated in adults without gastrointestinal disease; however, limited research has examined its selective utilization by the gut microbiota and associated health benefits. The aim of this analysis was to investigate the specific effects of yeast mannans on the composition and metabolic function of the gut microbiota.
METHODS: Fecal samples from an open-label study of adults consuming 15 g of yeast mannans per day were profiled for fecal microbiota composition using whole metagenome sequencing, and fecal and urine metabolomics were analyzed by direct infusion and liquid chromatography-tandem mass spectrometry (DI-MS/MS and LC-MS/MS).
RESULTS: Differential abundance testing identified 46 species whose relative abundances increased with the yeast mannan intervention, primarily in the genera Bacteroides and Parabacteroides. Thomasclavelia spiroformis showed the largest fold increase. Four species decreased in relative abundance, with a greater-than-2-fold decrease in Klebsiella pneumoniae, with 45% of participants showing no detection following the yeast mannan intervention. Minor metabolomic changes were noted, including increased urinary excretion of N-acetylalanine, alanine, and caproic acid, and decreases in fecal concentrations of ether-linked phosphatidylcholines C36:4 and C40:1.
DISCUSSION: Yeast mannan supplementation selectively enriched several Bacteroides and Parabacteroides, saccharolytic commensals that may outcompete Klebsiella pneumoniae, an opportunistic pathobiont capable of causing extraintestinal infections.
Additional Links: PMID-43032102
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032102,
year = {2026},
author = {Moreno, ML and Taft, DH and Tompkins, TA and Dahl, WJ},
title = {Nutritional intervention trial of yeast mannans derived from baker's yeast enriches Bacteroides and Parabacteroides with concurrent reduction of Klebsiella pneumoniae.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1958679},
pmid = {43032102},
issn = {1664-302X},
abstract = {INTRODUCTION: Supplementation with a yeast cell wall fraction rich in mannans is well tolerated in adults without gastrointestinal disease; however, limited research has examined its selective utilization by the gut microbiota and associated health benefits. The aim of this analysis was to investigate the specific effects of yeast mannans on the composition and metabolic function of the gut microbiota.
METHODS: Fecal samples from an open-label study of adults consuming 15 g of yeast mannans per day were profiled for fecal microbiota composition using whole metagenome sequencing, and fecal and urine metabolomics were analyzed by direct infusion and liquid chromatography-tandem mass spectrometry (DI-MS/MS and LC-MS/MS).
RESULTS: Differential abundance testing identified 46 species whose relative abundances increased with the yeast mannan intervention, primarily in the genera Bacteroides and Parabacteroides. Thomasclavelia spiroformis showed the largest fold increase. Four species decreased in relative abundance, with a greater-than-2-fold decrease in Klebsiella pneumoniae, with 45% of participants showing no detection following the yeast mannan intervention. Minor metabolomic changes were noted, including increased urinary excretion of N-acetylalanine, alanine, and caproic acid, and decreases in fecal concentrations of ether-linked phosphatidylcholines C36:4 and C40:1.
DISCUSSION: Yeast mannan supplementation selectively enriched several Bacteroides and Parabacteroides, saccharolytic commensals that may outcompete Klebsiella pneumoniae, an opportunistic pathobiont capable of causing extraintestinal infections.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Clinically suspected miliary tuberculosis unveiled as disseminated nocardiosis due to Nocardia transvalensis: A case report on the utility of metagenomic next-generation sequencing in difficult-to-diagnose cases.
IDCases, 46:e02823.
Disseminated nocardiosis is an uncommon but often underrecognized cause of granulomatous, miliary-type disease that can closely mimic tuberculosis (TB) radiologically and histopathologically. This report describes a postpartum woman with chronic, non-resolving pneumonia, miliary pulmonary and renal nodularities, and a central nervous system (CNS) lesion who was repeatedly managed as disseminated TB despite extensive negative mycobacterial investigations and lack of clinical improvement. Shotgun metagenomic next-generation sequencing (mNGS) of bronchoalveolar lavage (BAL) fluid ultimately detected Nocardia transvalensis, supporting a diagnosis of disseminated nocardiosis, and the patient responded to targeted therapy. No Nocardia sp. isolate was recovered in culture; species-level identification was therefore based on mNGS interpreted in its clinical context. This difficult-to-diagnose case highlights the diagnostic value of mNGS for culture-negative disseminated infections, the importance of critically interpreting sequencing results, and the need to consider nocardiosis when TB remains unconfirmed.
Additional Links: PMID-43032151
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032151,
year = {2026},
author = {Alhumaidhi, R and AlRashed, A and Alhussaini, M and Barry, MA and Albarrag, AM and Somily, AM and Barry, M},
title = {Clinically suspected miliary tuberculosis unveiled as disseminated nocardiosis due to Nocardia transvalensis: A case report on the utility of metagenomic next-generation sequencing in difficult-to-diagnose cases.},
journal = {IDCases},
volume = {46},
number = {},
pages = {e02823},
pmid = {43032151},
issn = {2214-2509},
abstract = {Disseminated nocardiosis is an uncommon but often underrecognized cause of granulomatous, miliary-type disease that can closely mimic tuberculosis (TB) radiologically and histopathologically. This report describes a postpartum woman with chronic, non-resolving pneumonia, miliary pulmonary and renal nodularities, and a central nervous system (CNS) lesion who was repeatedly managed as disseminated TB despite extensive negative mycobacterial investigations and lack of clinical improvement. Shotgun metagenomic next-generation sequencing (mNGS) of bronchoalveolar lavage (BAL) fluid ultimately detected Nocardia transvalensis, supporting a diagnosis of disseminated nocardiosis, and the patient responded to targeted therapy. No Nocardia sp. isolate was recovered in culture; species-level identification was therefore based on mNGS interpreted in its clinical context. This difficult-to-diagnose case highlights the diagnostic value of mNGS for culture-negative disseminated infections, the importance of critically interpreting sequencing results, and the need to consider nocardiosis when TB remains unconfirmed.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Metagenomic Next-Generation Sequencing as a Robust Diagnostic Approach for Human Orf Virus Infection Undetectable by Conventional Microbiological Methods.
Open forum infectious diseases, 13(11):ofag724.
BACKGROUND: Orf virus (ORFV) is a significant pathogen known for its highly contagious nature. This study reports the case of a 55-year-old male farmer who developed fever and wound infection following an untreated sheep bite. The hospital utilized metagenomic next-generation sequencing (mNGS) technology to identify the rare pathogen-orf virus. Since orf virus cannot be detected through conventional bacterial and fungal cultures, this study underscores the importance of mNGS in identifying rare pathogens.
METHOD: The patient developed fever and wound suppuration on day 11 following a sheep-related injury and was admitted to our hospital on day 17 because of persistent intermittent fever despite prior antibiotic treatment. Magnetic resonance imaging revealed soft tissue damage. Blood cultures, Gram and Ziehl-Neelsen staining, and bacterial and fungal cultures of wound secretions yielded negative results. Subsequently, mNGS detected orf virus in the wound secretion specimen. To verify the mNGS results, a quantitative polymerase chain reaction (qPCR) analysis targeting the B2L gene of parapoxvirus was performed on the wound secretion specimen.
RESULTS: mNGS identified ORFV in the wound secretion specimen. B2L-targeted qPCR performed on the same specimen yielded a positive result, while Sanger sequencing provided limited additional sequence-level support consistent with a parapoxvirus B2L target.
CONCLUSIONS: Orf virus cannot be cultured in most diagnostic laboratory cell lines, and there is no routine clinical detection for orf virus. Therefore, mNGS emerges as a powerful tool for detecting such infections. This case demonstrates that mNGS provides an effective diagnostic strategy for infections that cannot be identified by traditional culture methods.
Additional Links: PMID-43032153
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032153,
year = {2026},
author = {Wang, J and Wang, R and Yin, Y and Cheng, J and Wang, Y and Liu, S and Zhang, X and Tu, S and Guo, Y and Wang, Q and Wang, H and Chen, H},
title = {Metagenomic Next-Generation Sequencing as a Robust Diagnostic Approach for Human Orf Virus Infection Undetectable by Conventional Microbiological Methods.},
journal = {Open forum infectious diseases},
volume = {13},
number = {11},
pages = {ofag724},
pmid = {43032153},
issn = {2328-8957},
abstract = {BACKGROUND: Orf virus (ORFV) is a significant pathogen known for its highly contagious nature. This study reports the case of a 55-year-old male farmer who developed fever and wound infection following an untreated sheep bite. The hospital utilized metagenomic next-generation sequencing (mNGS) technology to identify the rare pathogen-orf virus. Since orf virus cannot be detected through conventional bacterial and fungal cultures, this study underscores the importance of mNGS in identifying rare pathogens.
METHOD: The patient developed fever and wound suppuration on day 11 following a sheep-related injury and was admitted to our hospital on day 17 because of persistent intermittent fever despite prior antibiotic treatment. Magnetic resonance imaging revealed soft tissue damage. Blood cultures, Gram and Ziehl-Neelsen staining, and bacterial and fungal cultures of wound secretions yielded negative results. Subsequently, mNGS detected orf virus in the wound secretion specimen. To verify the mNGS results, a quantitative polymerase chain reaction (qPCR) analysis targeting the B2L gene of parapoxvirus was performed on the wound secretion specimen.
RESULTS: mNGS identified ORFV in the wound secretion specimen. B2L-targeted qPCR performed on the same specimen yielded a positive result, while Sanger sequencing provided limited additional sequence-level support consistent with a parapoxvirus B2L target.
CONCLUSIONS: Orf virus cannot be cultured in most diagnostic laboratory cell lines, and there is no routine clinical detection for orf virus. Therefore, mNGS emerges as a powerful tool for detecting such infections. This case demonstrates that mNGS provides an effective diagnostic strategy for infections that cannot be identified by traditional culture methods.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Immune-mediated necrotizing myopathy complicated by rhabdomyolysis syndrome and central nervous system cytomegalovirus infection: A case report.
Biomedical reports, 25(6):149.
Immune-mediated necrotizing myopathy (IMNM) complicated by rhabdomyolysis (RM) syndrome and central nervous system (CNS) infection is extremely rare, difficult to diagnose and associated with a poor prognosis. The present study reports a case of anti-signal recognition particle (anti-SRP) antibody-positive IMNM complicated by RM and CNS cytomegalovirus (CMV) infection to provide a reference for its clinical diagnosis and treatment. The clinical data, laboratory findings, muscle pathology, imaging findings and electromyography results are retrospectively analyzed for a 19-year-old female patient who presented with progressive fatigue and muscle weakness, and the case is discussed in the context of the relevant literature. The patient's serum creatine kinase level peaked at 32,213 U/l, and anti-SRP antibodies were positive. Muscle biopsy revealed myogenic damage with muscle fiber necrosis and C5b-9 deposition. The diagnosis of IMNM was confirmed by muscle biopsy and strongly positive anti-SRP antibodies. Metagenomic next-generation sequencing of the cerebrospinal fluid detected CMV with a relative abundance of 100%; however, viral load quantification and read numbers were not available from the assay. Serum CMV-immunoglobulin (Ig)M (213.30 AU/ml; positive) and CMV-IgG (>1,000.00 AU/ml; positive) further supported active CMV infection. The patient's muscle strength progressively improved following combined immunotherapy, antiviral therapy and supportive care. At the 6-month follow-up, upper limb muscle strength had recovered to grade 4+, while lower limb muscle strength had recovered to grade 3. In conclusion, for patients presenting with rapidly progressive muscle weakness and increased muscle enzyme levels, myositis-specific antibodies should be tested for as early as possible. When complicated by a CNS infection, differentiation from Guillain-Barré syndrome is essential. Early comprehensive immunotherapy combined with antiviral therapy may improve the prognosis.
Additional Links: PMID-43032211
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032211,
year = {2026},
author = {Xu, Y and Zeng, N and Liu, J and Zhang, Q and Hu, Q},
title = {Immune-mediated necrotizing myopathy complicated by rhabdomyolysis syndrome and central nervous system cytomegalovirus infection: A case report.},
journal = {Biomedical reports},
volume = {25},
number = {6},
pages = {149},
pmid = {43032211},
issn = {2049-9442},
abstract = {Immune-mediated necrotizing myopathy (IMNM) complicated by rhabdomyolysis (RM) syndrome and central nervous system (CNS) infection is extremely rare, difficult to diagnose and associated with a poor prognosis. The present study reports a case of anti-signal recognition particle (anti-SRP) antibody-positive IMNM complicated by RM and CNS cytomegalovirus (CMV) infection to provide a reference for its clinical diagnosis and treatment. The clinical data, laboratory findings, muscle pathology, imaging findings and electromyography results are retrospectively analyzed for a 19-year-old female patient who presented with progressive fatigue and muscle weakness, and the case is discussed in the context of the relevant literature. The patient's serum creatine kinase level peaked at 32,213 U/l, and anti-SRP antibodies were positive. Muscle biopsy revealed myogenic damage with muscle fiber necrosis and C5b-9 deposition. The diagnosis of IMNM was confirmed by muscle biopsy and strongly positive anti-SRP antibodies. Metagenomic next-generation sequencing of the cerebrospinal fluid detected CMV with a relative abundance of 100%; however, viral load quantification and read numbers were not available from the assay. Serum CMV-immunoglobulin (Ig)M (213.30 AU/ml; positive) and CMV-IgG (>1,000.00 AU/ml; positive) further supported active CMV infection. The patient's muscle strength progressively improved following combined immunotherapy, antiviral therapy and supportive care. At the 6-month follow-up, upper limb muscle strength had recovered to grade 4+, while lower limb muscle strength had recovered to grade 3. In conclusion, for patients presenting with rapidly progressive muscle weakness and increased muscle enzyme levels, myositis-specific antibodies should be tested for as early as possible. When complicated by a CNS infection, differentiation from Guillain-Barré syndrome is essential. Early comprehensive immunotherapy combined with antiviral therapy may improve the prognosis.},
}
RevDate: 2026-11-15
CmpDate: 2026-11-14
Fulminant human rhinovirus C pneumonia after allogeneic hematopoietic cell transplantation supported with venovenous extracorporeal membrane oxygenation: a case report.
Respiratory medicine case reports, 64:102535.
Human rhinovirus C (HRV-C) can cause severe lower respiratory tract disease after allogeneic hematopoietic cell transplantation (allo-HCT), but evidence supporting extracorporeal membrane oxygenation (ECMO) in this setting is sparse. A 21-year-old Han Chinese man with Philadelphia chromosome-positive B-cell acute lymphoblastic leukemia developed rapidly progressive pneumonia approximately 3 months after paternal haploidentical allo-HCT. Bilateral ground-glass opacities progressed despite empirical antimicrobial therapy. Metagenomic next-generation sequencing of bronchoalveolar lavage fluid detected HRV-C (1136 sequence reads, 440 normalized reads; estimated concentration, 2.4 × 10[4] copies/mL) without another reported pathogen. Before extracorporeal support, a final arterial blood gas showed PaO2 40 mmHg and PaCO2 80 mmHg, consistent with severe combined hypoxemic and hypercapnic respiratory failure. He also had a blood pressure of 70/35 mmHg despite norepinephrine at 0.2-0.3 μg/kg/min, serum lactate of 3.6 mmol/L, and urine output of approximately 800 mL/24 h. He underwent intubation and venovenous ECMO. Off-label adjunctive therapy comprised intravenous ribavirin 500 mg twice daily for 7 days, subcutaneous interferon alfa-2b 3 million IU once daily for 7 days, and methylprednisolone. He was extubated on November 11, ECMO was discontinued on November 12 after 9 days, and he was discharged without supplemental oxygen. This case illustrates the potential severity of HRV-C after allo-HCT and time-limited venovenous ECMO support in a carefully selected patient with a potentially reversible pulmonary process.
Additional Links: PMID-43032274
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43032274,
year = {2026},
author = {Huang, Q and Peng, Y and Zhan, Y and Zhou, T and Song, B},
title = {Fulminant human rhinovirus C pneumonia after allogeneic hematopoietic cell transplantation supported with venovenous extracorporeal membrane oxygenation: a case report.},
journal = {Respiratory medicine case reports},
volume = {64},
number = {},
pages = {102535},
pmid = {43032274},
issn = {2213-0071},
abstract = {Human rhinovirus C (HRV-C) can cause severe lower respiratory tract disease after allogeneic hematopoietic cell transplantation (allo-HCT), but evidence supporting extracorporeal membrane oxygenation (ECMO) in this setting is sparse. A 21-year-old Han Chinese man with Philadelphia chromosome-positive B-cell acute lymphoblastic leukemia developed rapidly progressive pneumonia approximately 3 months after paternal haploidentical allo-HCT. Bilateral ground-glass opacities progressed despite empirical antimicrobial therapy. Metagenomic next-generation sequencing of bronchoalveolar lavage fluid detected HRV-C (1136 sequence reads, 440 normalized reads; estimated concentration, 2.4 × 10[4] copies/mL) without another reported pathogen. Before extracorporeal support, a final arterial blood gas showed PaO2 40 mmHg and PaCO2 80 mmHg, consistent with severe combined hypoxemic and hypercapnic respiratory failure. He also had a blood pressure of 70/35 mmHg despite norepinephrine at 0.2-0.3 μg/kg/min, serum lactate of 3.6 mmol/L, and urine output of approximately 800 mL/24 h. He underwent intubation and venovenous ECMO. Off-label adjunctive therapy comprised intravenous ribavirin 500 mg twice daily for 7 days, subcutaneous interferon alfa-2b 3 million IU once daily for 7 days, and methylprednisolone. He was extubated on November 11, ECMO was discontinued on November 12 after 9 days, and he was discharged without supplemental oxygen. This case illustrates the potential severity of HRV-C after allo-HCT and time-limited venovenous ECMO support in a carefully selected patient with a potentially reversible pulmonary process.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Chromium pollution and its effects on soil health, microbial diversity, and sugarcane production in the Vellore sugarcane belt: a metagenomic and geochemical assessment.
World journal of microbiology & biotechnology, 42(12):.
Disposal of tannery effluent has elevated soil chromium (Cr) across the Vellore sugarcane belt of southern India, yet its integrated consequences for soil health, bacterial communities and sugarcane productivity remain poorly characterised. Nine composite soil samples from a recognised tannery-affected belt were pre-stratified into High, Low and Near-threshold Cr sites. Soils and irrigation water were geochemically characterised for total and speciated heavy metals and profiled by full length (V1-V9) 16 S rRNA gene amplicon sequencing on an Oxford Nanopore GridION. Soil total Cr increased from 29.10 ± 0.14 to 180.51 ± 2.95 mg kg[- 1], the alkaline-extractable Cr (VI) fraction rising from 2.92 ± 0.09 to 52.07 ± 0.11 mg kg[-1] and co-occurring Cd and Ni increasing across strata. Soil pH shifted alkaline (7.74 to 8.48), organic carbon declined by up to 31.7% and exchangeable Ca and Mg by up to 40.4%. Tissue Cr followed root > shoot > leaf but peaked at Near-threshold rather than High Cr sites, indicating bioavailability-driven uptake. Cane and stalk fresh weight declined 25.8-31.3% at High Cr sites, whereas sucrose, Brix, purity and fibre remained largely unchanged. Alpha diversity followed a threshold structured rather than linear dose response, with observed richness in High Cr levels reduced to 74-76% that in Low Cr levels alongside parallel declines in Shannon diversity and Pielou's evenness. NMDS resolved three non-overlapping community clusters. A Cr-resilient core (Sphingomonas panacis, Nitrospira lenta) persisted alongside elevated prediction of azoR/acpD and chrA abundance, defining a chromium-tolerance signature in tannery-impacted sugarcane soils.
Additional Links: PMID-43024015
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43024015,
year = {2026},
author = {Gowtham, S and Sugumaran, MP and Parameswari, E and Pragadeesh, ARU and Kalaiselvi, T and Saravanan, NA and Elayarajan, M and Selva Kumar, M and Surya, K},
title = {Chromium pollution and its effects on soil health, microbial diversity, and sugarcane production in the Vellore sugarcane belt: a metagenomic and geochemical assessment.},
journal = {World journal of microbiology & biotechnology},
volume = {42},
number = {12},
pages = {},
pmid = {43024015},
issn = {1573-0972},
mesh = {*Saccharum/growth & development/microbiology ; *Chromium/analysis/toxicity ; *Soil Microbiology ; India ; *Soil/chemistry ; *Bacteria/classification/genetics/drug effects/isolation & purification ; *Soil Pollutants/analysis ; RNA, Ribosomal, 16S/genetics ; Metagenomics ; Metals, Heavy/analysis ; Hydrogen-Ion Concentration ; Biodiversity ; Tanning ; },
abstract = {Disposal of tannery effluent has elevated soil chromium (Cr) across the Vellore sugarcane belt of southern India, yet its integrated consequences for soil health, bacterial communities and sugarcane productivity remain poorly characterised. Nine composite soil samples from a recognised tannery-affected belt were pre-stratified into High, Low and Near-threshold Cr sites. Soils and irrigation water were geochemically characterised for total and speciated heavy metals and profiled by full length (V1-V9) 16 S rRNA gene amplicon sequencing on an Oxford Nanopore GridION. Soil total Cr increased from 29.10 ± 0.14 to 180.51 ± 2.95 mg kg[- 1], the alkaline-extractable Cr (VI) fraction rising from 2.92 ± 0.09 to 52.07 ± 0.11 mg kg[-1] and co-occurring Cd and Ni increasing across strata. Soil pH shifted alkaline (7.74 to 8.48), organic carbon declined by up to 31.7% and exchangeable Ca and Mg by up to 40.4%. Tissue Cr followed root > shoot > leaf but peaked at Near-threshold rather than High Cr sites, indicating bioavailability-driven uptake. Cane and stalk fresh weight declined 25.8-31.3% at High Cr sites, whereas sucrose, Brix, purity and fibre remained largely unchanged. Alpha diversity followed a threshold structured rather than linear dose response, with observed richness in High Cr levels reduced to 74-76% that in Low Cr levels alongside parallel declines in Shannon diversity and Pielou's evenness. NMDS resolved three non-overlapping community clusters. A Cr-resilient core (Sphingomonas panacis, Nitrospira lenta) persisted alongside elevated prediction of azoR/acpD and chrA abundance, defining a chromium-tolerance signature in tannery-impacted sugarcane soils.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Saccharum/growth & development/microbiology
*Chromium/analysis/toxicity
*Soil Microbiology
India
*Soil/chemistry
*Bacteria/classification/genetics/drug effects/isolation & purification
*Soil Pollutants/analysis
RNA, Ribosomal, 16S/genetics
Metagenomics
Metals, Heavy/analysis
Hydrogen-Ion Concentration
Biodiversity
Tanning
RevDate: 2026-11-13
Environmental Microbiomes as Sources of Postbiotic Therapeutics: A One Health Perspective on Translational Applications.
Probiotics and antimicrobial proteins [Epub ahead of print].
Environmental microbiomes are reservoirs of specialized microorganisms with implications for ecosystem functionality, host health, and therapeutic discovery. Soil, marine, plant-associated, wastewater, freshwater, and extreme-environment microbiomes contain communities capable of producing bioactive metabolites and functional biomolecules. Postbiotics-non-viable microorganisms and their functional components that confer health benefits-offer potential advantages in stability, safety, scalability, and pharmaceutical development. This review examines environmental microbiomes as sources of postbiotic therapeutics within a One Health framework. A critical integrative translational review synthesized literature on environmental microbiomes, postbiotic diversity, molecular mechanisms, therapeutic applications, technological advances, and barriers to clinical translation. Evidence was organized across ecological sources, postbiotic classes, human-health applications, discovery platforms, pharmaceutical development, and translational challenges. Environmental microbiomes represent chemically and functionally diverse reservoirs of postbiotic candidates shaped by ecological adaptation, microbial competition, and environmental selection. Major classes-including short-chain fatty acids, bacteriocins, extracellular vesicles, indole and tryptophan metabolites, exopolysaccharides, and microbial/lytic enzymes-act through receptor signaling, immune modulation, barrier protection, antimicrobial activity, metabolic regulation, intercellular communication, and enzymatic target modification. Metagenomics, metabolomics, artificial intelligence, synthetic biology, organoid and microfluidic models, advanced delivery systems, and biomanufacturing increasingly enable discovery and development. However, reproducibility, microbiome variability, manufacturing scalability, regulatory standardization, cost, and equitable translation remain barriers. Environmental microbiomes provide important reservoirs for postbiotic discovery. Integrating ecological microbiology with multi-omics, computational innovation, pharmaceutical engineering, and human-relevant validation could accelerate development of defined, reproducible, and scalable postbiotic therapeutics. Preserving microbial ecosystem diversity is relevant to environmental sustainability and future microbiome-derived therapeutic innovation within One Health medicine.
Additional Links: PMID-43024025
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43024025,
year = {2026},
author = {Chigozie, VU and Oghonyon, EI and Enyi, EO},
title = {Environmental Microbiomes as Sources of Postbiotic Therapeutics: A One Health Perspective on Translational Applications.},
journal = {Probiotics and antimicrobial proteins},
volume = {},
number = {},
pages = {},
pmid = {43024025},
issn = {1867-1314},
abstract = {Environmental microbiomes are reservoirs of specialized microorganisms with implications for ecosystem functionality, host health, and therapeutic discovery. Soil, marine, plant-associated, wastewater, freshwater, and extreme-environment microbiomes contain communities capable of producing bioactive metabolites and functional biomolecules. Postbiotics-non-viable microorganisms and their functional components that confer health benefits-offer potential advantages in stability, safety, scalability, and pharmaceutical development. This review examines environmental microbiomes as sources of postbiotic therapeutics within a One Health framework. A critical integrative translational review synthesized literature on environmental microbiomes, postbiotic diversity, molecular mechanisms, therapeutic applications, technological advances, and barriers to clinical translation. Evidence was organized across ecological sources, postbiotic classes, human-health applications, discovery platforms, pharmaceutical development, and translational challenges. Environmental microbiomes represent chemically and functionally diverse reservoirs of postbiotic candidates shaped by ecological adaptation, microbial competition, and environmental selection. Major classes-including short-chain fatty acids, bacteriocins, extracellular vesicles, indole and tryptophan metabolites, exopolysaccharides, and microbial/lytic enzymes-act through receptor signaling, immune modulation, barrier protection, antimicrobial activity, metabolic regulation, intercellular communication, and enzymatic target modification. Metagenomics, metabolomics, artificial intelligence, synthetic biology, organoid and microfluidic models, advanced delivery systems, and biomanufacturing increasingly enable discovery and development. However, reproducibility, microbiome variability, manufacturing scalability, regulatory standardization, cost, and equitable translation remain barriers. Environmental microbiomes provide important reservoirs for postbiotic discovery. Integrating ecological microbiology with multi-omics, computational innovation, pharmaceutical engineering, and human-relevant validation could accelerate development of defined, reproducible, and scalable postbiotic therapeutics. Preserving microbial ecosystem diversity is relevant to environmental sustainability and future microbiome-derived therapeutic innovation within One Health medicine.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Biotin Limitation Attenuates Streptococcus mutans Cariogenicity by Disrupting Metabolic Flux and Virulence Pathways.
Caries research, 60(4):344-358.
INTRODUCTION: Our previous metagenomic analysis revealed higher frequencies of biotin-related genes (i.e., bioY, bioM, bccP) in caries-active children, indicating a potential role of biotin in caries pathogenesis. This study investigated a biotin limitation strategy against the primary cariogenic bacterium Streptococcus mutans (S. mutans), including its effects on cariogenic phenotypes, gene expression, and metabolomics.
METHODS: S. mutans UA159 was cultured in biotin-free medium supplemented with different biotin concentrations. The cariogenic phenotypes of the strain, including growth kinetics, biofilm formation, exopolysaccharide production, lactate synthesis, acid/oxidative tolerance, and membrane fluidity, were measured and compared across biotin concentrations. Biofilm architecture was visualized via confocal laser-scanning microscopy and scanning electron microscopy. Quantitative real-time polymerase chain reaction was employed to analyze the expression of genes associated with virulence and biotin metabolism. Metabolomic analysis was performed to characterize metabolic perturbations induced by biotin limitation in S. mutans.
RESULTS: Under biotin limitation, S. mutans exhibited significantly reduced cariogenic phenotypes, accompanied by cell elongation and reduced membrane fluidity. At the molecular level, biotin limitation suppressed the expression of key virulence-associated genes and induced a compensatory upregulation of genes involved in biotin uptake and biotin-dependent carboxylases. Metabolomic analysis under biotin-limited conditions in S. mutans revealed perturbed pathways in central carbon metabolism and nucleotide metabolism.
CONCLUSION: Biotin limitation significantly reduced the cariogenic potential of S. mutans by disrupting metabolic flux and virulence gene expression, highlighting biotin uptake and metabolism as potential targets for anticaries therapies.
Additional Links: PMID-43024663
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43024663,
year = {2026},
author = {Qin, Z and Yang, Z and Zhang, Y and Qi, L and Peng, Y and Deng, S and Wang, Y},
title = {Biotin Limitation Attenuates Streptococcus mutans Cariogenicity by Disrupting Metabolic Flux and Virulence Pathways.},
journal = {Caries research},
volume = {60},
number = {4},
pages = {344-358},
doi = {10.1159/000548822},
pmid = {43024663},
issn = {1421-976X},
support = {82201035//National Natural Science Foundation of China/ ; 2022C03060//Key R&D Program of Zhejiang/ ; },
mesh = {*Streptococcus mutans/pathogenicity/metabolism/genetics/drug effects/growth & development ; *Biotin/metabolism/pharmacology ; Virulence/genetics ; Biofilms/growth & development/drug effects ; *Dental Caries/microbiology ; Microscopy, Confocal ; Polysaccharides, Bacterial/metabolism ; Phenotype ; Gene Expression Regulation, Bacterial ; Microscopy, Electron, Scanning ; Real-Time Polymerase Chain Reaction ; Metabolomics ; },
abstract = {INTRODUCTION: Our previous metagenomic analysis revealed higher frequencies of biotin-related genes (i.e., bioY, bioM, bccP) in caries-active children, indicating a potential role of biotin in caries pathogenesis. This study investigated a biotin limitation strategy against the primary cariogenic bacterium Streptococcus mutans (S. mutans), including its effects on cariogenic phenotypes, gene expression, and metabolomics.
METHODS: S. mutans UA159 was cultured in biotin-free medium supplemented with different biotin concentrations. The cariogenic phenotypes of the strain, including growth kinetics, biofilm formation, exopolysaccharide production, lactate synthesis, acid/oxidative tolerance, and membrane fluidity, were measured and compared across biotin concentrations. Biofilm architecture was visualized via confocal laser-scanning microscopy and scanning electron microscopy. Quantitative real-time polymerase chain reaction was employed to analyze the expression of genes associated with virulence and biotin metabolism. Metabolomic analysis was performed to characterize metabolic perturbations induced by biotin limitation in S. mutans.
RESULTS: Under biotin limitation, S. mutans exhibited significantly reduced cariogenic phenotypes, accompanied by cell elongation and reduced membrane fluidity. At the molecular level, biotin limitation suppressed the expression of key virulence-associated genes and induced a compensatory upregulation of genes involved in biotin uptake and biotin-dependent carboxylases. Metabolomic analysis under biotin-limited conditions in S. mutans revealed perturbed pathways in central carbon metabolism and nucleotide metabolism.
CONCLUSION: Biotin limitation significantly reduced the cariogenic potential of S. mutans by disrupting metabolic flux and virulence gene expression, highlighting biotin uptake and metabolism as potential targets for anticaries therapies.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Streptococcus mutans/pathogenicity/metabolism/genetics/drug effects/growth & development
*Biotin/metabolism/pharmacology
Virulence/genetics
Biofilms/growth & development/drug effects
*Dental Caries/microbiology
Microscopy, Confocal
Polysaccharides, Bacterial/metabolism
Phenotype
Gene Expression Regulation, Bacterial
Microscopy, Electron, Scanning
Real-Time Polymerase Chain Reaction
Metabolomics
RevDate: 2026-11-13
Mechanistic insights into static magnetic field-enhanced removal of polycyclic aromatic hydrocarbons from wastewater by constructed wetlands: Computational simulation and microbial community dynamics.
Water research, 309(Pt B):127275 pii:S0043-1354(26)01946-9 [Epub ahead of print].
Microbial degradation is the dominant pathway for pollutant removal in constructed wetlands (CWs). Static magnetic fields (SMFs) were employed as a promising strategy to enhance pollutant removal, as they can effectively promote microbial activity and metabolic capacity without additional chemical amendments. Here, COMSOL-based magnetic-field simulation, plant physiological assays, extracellular polymeric substance (EPS) characterization, and metagenomics were integrated to examine the regulatory effects of SMF intensity and spatial heterogeneity on microbial community structure, functional potential, and benzo[k]fluoranthene (B[k]F) biodegradation. B[k]F removal showed an intensity-dependent threshold response, with 150 mT increasing mean removal efficiency by 14.31% relative to the control. COMSOL simulations showed that 150 mT produced the largest effective SMF zone, covering 39.53% of the reactor volume. This treatment enhanced plant photosynthetic performance, potentially supporting rhizosphere microbial metabolism through plant-derived carbon and oxygen supply and increased EPS concentrations to 30.01 ± 3.19 and 23.59 ± 4.68 mg L[-1] in the upper and lower layers, respectively. Metagenomic analysis revealed enrichment of key polycyclic aromatic hydrocarbon (PAH) degrading bacteria affiliated with Actinomycetota (45.20%). At 150 mT, enhanced tricarboxylic acid cycling and oxidative phosphorylation accelerated microbial energy metabolism and promoted conversion of high-molecular-weight B[k]F into low-molecular-weight aromatic acids. These findings suggest that optimized SMFs enhance PAH removal through coordinated plant-microbe responses, thereby providing a basis for designing magnetically intensified CWs as a chemical-free wastewater treatment strategy.
Additional Links: PMID-43025069
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025069,
year = {2026},
author = {Dong, J and Liu, P and Kang, Y and Lu, J and Zhuang, L and Xie, H and Wu, H and Hu, Z and Guo, Z and Zhang, J},
title = {Mechanistic insights into static magnetic field-enhanced removal of polycyclic aromatic hydrocarbons from wastewater by constructed wetlands: Computational simulation and microbial community dynamics.},
journal = {Water research},
volume = {309},
number = {Pt B},
pages = {127275},
doi = {10.1016/j.watres.2026.127275},
pmid = {43025069},
issn = {1879-2448},
abstract = {Microbial degradation is the dominant pathway for pollutant removal in constructed wetlands (CWs). Static magnetic fields (SMFs) were employed as a promising strategy to enhance pollutant removal, as they can effectively promote microbial activity and metabolic capacity without additional chemical amendments. Here, COMSOL-based magnetic-field simulation, plant physiological assays, extracellular polymeric substance (EPS) characterization, and metagenomics were integrated to examine the regulatory effects of SMF intensity and spatial heterogeneity on microbial community structure, functional potential, and benzo[k]fluoranthene (B[k]F) biodegradation. B[k]F removal showed an intensity-dependent threshold response, with 150 mT increasing mean removal efficiency by 14.31% relative to the control. COMSOL simulations showed that 150 mT produced the largest effective SMF zone, covering 39.53% of the reactor volume. This treatment enhanced plant photosynthetic performance, potentially supporting rhizosphere microbial metabolism through plant-derived carbon and oxygen supply and increased EPS concentrations to 30.01 ± 3.19 and 23.59 ± 4.68 mg L[-1] in the upper and lower layers, respectively. Metagenomic analysis revealed enrichment of key polycyclic aromatic hydrocarbon (PAH) degrading bacteria affiliated with Actinomycetota (45.20%). At 150 mT, enhanced tricarboxylic acid cycling and oxidative phosphorylation accelerated microbial energy metabolism and promoted conversion of high-molecular-weight B[k]F into low-molecular-weight aromatic acids. These findings suggest that optimized SMFs enhance PAH removal through coordinated plant-microbe responses, thereby providing a basis for designing magnetically intensified CWs as a chemical-free wastewater treatment strategy.},
}
RevDate: 2026-11-13
Isolation of Dehalococcoides mccartyi JC20 and genome-centric metatranscriptomic identification of Aminivibrio sp. as a candidate vitamin B12 producer.
Water research, 309(Pt B):127267 pii:S0043-1354(26)01938-X [Epub ahead of print].
Vitamin B12 is an essential cofactor for reductive dehalogenases, yet many organohalide-respiring bacteria, including Dehalococcoides mccartyi, are B12-auxotrophic and rely on syntrophic partners for B12 supply. Here, we used genome-centric metatranscriptomics to investigate the B12-cross-feeding network sustaining 1,2-dichloroethane dechlorination in a defined enrichment consortium. Sustained dechlorination proceeded in both B12-free (with other vitamins retained) and completely vitamin‑free treatments at rates of 10.03 ± 0.73 and 9.66 ± 0.05 μM Cl[-] released·day[-1], respectively, with B12 detected at 1.7-3.3 µg∙L[-1] after successive transfers. Metagenomic and metatranscriptomic analyses revealed that Aminivibrio possesses a complete de novo B12 biosynthesis pathway, which was significantly upregulated 123.38-fold under B12-limited conditions, substantially higher than the 7.98-fold upregulation observed in the established B12 producer Acetobacterium. In response to B12 deprivation, D. mccartyi significantly upregulated its B12 transporter operon (btuFCD) and sustained dechlorination via the reductive dehalogenase TceA. Most recovered MAGs were fermentative bacteria that supplied acetate and hydrogen to D. mccartyi. A pure strain of D. mccartyi, designated JC20, was further isolated by dilution-to-extinction and achieved cell densities of 10[9] cells∙mL[-1]. Collectively, these findings identify the fermentative Aminivibrio as a previously unrecognized candidate for B12 production and highlight its potential role in supporting Dehalococcoides-mediated dechlorination. This work expands our understanding of the diversity of B12-producing partners in organohalide‑respiring communities and provides insights into the microbial interactions underpinning bioremediation in oligotrophic contaminated environments.
Additional Links: PMID-43025070
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025070,
year = {2026},
author = {Yu, X and Wu, Z and Liu, G and Wang, L and Wang, L and Liu, J and Bian, Z and Wu, X and Jiang, J and Qiao, W},
title = {Isolation of Dehalococcoides mccartyi JC20 and genome-centric metatranscriptomic identification of Aminivibrio sp. as a candidate vitamin B12 producer.},
journal = {Water research},
volume = {309},
number = {Pt B},
pages = {127267},
doi = {10.1016/j.watres.2026.127267},
pmid = {43025070},
issn = {1879-2448},
abstract = {Vitamin B12 is an essential cofactor for reductive dehalogenases, yet many organohalide-respiring bacteria, including Dehalococcoides mccartyi, are B12-auxotrophic and rely on syntrophic partners for B12 supply. Here, we used genome-centric metatranscriptomics to investigate the B12-cross-feeding network sustaining 1,2-dichloroethane dechlorination in a defined enrichment consortium. Sustained dechlorination proceeded in both B12-free (with other vitamins retained) and completely vitamin‑free treatments at rates of 10.03 ± 0.73 and 9.66 ± 0.05 μM Cl[-] released·day[-1], respectively, with B12 detected at 1.7-3.3 µg∙L[-1] after successive transfers. Metagenomic and metatranscriptomic analyses revealed that Aminivibrio possesses a complete de novo B12 biosynthesis pathway, which was significantly upregulated 123.38-fold under B12-limited conditions, substantially higher than the 7.98-fold upregulation observed in the established B12 producer Acetobacterium. In response to B12 deprivation, D. mccartyi significantly upregulated its B12 transporter operon (btuFCD) and sustained dechlorination via the reductive dehalogenase TceA. Most recovered MAGs were fermentative bacteria that supplied acetate and hydrogen to D. mccartyi. A pure strain of D. mccartyi, designated JC20, was further isolated by dilution-to-extinction and achieved cell densities of 10[9] cells∙mL[-1]. Collectively, these findings identify the fermentative Aminivibrio as a previously unrecognized candidate for B12 production and highlight its potential role in supporting Dehalococcoides-mediated dechlorination. This work expands our understanding of the diversity of B12-producing partners in organohalide‑respiring communities and provides insights into the microbial interactions underpinning bioremediation in oligotrophic contaminated environments.},
}
RevDate: 2026-11-13
Resolving the mechanisms on immobilized microspheres enhancing oil cleanup under typical deep-sea conditions via tracing the ecological and evolutionary dynamics of microbial generalists and specialists.
Marine pollution bulletin, 234(Pt 1):120561 pii:S0025-326X(26)01348-2 [Epub ahead of print].
Deep-sea oil pollution has garnered global concern yet. Immobilization-assisted bioremediation depending on microbial interactions represents a critical strategy for mitigating petroleum contamination. However, the behaviors of bacteria occupying distinct ecological niches during such remediation at deep-sea conditions remain inadequately understood. Herein, via multiple sequencing techniques and biochemical detection, the ecological and evolutionary characteristics of generalists and specialists were tracked under the microsphere-driven bioremediation of oily sediment at deep-sea microcosm. Ecologically, the microspheres increased the diversity of active niche bacteria (RNA-based consortium) in sediment and introduced more stochastic processes into community assembly. Meanwhile, they reinforced the community stability of the two niche taxa by increasing the number of nodes and edges, as well as the robustness of molecular networks in sediment. Genomic analyses revealed the division of labor between generalist and specialist communities. Notably, the microspheres increased the abundance of generalists and specialists that possessed higher gene number associated with petroleum decomposition. Evolutionarily, ecological equilibrium was maintained via a reduced transition rate from specialists to generalists and an elevated diversification rate within specialists. Furthermore, global dataset analysis showed the important niche bacteria activated by the microsphere were widely distributed in the deep sea, indicating the broad applicability of microsphere-based remediation in diverse deepwater habitats. Collectively, the findings highlight that the microspheres likely govern the ecological and evolutionary traits of niche bacteria to improve system stability and function, thereby enhancing oil cleanup. This work provides a novel mechanistic perspective to guide the application and design of microsphere-mediated remediation plan against in-situ pollution.
Additional Links: PMID-43025101
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025101,
year = {2026},
author = {Qiao, Y and Pan, H and Yue, C and Tian, W and Shi, J and Sun, Y and Tao, Z and Gao, Y and Li, Y and Zang, B and Xue, CX and Liu, Y},
title = {Resolving the mechanisms on immobilized microspheres enhancing oil cleanup under typical deep-sea conditions via tracing the ecological and evolutionary dynamics of microbial generalists and specialists.},
journal = {Marine pollution bulletin},
volume = {234},
number = {Pt 1},
pages = {120561},
doi = {10.1016/j.marpolbul.2026.120561},
pmid = {43025101},
issn = {1879-3363},
abstract = {Deep-sea oil pollution has garnered global concern yet. Immobilization-assisted bioremediation depending on microbial interactions represents a critical strategy for mitigating petroleum contamination. However, the behaviors of bacteria occupying distinct ecological niches during such remediation at deep-sea conditions remain inadequately understood. Herein, via multiple sequencing techniques and biochemical detection, the ecological and evolutionary characteristics of generalists and specialists were tracked under the microsphere-driven bioremediation of oily sediment at deep-sea microcosm. Ecologically, the microspheres increased the diversity of active niche bacteria (RNA-based consortium) in sediment and introduced more stochastic processes into community assembly. Meanwhile, they reinforced the community stability of the two niche taxa by increasing the number of nodes and edges, as well as the robustness of molecular networks in sediment. Genomic analyses revealed the division of labor between generalist and specialist communities. Notably, the microspheres increased the abundance of generalists and specialists that possessed higher gene number associated with petroleum decomposition. Evolutionarily, ecological equilibrium was maintained via a reduced transition rate from specialists to generalists and an elevated diversification rate within specialists. Furthermore, global dataset analysis showed the important niche bacteria activated by the microsphere were widely distributed in the deep sea, indicating the broad applicability of microsphere-based remediation in diverse deepwater habitats. Collectively, the findings highlight that the microspheres likely govern the ecological and evolutionary traits of niche bacteria to improve system stability and function, thereby enhancing oil cleanup. This work provides a novel mechanistic perspective to guide the application and design of microsphere-mediated remediation plan against in-situ pollution.},
}
RevDate: 2026-11-13
Dietary shifts across human evolution reveal a progressive loss of gut microbial diversity and functions.
Nutrition research (New York, N.Y.), 155:214-232 pii:S0271-5317(26)00129-6 [Epub ahead of print].
This narrative review summarizes the evolutionary, archaeological, and metagenomic evidence to illustrate how significant dietary shifts, from Paleolithic hunter-gatherer lifestyles to Neolithic agricultural practices and ultimately to modern industrial diets, have gradually transformed the human gut microbiome. The objective is to position diet as a dominant influence on gut microbial evolution, and to demonstrate that the contemporary Western diet signifies a profound divergence from the nutritional context to which our microbiota has coevolved over thousands of years. The available data uncovers three main conclusions. First, hunter-gatherer communities exhibit remarkably elevated α-diversity, a predominance of Prevotella and Treponema, and an abundance of cellulose-degrading Ruminococcus, which together facilitate substantial short-chain fatty acid synthesis. Second, comparative studies reveal a notable shift from Prevotella to Bacteroides in industrialized societies, along with a substantial decline in microbial diversity and a deterioration of carbohydrate-metabolizing pathways. Third, preindustrial gut microbiota were fundamentally different from Westernized ones, emphasizing that both dietary processing and food type influenced the evolution of human gut microbiota. Collectively, the modern industrial diet has created an evolutionary discord, leading to the gradual decline of ancient microbial lineages and their metabolic capabilities. This functional deterioration, especially in fiber fermentation and short-chain fatty acid production, is mechanistically associated with the increase of chronic inflammatory and metabolic disorders. These observations establish a historical reference point for gut microbiota composition, highlighting that dietary rewilding and targeted microbial restoration are empirically supported methods for mitigating contemporary dysbiosis.
Additional Links: PMID-43025213
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025213,
year = {2026},
author = {Dey, P},
title = {Dietary shifts across human evolution reveal a progressive loss of gut microbial diversity and functions.},
journal = {Nutrition research (New York, N.Y.)},
volume = {155},
number = {},
pages = {214-232},
doi = {10.1016/j.nutres.2026.10.009},
pmid = {43025213},
issn = {1879-0739},
abstract = {This narrative review summarizes the evolutionary, archaeological, and metagenomic evidence to illustrate how significant dietary shifts, from Paleolithic hunter-gatherer lifestyles to Neolithic agricultural practices and ultimately to modern industrial diets, have gradually transformed the human gut microbiome. The objective is to position diet as a dominant influence on gut microbial evolution, and to demonstrate that the contemporary Western diet signifies a profound divergence from the nutritional context to which our microbiota has coevolved over thousands of years. The available data uncovers three main conclusions. First, hunter-gatherer communities exhibit remarkably elevated α-diversity, a predominance of Prevotella and Treponema, and an abundance of cellulose-degrading Ruminococcus, which together facilitate substantial short-chain fatty acid synthesis. Second, comparative studies reveal a notable shift from Prevotella to Bacteroides in industrialized societies, along with a substantial decline in microbial diversity and a deterioration of carbohydrate-metabolizing pathways. Third, preindustrial gut microbiota were fundamentally different from Westernized ones, emphasizing that both dietary processing and food type influenced the evolution of human gut microbiota. Collectively, the modern industrial diet has created an evolutionary discord, leading to the gradual decline of ancient microbial lineages and their metabolic capabilities. This functional deterioration, especially in fiber fermentation and short-chain fatty acid production, is mechanistically associated with the increase of chronic inflammatory and metabolic disorders. These observations establish a historical reference point for gut microbiota composition, highlighting that dietary rewilding and targeted microbial restoration are empirically supported methods for mitigating contemporary dysbiosis.},
}
RevDate: 2026-11-13
Targeting F.nucleatum with metronidazole plus tislelizumab and fruquintinib in MSS colorectal cancer: A multicenter phase 2 study.
Cancer cell pii:S1535-6108(26)00478-2 [Epub ahead of print].
Fusobacterium nucleatum has been linked to chemoresistance and immune evasion in colorectal cancer (CRC), but therapeutic strategies targeting this bacterium remain limited. In this multicenter, single-arm, phase 2 trial (NCT06356597), 33 patients with Fusobacterium nucleatum-positive, microsatellite-stable metastatic CRC received the combination treatment of tislelizumab, fruquintinib, and the antimicrobial metronidazole. The primary endpoint was met, with a median progression-free survival of 6.0 months. The objective response rate and disease control rate were 19.4% and 83.9%, respectively. Grade 3 treatment-related adverse events occurred in 30.3% of patients, with no grade 4 or 5 events. Metagenomic analysis showed that metronidazole reduced Fusobacterium nucleatum abundance. Exploratory analyses associated clinical benefit with enrichment of Lactobacillus species and nonresponse with Enterococcus. Fecal microbiota transplantation from responders sensitized germ-free mice to the treatment, which enhanced cytotoxic CD8[+] T cell differentiation and reduced terminal exhaustion. These findings support further evaluation of this combination in Fusobacterium nucleatum-positive metastatic CRC.
Additional Links: PMID-43025422
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025422,
year = {2026},
author = {Jiang, S and Lin, X and Ni, L and Zhang, Y and Chen, H and Liu, C and Yuan, H and Kang, Z and Jiao, J and Han, T and Zhuo, M and Wu, X and Zhou, J and Sun, R and Hu, M and Deng, Z and Jiang, Y and Zhang, C and Xu, P and Li, P and Li, Q and Chen, YX and Xiao, XY and Fang, JY},
title = {Targeting F.nucleatum with metronidazole plus tislelizumab and fruquintinib in MSS colorectal cancer: A multicenter phase 2 study.},
journal = {Cancer cell},
volume = {},
number = {},
pages = {},
doi = {10.1016/j.ccell.2026.10.012},
pmid = {43025422},
issn = {1878-3686},
abstract = {Fusobacterium nucleatum has been linked to chemoresistance and immune evasion in colorectal cancer (CRC), but therapeutic strategies targeting this bacterium remain limited. In this multicenter, single-arm, phase 2 trial (NCT06356597), 33 patients with Fusobacterium nucleatum-positive, microsatellite-stable metastatic CRC received the combination treatment of tislelizumab, fruquintinib, and the antimicrobial metronidazole. The primary endpoint was met, with a median progression-free survival of 6.0 months. The objective response rate and disease control rate were 19.4% and 83.9%, respectively. Grade 3 treatment-related adverse events occurred in 30.3% of patients, with no grade 4 or 5 events. Metagenomic analysis showed that metronidazole reduced Fusobacterium nucleatum abundance. Exploratory analyses associated clinical benefit with enrichment of Lactobacillus species and nonresponse with Enterococcus. Fecal microbiota transplantation from responders sensitized germ-free mice to the treatment, which enhanced cytotoxic CD8[+] T cell differentiation and reduced terminal exhaustion. These findings support further evaluation of this combination in Fusobacterium nucleatum-positive metastatic CRC.},
}
RevDate: 2026-11-13
Descriptive genomic characterization of selected Klebsiella pneumoniae-positive clinical samples from two tertiary hospitals in China by mNGS.
Infection, genetics and evolution : journal of molecular epidemiology and evolutionary genetics in infectious diseases pii:S1567-1348(26)00188-7 [Epub ahead of print].
OBJECTIVE: Metagenomic next-generation sequencing (mNGS) is an attractive method for identifying diverse pathogens and has been used for diagnosing infectious diseases. This study aimed to perform a descriptive genomic characterization of AMR-associated and virulence factor genes in a selected analytic subset of Klebsiella pneumoniae-positive clinical samples detected by mNGS, rather than to estimate the prevalence of these genes among all mNGS-positive specimens.
METHODS: Clinical samples were collected from patients at two tertiary hospitals between July 1, 2021, and July 17, 2023. Sequence types, AMR-associated genes, and virulence factors of K. pneumoniae were identified using mNGS in the selected analytic subset of samples with detectable AMR/VF gene signals and sufficient sequencing coverage.
RESULTS: Thirty-three samples were collected from 31 patients as a pre-selected analytic subset enriched for AMR/VF gene signals and sufficient sequencing coverage, rather than a representative cohort of all K. pneumoniae-positive mNGS specimensAmong these, 25/31 (80.6%) of the patients underwent invasive procedures. Multilocus Sequence Typing (MLST) analysis identified eight STs, and a phylogenetic tree further categorized the samples into three major clusters. Within this selected subset, β-lactam resistance-associated genes were detected in 30/33 samples, indicating the presence of β-lactam resistance-associated genetic determinants identified from metagenomic sequences in the analyzed specimens. These values represent detection frequencies within the selected analytic subset and should not be interpreted as prevalence estimates among all K. pneumoniae-positive mNGS specimens. Notably, 48.5%(16/33) of the samples carried five or more types of virulence genes. These samples were observed more frequently among fatal cases; however, this exploratory observation should not be interpreted as evidence of a causal relationship or prognostic association.Based on mNGS findings, empirical antimicrobial therapy was modified in 64.5% of patients. The in-hospital survival rate was 74.2%.
CONCLUSION: mNGS provides descriptive genomic information on K. pneumoniae in clinical specimens and may help identify sequence types, AMR-associated genes, and virulence factors in selected samples.
Additional Links: PMID-43025576
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025576,
year = {2026},
author = {Shan, C and Geng, T and Wang, P and Wang, L and Huang, R and Zhu, X and Zhang, Q},
title = {Descriptive genomic characterization of selected Klebsiella pneumoniae-positive clinical samples from two tertiary hospitals in China by mNGS.},
journal = {Infection, genetics and evolution : journal of molecular epidemiology and evolutionary genetics in infectious diseases},
volume = {},
number = {},
pages = {106064},
doi = {10.1016/j.meegid.2026.106064},
pmid = {43025576},
issn = {1567-7257},
abstract = {OBJECTIVE: Metagenomic next-generation sequencing (mNGS) is an attractive method for identifying diverse pathogens and has been used for diagnosing infectious diseases. This study aimed to perform a descriptive genomic characterization of AMR-associated and virulence factor genes in a selected analytic subset of Klebsiella pneumoniae-positive clinical samples detected by mNGS, rather than to estimate the prevalence of these genes among all mNGS-positive specimens.
METHODS: Clinical samples were collected from patients at two tertiary hospitals between July 1, 2021, and July 17, 2023. Sequence types, AMR-associated genes, and virulence factors of K. pneumoniae were identified using mNGS in the selected analytic subset of samples with detectable AMR/VF gene signals and sufficient sequencing coverage.
RESULTS: Thirty-three samples were collected from 31 patients as a pre-selected analytic subset enriched for AMR/VF gene signals and sufficient sequencing coverage, rather than a representative cohort of all K. pneumoniae-positive mNGS specimensAmong these, 25/31 (80.6%) of the patients underwent invasive procedures. Multilocus Sequence Typing (MLST) analysis identified eight STs, and a phylogenetic tree further categorized the samples into three major clusters. Within this selected subset, β-lactam resistance-associated genes were detected in 30/33 samples, indicating the presence of β-lactam resistance-associated genetic determinants identified from metagenomic sequences in the analyzed specimens. These values represent detection frequencies within the selected analytic subset and should not be interpreted as prevalence estimates among all K. pneumoniae-positive mNGS specimens. Notably, 48.5%(16/33) of the samples carried five or more types of virulence genes. These samples were observed more frequently among fatal cases; however, this exploratory observation should not be interpreted as evidence of a causal relationship or prognostic association.Based on mNGS findings, empirical antimicrobial therapy was modified in 64.5% of patients. The in-hospital survival rate was 74.2%.
CONCLUSION: mNGS provides descriptive genomic information on K. pneumoniae in clinical specimens and may help identify sequence types, AMR-associated genes, and virulence factors in selected samples.},
}
RevDate: 2026-11-13
Chronic ethanol feeding drives gut microbiome remodeling and influences cholestatic liver injury in mice.
The American journal of pathology pii:S0002-9440(26)00322-6 [Epub ahead of print].
Alcohol-associated liver disease (ALD) is a major cause of liver-related mortality worldwide and encompasses steatosis, cirrhosis, and alcohol-associated hepatitis (AH). AH is frequently accompanied by cholestasis, characterized by impaired bile flow and disrupted bile acid homeostasis. Although gut microbiome (GM) dysbiosis has been linked to ALD progression and cholestatic injury, its functional role in alcohol-associated cholestasis remains unclear. Mice were subjected to chronic ethanol-feeding (EF) or pair-feeding (PF) for 4 weeks. Fecal samples collected at 10 days and 4 weeks underwent shotgun metagenomic sequencing. To assess the functional impact of microbiome alterations, cecal contents from male mice after 4 weeks of EF or PF were used for fecal microbiota transplantation (FMT). Recipient mice received antibiotics during acclimatization, followed by 10 days of EF and FMT every other day. Forty-eight hours before sacrifice, mice were challenged with α-naphthylisothiocyanate (ANIT; 25 mg/kg) to induce cholestatic liver injury. Liver injury, cholestasis, and inflammation were evaluated biochemically and histologically. EF induced progressive GM remodeling, whereas PF did not. Unexpectedly, FMT from PF donors aggravated ANIT-promoted liver injury, cholestasis, and hepatic inflammation in EF recipients of both sexes. In contrast, FMT from EF donors did not significantly alter injury compared with non-FMT controls. These findings suggest that GM derived from PF and chronic EF mice differentially influence susceptibility to cholestatic liver injury in EF recipients.
Additional Links: PMID-43025638
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025638,
year = {2026},
author = {Copes, J and Godeaux, E and Yan, S},
title = {Chronic ethanol feeding drives gut microbiome remodeling and influences cholestatic liver injury in mice.},
journal = {The American journal of pathology},
volume = {},
number = {},
pages = {},
doi = {10.1016/j.ajpath.2026.10.012},
pmid = {43025638},
issn = {1525-2191},
abstract = {Alcohol-associated liver disease (ALD) is a major cause of liver-related mortality worldwide and encompasses steatosis, cirrhosis, and alcohol-associated hepatitis (AH). AH is frequently accompanied by cholestasis, characterized by impaired bile flow and disrupted bile acid homeostasis. Although gut microbiome (GM) dysbiosis has been linked to ALD progression and cholestatic injury, its functional role in alcohol-associated cholestasis remains unclear. Mice were subjected to chronic ethanol-feeding (EF) or pair-feeding (PF) for 4 weeks. Fecal samples collected at 10 days and 4 weeks underwent shotgun metagenomic sequencing. To assess the functional impact of microbiome alterations, cecal contents from male mice after 4 weeks of EF or PF were used for fecal microbiota transplantation (FMT). Recipient mice received antibiotics during acclimatization, followed by 10 days of EF and FMT every other day. Forty-eight hours before sacrifice, mice were challenged with α-naphthylisothiocyanate (ANIT; 25 mg/kg) to induce cholestatic liver injury. Liver injury, cholestasis, and inflammation were evaluated biochemically and histologically. EF induced progressive GM remodeling, whereas PF did not. Unexpectedly, FMT from PF donors aggravated ANIT-promoted liver injury, cholestasis, and hepatic inflammation in EF recipients of both sexes. In contrast, FMT from EF donors did not significantly alter injury compared with non-FMT controls. These findings suggest that GM derived from PF and chronic EF mice differentially influence susceptibility to cholestatic liver injury in EF recipients.},
}
RevDate: 2026-11-13
Ecological trade-offs of S-nZVI-assisted phytoremediation in sulfamethoxazole-copper co-contaminated soil: Plant stress, contaminant attenuation and contrasting resistome responses.
Environmental research pii:S0013-9351(26)02524-7 [Epub ahead of print].
Antibiotic-metal co-contamination can select for environmental resistomes, but how nanomaterial-assisted phytoremediation alters resistance profiles remains uncertain. We conducted a pot experiment with Tradescantia pallida (T. pallida) to evaluate an integrated S-nZVI-plant remediation system in soil co-contaminated with sulfamethoxazole (SMX) and copper (Cu) and to identify environmental and microbial factors associated with resistome responses. S-nZVI showed context-dependent effects on plant performance: biomass increased in uncontaminated soil but decreased under pollutant exposure, mainly through root biomass loss. Nevertheless, S-nZVI decreased Cu bioavailability, partially restored root cell-wall integrity, and reduced SMX and Cu concentrations in plant tissues by 6.84-28.52% and 1.27-3.30%, respectively. Lipid peroxidation generally declined and antioxidant enzyme activities partially recovered, whereas H2O2 accumulation under co-contamination indicated a treatment-dependent redox response. Metagenomic profiling showed that Cu-containing treatments increased ARG subtype richness by 52.61-93.62%, with β-lactams predominating. The integrated S-nZVI-plant treatment reduced ARG subtype richness by 45.91% under Cu contamination and by 13.82% under co-contamination. However, ARG relative abundance decreased by 23.58% under Cu contamination but increased by 18.05% under co-contamination, indicating resistome restructuring rather than uniform attenuation. Network analysis suggested that S-nZVI application was associated with increased ARG-MRG-MGE co-occurrence connectivity, whereas planting substantially weakened MGE-associated statistical connections. Partial least squares path modeling explained 84.5% of the variation in ARG abundance and identified microbial α-diversity and MRG abundance as the major positive predictors, while β-diversity exerted the largest negative total effect. These findings reveal a trade-off among contaminant attenuation, plant stress, and compartment-specific resistome responses.
Additional Links: PMID-43025784
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025784,
year = {2026},
author = {Xue, W and Zhu, Y and Al-Dhabi, NA and He, Q and Xu, R and Huang, L and Li, J and Xu, Y and Tang, W},
title = {Ecological trade-offs of S-nZVI-assisted phytoremediation in sulfamethoxazole-copper co-contaminated soil: Plant stress, contaminant attenuation and contrasting resistome responses.},
journal = {Environmental research},
volume = {},
number = {},
pages = {126187},
doi = {10.1016/j.envres.2026.126187},
pmid = {43025784},
issn = {1096-0953},
abstract = {Antibiotic-metal co-contamination can select for environmental resistomes, but how nanomaterial-assisted phytoremediation alters resistance profiles remains uncertain. We conducted a pot experiment with Tradescantia pallida (T. pallida) to evaluate an integrated S-nZVI-plant remediation system in soil co-contaminated with sulfamethoxazole (SMX) and copper (Cu) and to identify environmental and microbial factors associated with resistome responses. S-nZVI showed context-dependent effects on plant performance: biomass increased in uncontaminated soil but decreased under pollutant exposure, mainly through root biomass loss. Nevertheless, S-nZVI decreased Cu bioavailability, partially restored root cell-wall integrity, and reduced SMX and Cu concentrations in plant tissues by 6.84-28.52% and 1.27-3.30%, respectively. Lipid peroxidation generally declined and antioxidant enzyme activities partially recovered, whereas H2O2 accumulation under co-contamination indicated a treatment-dependent redox response. Metagenomic profiling showed that Cu-containing treatments increased ARG subtype richness by 52.61-93.62%, with β-lactams predominating. The integrated S-nZVI-plant treatment reduced ARG subtype richness by 45.91% under Cu contamination and by 13.82% under co-contamination. However, ARG relative abundance decreased by 23.58% under Cu contamination but increased by 18.05% under co-contamination, indicating resistome restructuring rather than uniform attenuation. Network analysis suggested that S-nZVI application was associated with increased ARG-MRG-MGE co-occurrence connectivity, whereas planting substantially weakened MGE-associated statistical connections. Partial least squares path modeling explained 84.5% of the variation in ARG abundance and identified microbial α-diversity and MRG abundance as the major positive predictors, while β-diversity exerted the largest negative total effect. These findings reveal a trade-off among contaminant attenuation, plant stress, and compartment-specific resistome responses.},
}
RevDate: 2026-11-13
Polylactic acid microplastics alter dissolved organic matter transformation in karst and non-karst lake microcosms of the Tibetan Plateau.
Environmental pollution (Barking, Essex : 1987) pii:S0269-7491(26)01924-X [Epub ahead of print].
Biodegradable plastics are widely promoted as eco-friendly alternatives, but their degradation generates biodegradable microplastics (BMPs) that threaten aquatic systems. This study compared the impacts of polylactic acid (PLA) microplastics and conventional polyethylene (PE) microplastics on dissolved organic matter (DOM) dynamics and microbial ecology in microcosms established with water from a karst and a non-karst lake on the Tibetan Plateau, using FT-ICR MS, metagenomics, machine learning and dPMD network analysis. Under extreme exposure, PLA microplastics suppressed DOM photodegradation by 28% in the karst-lake microcosms, compared with 20% for PE microplastics. PLA microplastics reshaped DOM composition by reducing recalcitrant lignin-like formulae while increasing bioavailable lipid- and protein-like formulae by 4.8- and 2.0-fold, and shifted DOM assembly toward stochastic processes. Machine learning identified PLA microplastic treatment as the dominant predictor of DOM reactivity in the karst-lake microcosms. PLA microplastics were associated with a selective decline in Dormibacteraeota and with a restructuring of key microbial guilds and putative enzymatic pathways toward carbon and nitrogen lyase reactions, whereas microbial co-occurrence networks in the non-karst-lake microcosms remained stable. These microcosm results challenge the eco-friendly narrative of biodegradable plastics by showing that, under extreme exposure, PLA can alter DOM molecular composition and potential DOM-transforming microbial groups in Karst lake water, underscoring the need for further validation under environmentally relevant field conditions.
Additional Links: PMID-43025791
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025791,
year = {2026},
author = {Zeng, H and Wang, L and Muhammad, I and Zhao, D and Li, R and Lei, Y and Sun, G},
title = {Polylactic acid microplastics alter dissolved organic matter transformation in karst and non-karst lake microcosms of the Tibetan Plateau.},
journal = {Environmental pollution (Barking, Essex : 1987)},
volume = {},
number = {},
pages = {129554},
doi = {10.1016/j.envpol.2026.129554},
pmid = {43025791},
issn = {1873-6424},
abstract = {Biodegradable plastics are widely promoted as eco-friendly alternatives, but their degradation generates biodegradable microplastics (BMPs) that threaten aquatic systems. This study compared the impacts of polylactic acid (PLA) microplastics and conventional polyethylene (PE) microplastics on dissolved organic matter (DOM) dynamics and microbial ecology in microcosms established with water from a karst and a non-karst lake on the Tibetan Plateau, using FT-ICR MS, metagenomics, machine learning and dPMD network analysis. Under extreme exposure, PLA microplastics suppressed DOM photodegradation by 28% in the karst-lake microcosms, compared with 20% for PE microplastics. PLA microplastics reshaped DOM composition by reducing recalcitrant lignin-like formulae while increasing bioavailable lipid- and protein-like formulae by 4.8- and 2.0-fold, and shifted DOM assembly toward stochastic processes. Machine learning identified PLA microplastic treatment as the dominant predictor of DOM reactivity in the karst-lake microcosms. PLA microplastics were associated with a selective decline in Dormibacteraeota and with a restructuring of key microbial guilds and putative enzymatic pathways toward carbon and nitrogen lyase reactions, whereas microbial co-occurrence networks in the non-karst-lake microcosms remained stable. These microcosm results challenge the eco-friendly narrative of biodegradable plastics by showing that, under extreme exposure, PLA can alter DOM molecular composition and potential DOM-transforming microbial groups in Karst lake water, underscoring the need for further validation under environmentally relevant field conditions.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Humic Fractions Are Linked to Distinct Arsenic-Mobilizing Host-Virome Assemblages and Enrichment of Large Temperate Phages Carrying Receiver-Only AioR-Like Loci.
Environmental microbiology, 28(11):.
Arsenic mobilization in flooded rice paddies is driven by microbial redox reactions, but how redox-active organic matter coordinates arsenic-gene-bearing hosts and soil viromes remains unresolved. We examined the effects of fulvic acid (FA), humic acid (HA) and humin (HM) on arsenic-mobilizing host-virome assemblages using sequential anoxic slurry-transfer enrichments, genome-resolved metagenomics and curated virome analysis. FA and HA produced greater late-transfer arsenite accumulation than HM or unamended soil, while treatment-associated host genomes carried contrasting arsenic redox and detoxification repertoires. We identified two large temperate phages (> 100 kb) carrying receiver-only AioR-like loci in the FA and HA enrichments, with host-prediction signals converging on Bacillaceae. These loci were supported by hallmark-rich viral scaffold context and intact-locus read coverage and remained invariant despite genome-wide polymorphism. Reanalysis of 49,649 soil-derived viral genomes showed that arsenic-associated viral genes were rare among the 9518 quality-filtered genomes (0.29%) and occurred mainly as truncated regulatory architectures, particularly AioR-like receiver and AioS-like kinase-core fragments, on putatively lytic 30-70 kb genomes. By contrast, arsenic-associated viral-gene carriers were more frequent in our enrichment viromes (3.7%). These findings link distinct humic fractions to arsenic-mobilizing host-virome assemblages and reveal a candidate regulatory layer involving rare large temperate phages carrying receiver-only AioR-like loci.
Additional Links: PMID-43025862
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43025862,
year = {2026},
author = {Qiao, J and Chen, M and Ye, M and Chen, Y and Gao, R and Fang, L and Li, F},
title = {Humic Fractions Are Linked to Distinct Arsenic-Mobilizing Host-Virome Assemblages and Enrichment of Large Temperate Phages Carrying Receiver-Only AioR-Like Loci.},
journal = {Environmental microbiology},
volume = {28},
number = {11},
pages = {},
doi = {10.1111/1462-2920.70496},
pmid = {43025862},
issn = {1462-2920},
support = {42321005//National Natural Science Foundation of China/ ; 42277119//National Natural Science Foundation of China/ ; 42377243//National Natural Science Foundation of China/ ; 2025TQ09A637//Guangdong Special Support Program for High-Level Talents/ ; 2024A1515012654//Natural Science Foundation of Guangdong Province/ ; 2023B0303000006//Guangdong Major Project of Basic and Applied Basic Research/ ; },
mesh = {*Arsenic/metabolism ; *Humic Substances/analysis ; *Bacteriophages/genetics/metabolism ; Soil Microbiology ; Genome, Viral ; Soil/chemistry ; },
abstract = {Arsenic mobilization in flooded rice paddies is driven by microbial redox reactions, but how redox-active organic matter coordinates arsenic-gene-bearing hosts and soil viromes remains unresolved. We examined the effects of fulvic acid (FA), humic acid (HA) and humin (HM) on arsenic-mobilizing host-virome assemblages using sequential anoxic slurry-transfer enrichments, genome-resolved metagenomics and curated virome analysis. FA and HA produced greater late-transfer arsenite accumulation than HM or unamended soil, while treatment-associated host genomes carried contrasting arsenic redox and detoxification repertoires. We identified two large temperate phages (> 100 kb) carrying receiver-only AioR-like loci in the FA and HA enrichments, with host-prediction signals converging on Bacillaceae. These loci were supported by hallmark-rich viral scaffold context and intact-locus read coverage and remained invariant despite genome-wide polymorphism. Reanalysis of 49,649 soil-derived viral genomes showed that arsenic-associated viral genes were rare among the 9518 quality-filtered genomes (0.29%) and occurred mainly as truncated regulatory architectures, particularly AioR-like receiver and AioS-like kinase-core fragments, on putatively lytic 30-70 kb genomes. By contrast, arsenic-associated viral-gene carriers were more frequent in our enrichment viromes (3.7%). These findings link distinct humic fractions to arsenic-mobilizing host-virome assemblages and reveal a candidate regulatory layer involving rare large temperate phages carrying receiver-only AioR-like loci.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Arsenic/metabolism
*Humic Substances/analysis
*Bacteriophages/genetics/metabolism
Soil Microbiology
Genome, Viral
Soil/chemistry
RevDate: 2026-11-14
CmpDate: 2026-11-13
Integrating whole-genome sequencing and metagenomics improves phenotype prediction of fat deposition traits in pigs.
Genetics, selection, evolution : GSE, 58(1):.
BACKGROUND: Fat deposition is a key economic trait affecting production efficiency and carcass quality in pigs and is influenced by both genetic and environmental factors. Although increasing evidence indicates that the gut microbiome contributes to host phenotypic variation, current genomic evaluation and prediction models largely rely on host genotype information and rarely account for microbiome effects. This study aimed to quantify the contribution of the gut microbiome to fat deposition traits and to evaluate whether incorporating microbiome information could improve phenotypic prediction in pigs.
RESULTS: We collected phenotypic characteristics of six fat deposition traits including the shoulder backfat thickness, the 6th-7th rib backfat thickness, the last rib backfat thickness, waist backfat thickness, the 3rd-4th last rib backfat thickness, and the average backfat thickness from 478 Duroc × Landrace × Yorkshire pigs. Using whole-genome resequencing and metagenomic sequencing, we identified 21 million single nucleotide variants and quantified the abundances of 20,336 gut microbial species across 478 pigs. Distinct fecal microbial community structures were observed between pigs with high- and low-fat deposition, and 27 bacterial species were identified as potential biomarkers associated with fat deposition. Microbiability estimates for the six backfat thickness traits ranged from 0.17 to 0.44, indicating that the gut microbiome accounted for a considerable proportion of phenotypic variance beyond host genetic contributions. In contrast, the interaction effects between the host genome and gut microbiome accounted for only a negligible proportion of phenotypic variance across the six traits. Incorporating overall microbiome information into genomic prediction models resulted in mean phenotypic predictive abilities ranging from 0.082 to 0.292, representing increases in predictive ability of 0.023 to 0.084 across traits and reductions in the mean squared error of prediction of 0.001 to 0.009 compared with the additive genetic model.
CONCLUSIONS: Our findings demonstrate that the gut microbiome makes a substantial contribution to variation in fat deposition traits in pigs. Integrating host whole-genome resequencing and gut metagenomic information yielded modest absolute gains in phenotypic predictive ability, suggesting that the gut microbiome may provide complementary information for prediction models of complex traits in pigs.
Additional Links: PMID-43026532
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43026532,
year = {2026},
author = {Liu, Y and Lin, M and Luo, J and Yang, J and Yao, Z and Lin, D and Qiu, H and Zhang, Z and Huang, S and Zheng, E and Cai, G and Wu, Z and Liu, L and Wu, JJ},
title = {Integrating whole-genome sequencing and metagenomics improves phenotype prediction of fat deposition traits in pigs.},
journal = {Genetics, selection, evolution : GSE},
volume = {58},
number = {1},
pages = {},
pmid = {43026532},
issn = {1297-9686},
support = {2025B0202080002//Natural Science Foundation of Guangdong Province project/ ; 2025-XPY-01-002//Guangdong Provincial Special Finance Project for Seed Industry Revitalization Action/ ; NYQN2025004//Guangdong Special Support Program/ ; 2019BT02N630//Local Innovative and Research Teams Project of Guangdong Province/ ; 32502844//National Natural Science Foundation of China/ ; GZB20230230//the Postdoctoral Fellowship Program of CPSF/ ; GZB20250817//the Postdoctoral Fellowship Program of CPSF/ ; },
mesh = {Animals ; *Metagenomics/methods ; Phenotype ; *Whole Genome Sequencing/methods ; Swine/genetics ; *Gastrointestinal Microbiome/genetics ; Polymorphism, Single Nucleotide ; *Adipose Tissue ; Genotype ; },
abstract = {BACKGROUND: Fat deposition is a key economic trait affecting production efficiency and carcass quality in pigs and is influenced by both genetic and environmental factors. Although increasing evidence indicates that the gut microbiome contributes to host phenotypic variation, current genomic evaluation and prediction models largely rely on host genotype information and rarely account for microbiome effects. This study aimed to quantify the contribution of the gut microbiome to fat deposition traits and to evaluate whether incorporating microbiome information could improve phenotypic prediction in pigs.
RESULTS: We collected phenotypic characteristics of six fat deposition traits including the shoulder backfat thickness, the 6th-7th rib backfat thickness, the last rib backfat thickness, waist backfat thickness, the 3rd-4th last rib backfat thickness, and the average backfat thickness from 478 Duroc × Landrace × Yorkshire pigs. Using whole-genome resequencing and metagenomic sequencing, we identified 21 million single nucleotide variants and quantified the abundances of 20,336 gut microbial species across 478 pigs. Distinct fecal microbial community structures were observed between pigs with high- and low-fat deposition, and 27 bacterial species were identified as potential biomarkers associated with fat deposition. Microbiability estimates for the six backfat thickness traits ranged from 0.17 to 0.44, indicating that the gut microbiome accounted for a considerable proportion of phenotypic variance beyond host genetic contributions. In contrast, the interaction effects between the host genome and gut microbiome accounted for only a negligible proportion of phenotypic variance across the six traits. Incorporating overall microbiome information into genomic prediction models resulted in mean phenotypic predictive abilities ranging from 0.082 to 0.292, representing increases in predictive ability of 0.023 to 0.084 across traits and reductions in the mean squared error of prediction of 0.001 to 0.009 compared with the additive genetic model.
CONCLUSIONS: Our findings demonstrate that the gut microbiome makes a substantial contribution to variation in fat deposition traits in pigs. Integrating host whole-genome resequencing and gut metagenomic information yielded modest absolute gains in phenotypic predictive ability, suggesting that the gut microbiome may provide complementary information for prediction models of complex traits in pigs.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Metagenomics/methods
Phenotype
*Whole Genome Sequencing/methods
Swine/genetics
*Gastrointestinal Microbiome/genetics
Polymorphism, Single Nucleotide
*Adipose Tissue
Genotype
RevDate: 2026-11-14
Correction: Metagenomic next-generation sequencing for pathogen detection of pulmonary infections in persons living with HIV.
AIDS research and therapy, 23(1):.
Additional Links: PMID-43026571
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43026571,
year = {2026},
author = {Zhang, Y and Guo, S and Lu, L and Li, Y and Wu, Y},
title = {Correction: Metagenomic next-generation sequencing for pathogen detection of pulmonary infections in persons living with HIV.},
journal = {AIDS research and therapy},
volume = {23},
number = {1},
pages = {},
pmid = {43026571},
issn = {1742-6405},
}
RevDate: 2026-11-13
CmpDate: 2026-11-14
A specialized microbiome enriched in nitrification and halogen cycling potential resolved from multi-omics investigation of a Caribbean suboxic and acidic marine sinkhole.
Microbiome, 14(1):.
BACKGROUND: The Luymes Sinkholes, located on the Luymes Bank northwest of Saba, represent a unique marine environment characterized by distinct biogeochemical gradients. Most noticeable is an acidic (pH ≈ 6.25) and suboxic (O2 < 60 µmol/kg) submerged 'lake' at ~ 220 m depth exhibiting extreme deviations from standard oceanographic patterns, including elevated inorganic carbon, nitrate, and phosphate, and a lower temperature (20.3 ºC vs 26.4 ºC). To determine how the microbiome within the submerged acid lake adapted to these unique conditions, we collected 59 coupled samples of size-filtered (0.22 µm) planktonic microbes for shotgun metagenomics and solid-phase extracted dissolved organic matter for untargeted metabolomics from 21 stations at depths ranging from 2 to 600 m during a 10-day research cruise in December 2019.
RESULTS: We identified a distinct microbial community within the submerged acid lake different from the non-acidic sinkholes, with increased abundances of Nitrospinota, Thermoproteota, Planctomycetota, and Actinobacteriota. We also found three enriched classes of dissolved compounds within the submerged acid lake: (1) large, (2) low-polarity, and (3) halogenated metabolites, including iodides and halobenzoic acids. Correlations between microbial taxa and metabolite classes show potential links between nitrifying archaea and halogenated compound cycling. We further supported these findings using the functional annotation of metagenome-assembled genomes (MAGs) and protein clusters showing enrichment of nitrification and halogen cycling genes, suggesting the presence of specialized degradation and biosynthetic pathways, directly connecting microbial metabolism to the observed geochemical conditions.
CONCLUSION: These findings demonstrate that the submerged acid lake contains specific microbially driven biogeochemical processes under acidified and low-oxygen conditions, providing a natural model for future extreme ocean acidification scenarios. With this multi‑omics approach, we were able to connect the microbial metabolism with this unique environment, thereby expanding our understanding of how both are interconnected. Video Abstract.
Additional Links: PMID-43026629
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43026629,
year = {2026},
author = {Boer, MD and Peets, P and van Eijnatten, AL and Nelson, CE and do Canto Brum de Menezes Pena, MM and Karancz, S and Humphreys, MP and van Duyl, FC and Meesters, EH and Snoek, BL and Melkonian, C and Haas, AF and Dutilh, BE},
title = {A specialized microbiome enriched in nitrification and halogen cycling potential resolved from multi-omics investigation of a Caribbean suboxic and acidic marine sinkhole.},
journal = {Microbiome},
volume = {14},
number = {1},
pages = {},
pmid = {43026629},
issn = {2049-2618},
mesh = {*Nitrification ; *Microbiota ; Multiomics ; Metagenomics/methods ; *Bacteria/classification/metabolism/genetics/isolation & purification ; *Halogens/metabolism ; *Lakes/microbiology/chemistry ; Metabolomics ; Hydrogen-Ion Concentration ; },
abstract = {BACKGROUND: The Luymes Sinkholes, located on the Luymes Bank northwest of Saba, represent a unique marine environment characterized by distinct biogeochemical gradients. Most noticeable is an acidic (pH ≈ 6.25) and suboxic (O2 < 60 µmol/kg) submerged 'lake' at ~ 220 m depth exhibiting extreme deviations from standard oceanographic patterns, including elevated inorganic carbon, nitrate, and phosphate, and a lower temperature (20.3 ºC vs 26.4 ºC). To determine how the microbiome within the submerged acid lake adapted to these unique conditions, we collected 59 coupled samples of size-filtered (0.22 µm) planktonic microbes for shotgun metagenomics and solid-phase extracted dissolved organic matter for untargeted metabolomics from 21 stations at depths ranging from 2 to 600 m during a 10-day research cruise in December 2019.
RESULTS: We identified a distinct microbial community within the submerged acid lake different from the non-acidic sinkholes, with increased abundances of Nitrospinota, Thermoproteota, Planctomycetota, and Actinobacteriota. We also found three enriched classes of dissolved compounds within the submerged acid lake: (1) large, (2) low-polarity, and (3) halogenated metabolites, including iodides and halobenzoic acids. Correlations between microbial taxa and metabolite classes show potential links between nitrifying archaea and halogenated compound cycling. We further supported these findings using the functional annotation of metagenome-assembled genomes (MAGs) and protein clusters showing enrichment of nitrification and halogen cycling genes, suggesting the presence of specialized degradation and biosynthetic pathways, directly connecting microbial metabolism to the observed geochemical conditions.
CONCLUSION: These findings demonstrate that the submerged acid lake contains specific microbially driven biogeochemical processes under acidified and low-oxygen conditions, providing a natural model for future extreme ocean acidification scenarios. With this multi‑omics approach, we were able to connect the microbial metabolism with this unique environment, thereby expanding our understanding of how both are interconnected. Video Abstract.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Nitrification
*Microbiota
Multiomics
Metagenomics/methods
*Bacteria/classification/metabolism/genetics/isolation & purification
*Halogens/metabolism
*Lakes/microbiology/chemistry
Metabolomics
Hydrogen-Ion Concentration
RevDate: 2026-11-14
CmpDate: 2026-11-13
A fatal case of concurrent EBV-induced meningitis, ARDS, and HLH in an immunocompetent adult: a case report.
Virology journal, 23(1):.
RATIONALE: Epstein-Barr virus (EBV) typically causes self-limited infections in immunocompetent adults. Life-threatening complications such as hemophagocytic lymphohistiocytosis (HLH), meningitis, and acute respiratory distress syndrome (ARDS) are exceptionally rare and have not been previously reported to occur concurrently in a single patient. This case highlights the catastrophic potential of EBV and underscores a critical therapeutic dilemma: balancing the need for aggressive immunosuppressive therapy to control life‑threatening HLH against the risk of exacerbating uncontrolled EBV viremia and severe infections in a critically ill patient.
PATIENT CONCERNS: A 34-year-old immunocompetent woman presented with a 10-day history of high-grade fever (39.9 °C) and persistent headache.
DIAGNOSES: The diagnosis of Epstein-Barr virus (EBV)-associated hemophagocytic lymphohistiocytosis (HLH) was confirmed by meeting 5 of 8 HLH-2004 criteria in the setting of extreme EBV viremia (1.48 × 10⁸copies/mL), with high EBV DNA load additionally detected by metagenomic next-generation sequencing (mNGS) in cerebrospinal fluid (239 EBV reads) and lower respiratory tract samples (145,386 EBV reads). Concurrently, EBV meningitis was confirmed by CSF findings (lymphocytic pleocytosis, elevated protein, and hypoglycorrhachia) alongside positive mNGS. She also met the Berlin criteria for ARDS, with bilateral pulmonary infiltrates on imaging and a PaO₂/FiO₂ ratio of 200 prior to ICU admission (on low-flow oxygen, consistent with moderate ARDS). Shortly after ICU admission, her oxygenation deteriorated to a PaO₂/FiO₂ ratio of 81 on mechanical ventilation with PEEP of 10 cmH₂O, meeting criteria for severe ARDS.
INTERVENTIONS: Management was restricted to organ support (mechanical ventilation, continuous renal replacement therapy), ganciclovir antiviral therapy, and broad-spectrum antibiotics. Crucially, guideline-directed immunomodulatory therapy for HLH (e.g., corticosteroids/etoposide per HLH-2004) was not administered due to delayed diagnosis, lack of institutional expertise, and socioeconomic constraints.
OUTCOMES: The patient's condition deteriorated rapidly, with progression to fatal multi-organ failure and death on hospital day 5 (illness day 15).
LESSONS: This case demonstrates that EBV can trigger a fulminant, lethal triad of HLH, meningitis, and ARDS even in immunocompetent hosts, which has not been previously reported in a single patient. It underscores that high-risk EBV-HLH mandates immediate immunomodulation, as antiviral and supportive care alone are catastrophically insufficient. Early recognition and overcoming barriers to administering complex, urgent therapies are essential to improving outcomes.
Additional Links: PMID-43026631
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43026631,
year = {2026},
author = {Tian, J and Dong, J and Guan, W},
title = {A fatal case of concurrent EBV-induced meningitis, ARDS, and HLH in an immunocompetent adult: a case report.},
journal = {Virology journal},
volume = {23},
number = {1},
pages = {},
pmid = {43026631},
issn = {1743-422X},
mesh = {Humans ; Female ; *Lymphohistiocytosis, Hemophagocytic/diagnosis/complications/pathology/virology ; *Epstein-Barr Virus Infections/complications/diagnosis/pathology/virology ; Adult ; *Respiratory Distress Syndrome/diagnosis/pathology/complications/virology ; Fatal Outcome ; *Herpesvirus 4, Human/isolation & purification/genetics ; Immunocompetence ; },
abstract = {RATIONALE: Epstein-Barr virus (EBV) typically causes self-limited infections in immunocompetent adults. Life-threatening complications such as hemophagocytic lymphohistiocytosis (HLH), meningitis, and acute respiratory distress syndrome (ARDS) are exceptionally rare and have not been previously reported to occur concurrently in a single patient. This case highlights the catastrophic potential of EBV and underscores a critical therapeutic dilemma: balancing the need for aggressive immunosuppressive therapy to control life‑threatening HLH against the risk of exacerbating uncontrolled EBV viremia and severe infections in a critically ill patient.
PATIENT CONCERNS: A 34-year-old immunocompetent woman presented with a 10-day history of high-grade fever (39.9 °C) and persistent headache.
DIAGNOSES: The diagnosis of Epstein-Barr virus (EBV)-associated hemophagocytic lymphohistiocytosis (HLH) was confirmed by meeting 5 of 8 HLH-2004 criteria in the setting of extreme EBV viremia (1.48 × 10⁸copies/mL), with high EBV DNA load additionally detected by metagenomic next-generation sequencing (mNGS) in cerebrospinal fluid (239 EBV reads) and lower respiratory tract samples (145,386 EBV reads). Concurrently, EBV meningitis was confirmed by CSF findings (lymphocytic pleocytosis, elevated protein, and hypoglycorrhachia) alongside positive mNGS. She also met the Berlin criteria for ARDS, with bilateral pulmonary infiltrates on imaging and a PaO₂/FiO₂ ratio of 200 prior to ICU admission (on low-flow oxygen, consistent with moderate ARDS). Shortly after ICU admission, her oxygenation deteriorated to a PaO₂/FiO₂ ratio of 81 on mechanical ventilation with PEEP of 10 cmH₂O, meeting criteria for severe ARDS.
INTERVENTIONS: Management was restricted to organ support (mechanical ventilation, continuous renal replacement therapy), ganciclovir antiviral therapy, and broad-spectrum antibiotics. Crucially, guideline-directed immunomodulatory therapy for HLH (e.g., corticosteroids/etoposide per HLH-2004) was not administered due to delayed diagnosis, lack of institutional expertise, and socioeconomic constraints.
OUTCOMES: The patient's condition deteriorated rapidly, with progression to fatal multi-organ failure and death on hospital day 5 (illness day 15).
LESSONS: This case demonstrates that EBV can trigger a fulminant, lethal triad of HLH, meningitis, and ARDS even in immunocompetent hosts, which has not been previously reported in a single patient. It underscores that high-risk EBV-HLH mandates immediate immunomodulation, as antiviral and supportive care alone are catastrophically insufficient. Early recognition and overcoming barriers to administering complex, urgent therapies are essential to improving outcomes.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
Female
*Lymphohistiocytosis, Hemophagocytic/diagnosis/complications/pathology/virology
*Epstein-Barr Virus Infections/complications/diagnosis/pathology/virology
Adult
*Respiratory Distress Syndrome/diagnosis/pathology/complications/virology
Fatal Outcome
*Herpesvirus 4, Human/isolation & purification/genetics
Immunocompetence
RevDate: 2026-11-14
CmpDate: 2026-11-14
Modulatory Effects of Water Extract from Eleutherine americana on Intestinal Health: Insights from Growth, Morphology, Microbiota and Metabolomics in Plectropomus leopardus.
Animals : an open access journal from MDPI, 16(21):.
This study evaluated dietary Eleutherine americana water extract (EAW, 0 [NC], 1 [LD], 5 [MD], and 10 [HD] mL/kg) in 480 juvenile leopard coral grouper (Plectropomus leopardus; initial body weight, 11.72 ± 0.34 g) over 8 weeks, with four replicate tanks of 30 fish per treatment. EAW did not significantly improve growth performance, but survival was higher in MD and HD than in NC (p < 0.05). Histological observations indicated more intact mucosal folds, greater crypt depth, and higher goblet-cell counts. Metabolomic analysis showed lower carnitine levels and higher phospholipid and prostaglandin metabolites, with annotated differences in bile acid biosynthesis, amino acid metabolism, and carbon metabolism pathways. Intestinal microbiota profiles showed lower relative abundances of Pseudomonadota and Basidiomycota in EAW groups, lower Actinomycetota in LD and MD, and higher Bacteroidota and Bacillota in MD. At the genus level, Bradyrhizobium, Microbacterium, and Sphingobium abundances were lower, and Afipia abundance was higher, in EAW groups. Functional profiles varied in carbohydrate, amino acid, and lipid metabolism categories. Microbiota-metabolite correlation networks identified Pseudomonadota, Actinomycetota, hexosylceramides, and carnitines as highly connected features. Network pharmacology identified pdgfra, pdgfrb, and sirt1 as candidate targets of the major EAW components eleutherol, eleutherin, and isoeleutherin. The survival and intestinal findings support further evaluation of EAW as a feed additive for intestinal health in leopard coral grouper.
Additional Links: PMID-43027094
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027094,
year = {2026},
author = {Dong, H and Zeng, X and Hu, Y and Zheng, X and Zhang, J},
title = {Modulatory Effects of Water Extract from Eleutherine americana on Intestinal Health: Insights from Growth, Morphology, Microbiota and Metabolomics in Plectropomus leopardus.},
journal = {Animals : an open access journal from MDPI},
volume = {16},
number = {21},
pages = {},
pmid = {43027094},
issn = {2076-2615},
abstract = {This study evaluated dietary Eleutherine americana water extract (EAW, 0 [NC], 1 [LD], 5 [MD], and 10 [HD] mL/kg) in 480 juvenile leopard coral grouper (Plectropomus leopardus; initial body weight, 11.72 ± 0.34 g) over 8 weeks, with four replicate tanks of 30 fish per treatment. EAW did not significantly improve growth performance, but survival was higher in MD and HD than in NC (p < 0.05). Histological observations indicated more intact mucosal folds, greater crypt depth, and higher goblet-cell counts. Metabolomic analysis showed lower carnitine levels and higher phospholipid and prostaglandin metabolites, with annotated differences in bile acid biosynthesis, amino acid metabolism, and carbon metabolism pathways. Intestinal microbiota profiles showed lower relative abundances of Pseudomonadota and Basidiomycota in EAW groups, lower Actinomycetota in LD and MD, and higher Bacteroidota and Bacillota in MD. At the genus level, Bradyrhizobium, Microbacterium, and Sphingobium abundances were lower, and Afipia abundance was higher, in EAW groups. Functional profiles varied in carbohydrate, amino acid, and lipid metabolism categories. Microbiota-metabolite correlation networks identified Pseudomonadota, Actinomycetota, hexosylceramides, and carnitines as highly connected features. Network pharmacology identified pdgfra, pdgfrb, and sirt1 as candidate targets of the major EAW components eleutherol, eleutherin, and isoeleutherin. The survival and intestinal findings support further evaluation of EAW as a feed additive for intestinal health in leopard coral grouper.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Testicular and Rumen Microbial Responses to Selenium-Enriched Malt in Heat-Exposed Simmental Bulls.
Animals : an open access journal from MDPI, 16(21):.
This study evaluated testicular and rumen microbial responses to selenium-enriched malt in heat-exposed Simmental bulls. Eighteen bulls received a basal diet alone or supplemented to provide target supplemental Se levels of 0.3 or 0.5 mg/kg dietary DM. Metagenomics included six bulls per group, host assays four per group, and RNA sequencing three per group. Compared with controls, the high-dose group had higher serum antioxidant indices and reproductive hormone concentrations, lower inflammatory markers, higher Johnsen scores, and lower TUNEL positivity. Gene-level FDR control identified 23, 1, and 4 differentially expressed genes in the low-dose versus control, high-dose versus control, and high-dose versus low-dose contrasts, respectively. Provider-reported GSEA showed negative normalized enrichment scores for steroid biosynthesis in the control-versus-low-dose and control-versus-high-dose contrasts, indicating enrichment toward the supplemented side under the provider's contrast convention. qPCR showed lower CYP11A1, CYP17A1, SCARB1, and LDLR expression at the low dose, whereas Bax, Bcl-2, and CYP17A1 protein abundance did not differ. Rumen microbial diversity and community composition differed among groups, and two post hoc selected cofactor-related pathways had lower abundance at the low dose. None of the 160 microbiome-testis or 720 microbiome-serum associations survived FDR correction. Supplementation was associated with favorable systemic and testicular histological responses and dose-specific molecular and microbial changes under heat exposure.
Additional Links: PMID-43027216
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027216,
year = {2026},
author = {Li, N and Li, F and Zhang, M and Zhang, F and Wu, C and Hu, L and Xu, J},
title = {Testicular and Rumen Microbial Responses to Selenium-Enriched Malt in Heat-Exposed Simmental Bulls.},
journal = {Animals : an open access journal from MDPI},
volume = {16},
number = {21},
pages = {},
pmid = {43027216},
issn = {2076-2615},
support = {JXSNKYJCRC202442//Jiangxi Academy of Agricultural Sciences/ ; },
abstract = {This study evaluated testicular and rumen microbial responses to selenium-enriched malt in heat-exposed Simmental bulls. Eighteen bulls received a basal diet alone or supplemented to provide target supplemental Se levels of 0.3 or 0.5 mg/kg dietary DM. Metagenomics included six bulls per group, host assays four per group, and RNA sequencing three per group. Compared with controls, the high-dose group had higher serum antioxidant indices and reproductive hormone concentrations, lower inflammatory markers, higher Johnsen scores, and lower TUNEL positivity. Gene-level FDR control identified 23, 1, and 4 differentially expressed genes in the low-dose versus control, high-dose versus control, and high-dose versus low-dose contrasts, respectively. Provider-reported GSEA showed negative normalized enrichment scores for steroid biosynthesis in the control-versus-low-dose and control-versus-high-dose contrasts, indicating enrichment toward the supplemented side under the provider's contrast convention. qPCR showed lower CYP11A1, CYP17A1, SCARB1, and LDLR expression at the low dose, whereas Bax, Bcl-2, and CYP17A1 protein abundance did not differ. Rumen microbial diversity and community composition differed among groups, and two post hoc selected cofactor-related pathways had lower abundance at the low dose. None of the 160 microbiome-testis or 720 microbiome-serum associations survived FDR correction. Supplementation was associated with favorable systemic and testicular histological responses and dose-specific molecular and microbial changes under heat exposure.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Microbial Functional Carriers and Metabolic Background Associated with Natural Musk Quality in Forest Musk Deer.
Animals : an open access journal from MDPI, 16(21):.
Whether differences in natural musk quality are associated with identifiable microbial ecological and metabolic configurations remains unclear, as studies integrating phenotype, community composition, functional sources, and non-muscone metabolic background are lacking. In this study, integrated metagenomic and GC-MS/LC-MS analyses were conducted on six natural musk samples from forest musk deer on a single farm, categorized into low-quality-associated (S1, S2, and S6) and high-quality-associated (S3, S4, and S5) groups. The two groups exhibited phenotypic differentiation in muscone, moisture, and sensory characteristics. Genus-level composition revealed quality-associated consortia with individual heterogeneity. KEGG Level 3 pathway decomposition into taxonomic carriers showed that similar functional potential could be achieved across genera, particularly lipid, ketone-body, and terpenoid pathways. After excluding muscone, candidate features related to lipid/membrane components, sterols, macrocyclic ketones, and volatiles changed coordinately along the quality axis. These results outline a multi-omics evidence chain of "phenotypic differentiation-consortium reorganization-functional carrier redistribution-background metabolic remodeling", in which the identity of the organisms carrying each function provides an additional explanatory layer beyond total pathway abundance. This exploratory cross-sectional study provides testable hypotheses on the quality-associated ecological-metabolic organization of natural musk and targets for subsequent mechanistic validation.
Additional Links: PMID-43027271
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027271,
year = {2026},
author = {Sun, S and Zhang, Z and Yang, P and Ye, S and Hu, J and Zheng, J and Zhao, Y and Li, X and Zhi, R and Dou, X and Zhang, H and Song, Y and Dou, X and An, X and Bati, H and Yao, S and Tao, Y and Tan, R and Liu, S},
title = {Microbial Functional Carriers and Metabolic Background Associated with Natural Musk Quality in Forest Musk Deer.},
journal = {Animals : an open access journal from MDPI},
volume = {16},
number = {21},
pages = {},
pmid = {43027271},
issn = {2076-2615},
support = {2025HXFWBH-LSQ-01//Zhangzhou Pientzehuang Pharmaceutical Co., Ltd./ ; },
abstract = {Whether differences in natural musk quality are associated with identifiable microbial ecological and metabolic configurations remains unclear, as studies integrating phenotype, community composition, functional sources, and non-muscone metabolic background are lacking. In this study, integrated metagenomic and GC-MS/LC-MS analyses were conducted on six natural musk samples from forest musk deer on a single farm, categorized into low-quality-associated (S1, S2, and S6) and high-quality-associated (S3, S4, and S5) groups. The two groups exhibited phenotypic differentiation in muscone, moisture, and sensory characteristics. Genus-level composition revealed quality-associated consortia with individual heterogeneity. KEGG Level 3 pathway decomposition into taxonomic carriers showed that similar functional potential could be achieved across genera, particularly lipid, ketone-body, and terpenoid pathways. After excluding muscone, candidate features related to lipid/membrane components, sterols, macrocyclic ketones, and volatiles changed coordinately along the quality axis. These results outline a multi-omics evidence chain of "phenotypic differentiation-consortium reorganization-functional carrier redistribution-background metabolic remodeling", in which the identity of the organisms carrying each function provides an additional explanatory layer beyond total pathway abundance. This exploratory cross-sectional study provides testable hypotheses on the quality-associated ecological-metabolic organization of natural musk and targets for subsequent mechanistic validation.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Hen Egg-Derived Bioactive Proteins and Peptides at the Gut Interface: From Molecular Characterization and Digestion to Microbiome Interactions and Prospective Precision Nutrition.
Foods (Basel, Switzerland), 15(21):.
Hen eggs (Gallus gallus domesticus) contain bioactive proteins, peptides and lipids, but compound characterization, mechanistic cell and animal studies, and human consumption trials have developed largely in isolation. This narrative review, supported by a documented literature search, follows one pathway: egg composition, molecular characterization, processing and digestion, bioaccessibility, intestinal and microbial interaction, host signaling, and a prospective precision-nutrition outlook. Findings are classified into translational evidence categories (a heuristic classification of experimental systems, not a ranking of study quality) and summarized in a multidimensional evidence matrix. Egg white and yolk proteins, including ovotransferrin, ovomucin and lysozyme, yield characterized peptides, notably the ovotransferrin tripeptides IRW (Ile-Arg-Trp), IQW (Ile-Gln-Trp) and LKP (Leu-Lys-Pro), and ovomucin hydrolysates, whose activities in cell and animal models are compound-specific: LKP, for example, did not reproduce the effect of IRW on insulin resistance in muscle cells. In humans, one small ileostomy study (n = 5) reports higher ileal protein digestibility for cooked than raw egg, short-term trials show that whole-egg intake raises plasma choline, and an antibiotic-suppression study shows that trimethylamine-N-oxide formation after eggs plus phosphatidylcholine depends on gut microbiota. No study identified shows that IRW, IQW or LKP are generated, survive digestion or reach the circulation after ordinary egg consumption, and only six human studies address egg intake and the gut microbiome. Dose-response, delivery-system and safety data are largely missing, and precision nutrition for egg bioactives remains a research framework. Priorities are human pharmacokinetic confirmation, longer trials with shotgun metagenomics and functional readouts, and independent safety testing.
Additional Links: PMID-43027480
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027480,
year = {2026},
author = {Abou-Emera, OK and Aladhadh, M},
title = {Hen Egg-Derived Bioactive Proteins and Peptides at the Gut Interface: From Molecular Characterization and Digestion to Microbiome Interactions and Prospective Precision Nutrition.},
journal = {Foods (Basel, Switzerland)},
volume = {15},
number = {21},
pages = {},
pmid = {43027480},
issn = {2304-8158},
support = {QU-APC-2026//Qassim University/ ; },
abstract = {Hen eggs (Gallus gallus domesticus) contain bioactive proteins, peptides and lipids, but compound characterization, mechanistic cell and animal studies, and human consumption trials have developed largely in isolation. This narrative review, supported by a documented literature search, follows one pathway: egg composition, molecular characterization, processing and digestion, bioaccessibility, intestinal and microbial interaction, host signaling, and a prospective precision-nutrition outlook. Findings are classified into translational evidence categories (a heuristic classification of experimental systems, not a ranking of study quality) and summarized in a multidimensional evidence matrix. Egg white and yolk proteins, including ovotransferrin, ovomucin and lysozyme, yield characterized peptides, notably the ovotransferrin tripeptides IRW (Ile-Arg-Trp), IQW (Ile-Gln-Trp) and LKP (Leu-Lys-Pro), and ovomucin hydrolysates, whose activities in cell and animal models are compound-specific: LKP, for example, did not reproduce the effect of IRW on insulin resistance in muscle cells. In humans, one small ileostomy study (n = 5) reports higher ileal protein digestibility for cooked than raw egg, short-term trials show that whole-egg intake raises plasma choline, and an antibiotic-suppression study shows that trimethylamine-N-oxide formation after eggs plus phosphatidylcholine depends on gut microbiota. No study identified shows that IRW, IQW or LKP are generated, survive digestion or reach the circulation after ordinary egg consumption, and only six human studies address egg intake and the gut microbiome. Dose-response, delivery-system and safety data are largely missing, and precision nutrition for egg bioactives remains a research framework. Priorities are human pharmacokinetic confirmation, longer trials with shotgun metagenomics and functional readouts, and independent safety testing.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Exploratory Amplicon-Based Metagenomic Profiling and Chemical Fingerprinting of Three Spontaneously Fermented Southern African Traditional Products.
Foods (Basel, Switzerland), 15(21):.
Traditional fermented foods are integral to African culinary heritage and nutrition, yet their microbial ecology remains underexplored in many regions. This study employed amplicon sequencing (16S rRNA and ITS) and Fourier Transform Infrared (FTIR) spectroscopy to characterize the microbial communities and biochemical profiles of spontaneously fermented cow's milk (amasi), sorghum-maize slurry (ting-type gruel), and sour maize gruel (mahewu). Amasi displayed the highest diversity, dominated by Lactococcus and Malassezia restricta. Mahewu was dominated by Clostridium sensu stricto and Malassezia restricta. The ting-variant showed low diversity with Weissella dominance and Symbiotaphrina species. FTIR spectra corroborated these patterns, highlighting lipid/protein bands in amasi and carbohydrate/organic acid signatures in the fermented cereal products. Beta-diversity analyses confirmed strong substrate-driven separation, with limited shared taxa underscoring ecological specialization. These findings reveal how raw material composition and processing shape spontaneous fermentation outcomes, while highlighting potential safety risks from opportunistic taxa and opportunities for probiotic strain isolation from beneficial taxa. Future studies are required to determine the viability and characteristics of the microbial flora in these fermented beverages to determine if their occurrence is significant. The results in this study show that metagenomic and FT-IR spectroscopy can be used to profile traditional fermented products, but this must be coupled with in-depth metabolite analyses using quantitative instruments and machine learning technologies for predictive fingerprinting to determine the identity, safety, and functionality of these culturally significant foods.
Additional Links: PMID-43027566
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027566,
year = {2026},
author = {Lynett, ML and Dickson, CA and Memory, T},
title = {Exploratory Amplicon-Based Metagenomic Profiling and Chemical Fingerprinting of Three Spontaneously Fermented Southern African Traditional Products.},
journal = {Foods (Basel, Switzerland)},
volume = {15},
number = {21},
pages = {},
pmid = {43027566},
issn = {2304-8158},
abstract = {Traditional fermented foods are integral to African culinary heritage and nutrition, yet their microbial ecology remains underexplored in many regions. This study employed amplicon sequencing (16S rRNA and ITS) and Fourier Transform Infrared (FTIR) spectroscopy to characterize the microbial communities and biochemical profiles of spontaneously fermented cow's milk (amasi), sorghum-maize slurry (ting-type gruel), and sour maize gruel (mahewu). Amasi displayed the highest diversity, dominated by Lactococcus and Malassezia restricta. Mahewu was dominated by Clostridium sensu stricto and Malassezia restricta. The ting-variant showed low diversity with Weissella dominance and Symbiotaphrina species. FTIR spectra corroborated these patterns, highlighting lipid/protein bands in amasi and carbohydrate/organic acid signatures in the fermented cereal products. Beta-diversity analyses confirmed strong substrate-driven separation, with limited shared taxa underscoring ecological specialization. These findings reveal how raw material composition and processing shape spontaneous fermentation outcomes, while highlighting potential safety risks from opportunistic taxa and opportunities for probiotic strain isolation from beneficial taxa. Future studies are required to determine the viability and characteristics of the microbial flora in these fermented beverages to determine if their occurrence is significant. The results in this study show that metagenomic and FT-IR spectroscopy can be used to profile traditional fermented products, but this must be coupled with in-depth metabolite analyses using quantitative instruments and machine learning technologies for predictive fingerprinting to determine the identity, safety, and functionality of these culturally significant foods.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Unlocking the Power of Metaproteomics: A Novel Tool for Enhancing Shorebird Conservation and Blue-Economy Management.
International journal of molecular sciences, 27(21):.
Metaproteomics, the large-scale analysis of proteins from environmental or host-associated samples, has emerged as a transformative tool for characterizing microbial communities and their functional roles. Unlike metagenomics or metatranscriptomics, metaproteomics directly captures phenotypic expression, linking molecular function to ecological and physiological outcomes. Beyond its established applications in human microbiome and dietary research, metaproteomics enables the precise identification of food-derived protein biomarkers, including the discrimination of tissue-specific origins. This level of resolution is not attainable through DNA-based approaches. Here, we applied metaproteomics for the first time to analyze the feces of shorebirds in the salt flats and salt marshes of Cadiz Bay, aiming to determine their diet with unprecedented resolution. Conventional methods, based on direct observation or fecal inspection, are labor-intensive and risk losing critical dietary information. By contrast, metaproteomics provides rapid, high-resolution insights, offering a robust alternative for ecological monitoring. Our results demonstrate the potential of this technique to transform natural area management, optimize conservation strategies for avian species, and support the sustainable development of the blue economy. This study highlights metaproteomics as an innovative, cross-sectoral approach capable of advancing both fundamental ecological research and practical conservation outcomes.
Additional Links: PMID-43027762
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027762,
year = {2026},
author = {Escobar-Niño, A and Calcis-Marzán, MV and Forján-Miguéns, A and Punta-Sánchez, I and Coste, O and Pérez-Hurtado, A and Fernández-Acero, FJ},
title = {Unlocking the Power of Metaproteomics: A Novel Tool for Enhancing Shorebird Conservation and Blue-Economy Management.},
journal = {International journal of molecular sciences},
volume = {27},
number = {21},
pages = {},
pmid = {43027762},
issn = {1422-0067},
support = {RESBIOMAR LiA 4, SP18//Plan Complementario de I+D+i en el área de Biodiversidad (PCBIO)/ ; FEDER-UCA-2024-A2-14 (2024-095 / PVA / R&D+I FEDER Junta Andalucía / PR)(BIOPAB)//Universidad de Cádiz/ ; },
mesh = {Animals ; *Proteomics/methods ; *Conservation of Natural Resources/methods ; *Birds/metabolism/microbiology ; Feces/microbiology/chemistry ; Metagenomics/methods ; Proteome ; Microbiota ; },
abstract = {Metaproteomics, the large-scale analysis of proteins from environmental or host-associated samples, has emerged as a transformative tool for characterizing microbial communities and their functional roles. Unlike metagenomics or metatranscriptomics, metaproteomics directly captures phenotypic expression, linking molecular function to ecological and physiological outcomes. Beyond its established applications in human microbiome and dietary research, metaproteomics enables the precise identification of food-derived protein biomarkers, including the discrimination of tissue-specific origins. This level of resolution is not attainable through DNA-based approaches. Here, we applied metaproteomics for the first time to analyze the feces of shorebirds in the salt flats and salt marshes of Cadiz Bay, aiming to determine their diet with unprecedented resolution. Conventional methods, based on direct observation or fecal inspection, are labor-intensive and risk losing critical dietary information. By contrast, metaproteomics provides rapid, high-resolution insights, offering a robust alternative for ecological monitoring. Our results demonstrate the potential of this technique to transform natural area management, optimize conservation strategies for avian species, and support the sustainable development of the blue economy. This study highlights metaproteomics as an innovative, cross-sectoral approach capable of advancing both fundamental ecological research and practical conservation outcomes.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Proteomics/methods
*Conservation of Natural Resources/methods
*Birds/metabolism/microbiology
Feces/microbiology/chemistry
Metagenomics/methods
Proteome
Microbiota
RevDate: 2026-11-14
CmpDate: 2026-11-14
Gut Microbiome Alterations in Alzheimer's Disease: A Shotgun Metagenomic Case-Control Study.
International journal of molecular sciences, 27(21):.
The gut microbiome has been proposed as a contributor to Alzheimer's disease (AD) pathogenesis via the microbiota-gut-brain axis, but findings across cohorts are inconsistent and scarce data exist from Central Asian populations. We used shotgun metagenomic sequencing to compare the gut microbiome of AD patients and cognitively healthy controls from Kazakhstan. In a cross-sectional case-control study (35 AD, 39 Control), we profiled taxonomic and functional (HUMAnN pathway) composition, and compared alpha/beta diversity, individual taxa and pathways, and their association with clinical and demographic variables (Mann-Whitney U test, PERMANOVA/PERMDISP, OLS regression, all BH-FDR corrected, sensitivity analysis using ANCOM-BC2), with sensitivity analyses adjusting for age and BMI and applying Bonferroni correction. Within the AD group, circulating adiponectin-measured only in this group-was examined for associations with microbiome features in an exploratory analysis. Alpha diversity was lower in AD (q < 0.05). Community composition also separated by group on PERMANOVA (R[2] = 8.6%, p = 0.001), and this separation held after adjusting for age and BMI. A total of 178 taxa differed significantly between groups (BH-FDR; 37 were also Bonferroni-significant), most prominently depletion of Faecalibacterium and enrichment of Eisenbergiella and Desulfovibrio in AD; 100 pathways differed significantly (20 Bonferroni-significant). Three clinical/demographic variables (total bilirubin, atherogenic index, BMI) were associated with specific taxa or pathways. Within the AD group, adiponectin correlated negatively with the pathway assimilatory sulfate reduction IV (rho = -0.60, q = 0.015). AD in this Central Asian cohort was associated with a less diverse, compositionally distinct gut microbiome, with depletion of butyrate-producing Firmicutes and enrichment of sulfate-reducing taxa, alongside other taxa whose direction of association differed from prior reports.
Additional Links: PMID-43027795
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027795,
year = {2026},
author = {Issilbayeva, A and Kozhakhmetov, S and Kushugulova, A and Askarova, S and Kaiyrlykyzy, A and Tuigunov, D and Dolmatova, O and Zholdassova, Z and Omarov, Y and Nurgozhin, T},
title = {Gut Microbiome Alterations in Alzheimer's Disease: A Shotgun Metagenomic Case-Control Study.},
journal = {International journal of molecular sciences},
volume = {27},
number = {21},
pages = {},
pmid = {43027795},
issn = {1422-0067},
support = {AP23489983//Ministry of Science and Higher Education of the Republic of Kazakhstan/ ; },
mesh = {Humans ; *Alzheimer Disease/microbiology ; Case-Control Studies ; Female ; *Gastrointestinal Microbiome/genetics ; *Metagenomics/methods ; Male ; Aged ; Metagenome ; Shotgun Sequencing ; Cross-Sectional Studies ; Bacteria/classification/genetics ; Kazakhstan ; },
abstract = {The gut microbiome has been proposed as a contributor to Alzheimer's disease (AD) pathogenesis via the microbiota-gut-brain axis, but findings across cohorts are inconsistent and scarce data exist from Central Asian populations. We used shotgun metagenomic sequencing to compare the gut microbiome of AD patients and cognitively healthy controls from Kazakhstan. In a cross-sectional case-control study (35 AD, 39 Control), we profiled taxonomic and functional (HUMAnN pathway) composition, and compared alpha/beta diversity, individual taxa and pathways, and their association with clinical and demographic variables (Mann-Whitney U test, PERMANOVA/PERMDISP, OLS regression, all BH-FDR corrected, sensitivity analysis using ANCOM-BC2), with sensitivity analyses adjusting for age and BMI and applying Bonferroni correction. Within the AD group, circulating adiponectin-measured only in this group-was examined for associations with microbiome features in an exploratory analysis. Alpha diversity was lower in AD (q < 0.05). Community composition also separated by group on PERMANOVA (R[2] = 8.6%, p = 0.001), and this separation held after adjusting for age and BMI. A total of 178 taxa differed significantly between groups (BH-FDR; 37 were also Bonferroni-significant), most prominently depletion of Faecalibacterium and enrichment of Eisenbergiella and Desulfovibrio in AD; 100 pathways differed significantly (20 Bonferroni-significant). Three clinical/demographic variables (total bilirubin, atherogenic index, BMI) were associated with specific taxa or pathways. Within the AD group, adiponectin correlated negatively with the pathway assimilatory sulfate reduction IV (rho = -0.60, q = 0.015). AD in this Central Asian cohort was associated with a less diverse, compositionally distinct gut microbiome, with depletion of butyrate-producing Firmicutes and enrichment of sulfate-reducing taxa, alongside other taxa whose direction of association differed from prior reports.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
*Alzheimer Disease/microbiology
Case-Control Studies
Female
*Gastrointestinal Microbiome/genetics
*Metagenomics/methods
Male
Aged
Metagenome
Shotgun Sequencing
Cross-Sectional Studies
Bacteria/classification/genetics
Kazakhstan
RevDate: 2026-11-14
CmpDate: 2026-11-14
A Tailored Sequencing Approach for Identifying SARS-CoV-2 Variants in Challenging Samples from Mink.
International journal of molecular sciences, 27(21):.
High-throughput sequencing (HTS) is an essential tool for genomic surveillance of infectious agents, including SARS-CoV-2 in farmed mink. However, sample quality varies, hindering high-quality sequencing results. In this study, six mink-origin SARS-CoV-2-positive samples were sequenced using two hybrid-capture methods (commercial and custom) and a shotgun metagenomic approach. All methods consistently identified SARS-CoV-2 variants. The metagenomic approach yielded the lowest mapping rate (up to 0.35%), breadth of coverage (97-100%), depth of coverage (9-932×), and mean mapq (30-59), and one sequence did not meet the quality criteria. In contrast, both hybrid-capture methods achieved higher mapping rates (47-99%), breadth of coverage (99.9-100%), and depth of coverage (508-115,846×). The custom hybrid-capture approach was selected because of its superior coverage uniformity, the lowest number of ambiguous bases (N), adjustability to low RNA levels and degraded material, and streamlined laboratory workflow. To further evaluate its applicability, it was applied to 35 clinical samples collected as part of routine genomic surveillance. This method generated sequencing data with high mapping rates (median 95%), depth of coverage (median 13,238×), breadth of coverage (median 100%), and mean mapq (median 60). These findings demonstrate that the custom hybrid-capture approach is a useful tool for genomic surveillance of SARS-CoV-2 in mink-origin samples.
Additional Links: PMID-43027887
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43027887,
year = {2026},
author = {Giza, A and Iwan, E and Bomba, A and Orłowska, A and Smreczak, M and Pietruk, M and Chudzik, A and Olejnik, M and Domańska-Blicharz, K},
title = {A Tailored Sequencing Approach for Identifying SARS-CoV-2 Variants in Challenging Samples from Mink.},
journal = {International journal of molecular sciences},
volume = {27},
number = {21},
pages = {},
pmid = {43027887},
issn = {1422-0067},
support = {S/445//National Veterinary Research Institute/ ; No.773830//European Union/ ; },
mesh = {Animals ; *Mink/virology ; *SARS-CoV-2/genetics/isolation & purification ; *High-Throughput Nucleotide Sequencing/methods ; *COVID-19/virology/veterinary ; Metagenomics/methods ; Genome, Viral ; },
abstract = {High-throughput sequencing (HTS) is an essential tool for genomic surveillance of infectious agents, including SARS-CoV-2 in farmed mink. However, sample quality varies, hindering high-quality sequencing results. In this study, six mink-origin SARS-CoV-2-positive samples were sequenced using two hybrid-capture methods (commercial and custom) and a shotgun metagenomic approach. All methods consistently identified SARS-CoV-2 variants. The metagenomic approach yielded the lowest mapping rate (up to 0.35%), breadth of coverage (97-100%), depth of coverage (9-932×), and mean mapq (30-59), and one sequence did not meet the quality criteria. In contrast, both hybrid-capture methods achieved higher mapping rates (47-99%), breadth of coverage (99.9-100%), and depth of coverage (508-115,846×). The custom hybrid-capture approach was selected because of its superior coverage uniformity, the lowest number of ambiguous bases (N), adjustability to low RNA levels and degraded material, and streamlined laboratory workflow. To further evaluate its applicability, it was applied to 35 clinical samples collected as part of routine genomic surveillance. This method generated sequencing data with high mapping rates (median 95%), depth of coverage (median 13,238×), breadth of coverage (median 100%), and mean mapq (median 60). These findings demonstrate that the custom hybrid-capture approach is a useful tool for genomic surveillance of SARS-CoV-2 in mink-origin samples.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Mink/virology
*SARS-CoV-2/genetics/isolation & purification
*High-Throughput Nucleotide Sequencing/methods
*COVID-19/virology/veterinary
Metagenomics/methods
Genome, Viral
RevDate: 2026-11-14
CmpDate: 2026-11-14
Global Analysis of Public Cheese Microbiomes Reveals Widespread Bacterial Taxa and Geographic Community Structure.
International journal of molecular sciences, 27(21):.
Cheese microbiomes remain difficult to compare globally because public sequencing datasets are fragmented and methodologically heterogeneous. Here, we curated 14,203 sequencing records from 233 BioProjects and retained 8910 records from 112 BioProjects for standardized reanalysis, establishing a global comparative framework for cheese microbiome research. Three taxonomic profiling pipelines (QIIME2, Kraken2 and mOTUs) recovered broadly consistent genus-level patterns dominated by lactic acid bacteria, although quantitative concordance varied between methods and study collections. Whole-genome shotgun analysis identified 12 widespread bacterial species independently detected in at least 50% of BioProjects by both Kraken2 and mOTUs, with Lactococcus lactis, Streptococcus thermophilus and Lactococcus cremoris detected in at least 84% of BioProjects in both analyses. Geographic structure remained evident after accounting for dominant community composition: sample-origin country explained 36.6% of community variation in the primary analysis and remained associated with community composition in balanced subsampling and analyses removing or continuously adjusting for five dominant lactic acid bacterial genera. However, public datasets were strongly geographically imbalanced, with European BioProjects dominating the available collection. Together, these results establish a reproducible framework and comparative baseline for investigating cheese microbiome diversity, geographic structure and cross-study patterns across globally distributed public sequencing datasets.
Additional Links: PMID-43028045
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43028045,
year = {2026},
author = {Sadurski, J and Staniszewski, A and Muyanlı, EB and Yılmaz, R and Waśko, A},
title = {Global Analysis of Public Cheese Microbiomes Reveals Widespread Bacterial Taxa and Geographic Community Structure.},
journal = {International journal of molecular sciences},
volume = {27},
number = {21},
pages = {},
pmid = {43028045},
issn = {1422-0067},
support = {SD/81/TŻ/2023//Doctoral School of the University of Life Sciences in Lublin/ ; },
mesh = {*Cheese/microbiology ; *Microbiota/genetics ; *Bacteria/classification/genetics ; Metagenomics/methods ; Phylogeny ; RNA, Ribosomal, 16S/genetics ; Geography ; Biodiversity ; },
abstract = {Cheese microbiomes remain difficult to compare globally because public sequencing datasets are fragmented and methodologically heterogeneous. Here, we curated 14,203 sequencing records from 233 BioProjects and retained 8910 records from 112 BioProjects for standardized reanalysis, establishing a global comparative framework for cheese microbiome research. Three taxonomic profiling pipelines (QIIME2, Kraken2 and mOTUs) recovered broadly consistent genus-level patterns dominated by lactic acid bacteria, although quantitative concordance varied between methods and study collections. Whole-genome shotgun analysis identified 12 widespread bacterial species independently detected in at least 50% of BioProjects by both Kraken2 and mOTUs, with Lactococcus lactis, Streptococcus thermophilus and Lactococcus cremoris detected in at least 84% of BioProjects in both analyses. Geographic structure remained evident after accounting for dominant community composition: sample-origin country explained 36.6% of community variation in the primary analysis and remained associated with community composition in balanced subsampling and analyses removing or continuously adjusting for five dominant lactic acid bacterial genera. However, public datasets were strongly geographically imbalanced, with European BioProjects dominating the available collection. Together, these results establish a reproducible framework and comparative baseline for investigating cheese microbiome diversity, geographic structure and cross-study patterns across globally distributed public sequencing datasets.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Cheese/microbiology
*Microbiota/genetics
*Bacteria/classification/genetics
Metagenomics/methods
Phylogeny
RNA, Ribosomal, 16S/genetics
Geography
Biodiversity
RevDate: 2026-11-14
CmpDate: 2026-11-14
Genotype-Associated Molecular Phenotypes Across Brain and Gut Compartments in Male 5xFAD Mice: Shared Latent Structure and the Limits of Individual-Level Inference.
International journal of molecular sciences, 27(21):.
Whether gut and brain molecular states covary within individual animals, rather than merely co-occurring at the group level, has rarely been tested in Alzheimer's disease models. We profiled the hippocampus, cortex, cerebellum, intestine and feces of 18 male 5xFAD (5 familial AD mutations) (B6SJL background) mice and wild-type littermates at 3, 6 and 9 months by label-free proteomics, and feces additionally by shotgun metagenomics. The hippocampal proteome reproduced the amyloid-associated signature (App, Apoe, Hexb, Clu, Gfap), and protein-set analysis showed elevated microglial, lysosomal, astrocytic and endoplasmic reticulum chaperone proteins in the hippocampus and cortex, with no detectable difference in synaptic or mitochondrial sets. Fecal Shannon diversity was reduced, and the bile-acid-modifying genera Romboutsia and Turicibacter were nearly absent in all 5xFAD animals from 3 months (cage nested within genotype). Multi-omics factor analysis recovered a genotype-associated factor shared across brain and gut views (exact permutation p = 0.00125) that explained at most 6.3% of variance in any view and was preprocessing-dependent. After removal of group structure, correlation burden did not exceed a within-cell permutation null, and neither did same-animal positive controls. The 5xFAD genotype thus alters every compartment examined, and a modest latent structure links them; because the design excludes only strong or dense individual-level coupling, the null result bounds design resolution rather than showing that gut and brain are uncoupled.
Additional Links: PMID-43028125
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43028125,
year = {2026},
author = {Sahin, B and Eken, Y and Sahin, F and Karaman, T and Sezerman, OU and Ozer, S and Kaya, ZZ and Karagözlü, MZ and Serdar, MA and Serteser, M and Baykal, AT},
title = {Genotype-Associated Molecular Phenotypes Across Brain and Gut Compartments in Male 5xFAD Mice: Shared Latent Structure and the Limits of Individual-Level Inference.},
journal = {International journal of molecular sciences},
volume = {27},
number = {21},
pages = {},
pmid = {43028125},
issn = {1422-0067},
support = {TSA-2024-2169//Acıbadem Mehmet Ali Aydınlar University Scientific Research Projects Commission (ABAPKO)/ ; },
mesh = {Animals ; Male ; Mice ; *Alzheimer Disease/genetics/metabolism ; *Brain/metabolism ; Phenotype ; Genotype ; Proteomics/methods ; Disease Models, Animal ; Proteome ; *Gastrointestinal Microbiome ; Mice, Transgenic ; Feces/microbiology ; Hippocampus/metabolism ; Metagenomics ; Multiomics ; },
abstract = {Whether gut and brain molecular states covary within individual animals, rather than merely co-occurring at the group level, has rarely been tested in Alzheimer's disease models. We profiled the hippocampus, cortex, cerebellum, intestine and feces of 18 male 5xFAD (5 familial AD mutations) (B6SJL background) mice and wild-type littermates at 3, 6 and 9 months by label-free proteomics, and feces additionally by shotgun metagenomics. The hippocampal proteome reproduced the amyloid-associated signature (App, Apoe, Hexb, Clu, Gfap), and protein-set analysis showed elevated microglial, lysosomal, astrocytic and endoplasmic reticulum chaperone proteins in the hippocampus and cortex, with no detectable difference in synaptic or mitochondrial sets. Fecal Shannon diversity was reduced, and the bile-acid-modifying genera Romboutsia and Turicibacter were nearly absent in all 5xFAD animals from 3 months (cage nested within genotype). Multi-omics factor analysis recovered a genotype-associated factor shared across brain and gut views (exact permutation p = 0.00125) that explained at most 6.3% of variance in any view and was preprocessing-dependent. After removal of group structure, correlation burden did not exceed a within-cell permutation null, and neither did same-animal positive controls. The 5xFAD genotype thus alters every compartment examined, and a modest latent structure links them; because the design excludes only strong or dense individual-level coupling, the null result bounds design resolution rather than showing that gut and brain are uncoupled.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
Male
Mice
*Alzheimer Disease/genetics/metabolism
*Brain/metabolism
Phenotype
Genotype
Proteomics/methods
Disease Models, Animal
Proteome
*Gastrointestinal Microbiome
Mice, Transgenic
Feces/microbiology
Hippocampus/metabolism
Metagenomics
Multiomics
RevDate: 2026-11-14
CmpDate: 2026-11-14
Paper-Mulberry-Silage-Enhanced Lipid Metabolism and Milk Polyunsaturated Fatty Acid Concentration in Dairy Cows: A Multi-Omics Study.
International journal of molecular sciences, 27(21):.
Developing alternative forages that modify milk fatty acid composition is important for improving milk nutritional quality and diversifying feed resources in dairy production. Previous studies have demonstrated the potential of paper mulberry (PM, Broussonetia papyrifera) silage to partially replace conventional roughages and influence antioxidant status and milk fatty acid composition. However, its effects on milk polyunsaturated fatty acids (PUFAs) and the associated ruminal microbial and metabolic mechanisms remain insufficiently characterized. This study aimed to evaluate the effects of feeding PM silage on the concentration of polyunsaturated fatty acids in milk, the structure of the rumen microbial community, and lipid metabolic functions, as well as to analyze the associations between rumen microbial and metabolic profiles and the fatty acid composition of milk. Thirty lactating Holstein cows were randomly assigned to control and PM groups, with 15 cows per group. The PM group received diets containing 9% PM silage for the first 28 days and 18% for the subsequent 28 days on a dry matter basis. Rumen fluid, plasma, and milk were collected on day 56 for integrated analyses combining fatty acid profiling, ruminal fermentation measurements, 16S rRNA gene sequencing, metagenomics, and metabolomics. Milk PUFA concentration was significantly higher in the PM group than in controls, with no significant differences in dry matter intake or milk yield. PM feeding also altered ruminal microbial community composition and lipid-metabolism-related functional gene abundances, accompanied by increased glycerophosphocholine and glycerophosphoethanolamine levels. Integrated analyses linked these changes to fatty acid profiles in rumen fluid, plasma, and milk, supporting ruminal lipid remodeling as a candidate mechanism underlying the increase in milk PUFA concentrations. These findings provide evidence for the microbial and metabolic basis of the effects of PM silage on milk fatty acid composition and inform its development as a functional forage and the optimization of dairy cow feeding strategies.
Additional Links: PMID-43028184
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43028184,
year = {2026},
author = {Yang, S and Tang, W and Wang, Y and Yang, H and Wang, W and Cao, Z and Wang, W and Li, S and Bai, Y and Hao, Y},
title = {Paper-Mulberry-Silage-Enhanced Lipid Metabolism and Milk Polyunsaturated Fatty Acid Concentration in Dairy Cows: A Multi-Omics Study.},
journal = {International journal of molecular sciences},
volume = {27},
number = {21},
pages = {},
pmid = {43028184},
issn = {1422-0067},
support = {BX2025003//China Postdoctoral Science Foundation/ ; 2022YFD1301001//National Natural Science Foundation of China/ ; 241111110100//Henan Science and Technology Department/ ; 242102111039//Henan Science and Technology Department/ ; },
mesh = {Animals ; *Milk/chemistry/metabolism ; Cattle ; *Fatty Acids, Unsaturated/metabolism/analysis ; Female ; *Lipid Metabolism ; *Silage/analysis ; *Morus/chemistry ; Rumen/microbiology/metabolism ; Multiomics ; Lactation ; Animal Feed ; },
abstract = {Developing alternative forages that modify milk fatty acid composition is important for improving milk nutritional quality and diversifying feed resources in dairy production. Previous studies have demonstrated the potential of paper mulberry (PM, Broussonetia papyrifera) silage to partially replace conventional roughages and influence antioxidant status and milk fatty acid composition. However, its effects on milk polyunsaturated fatty acids (PUFAs) and the associated ruminal microbial and metabolic mechanisms remain insufficiently characterized. This study aimed to evaluate the effects of feeding PM silage on the concentration of polyunsaturated fatty acids in milk, the structure of the rumen microbial community, and lipid metabolic functions, as well as to analyze the associations between rumen microbial and metabolic profiles and the fatty acid composition of milk. Thirty lactating Holstein cows were randomly assigned to control and PM groups, with 15 cows per group. The PM group received diets containing 9% PM silage for the first 28 days and 18% for the subsequent 28 days on a dry matter basis. Rumen fluid, plasma, and milk were collected on day 56 for integrated analyses combining fatty acid profiling, ruminal fermentation measurements, 16S rRNA gene sequencing, metagenomics, and metabolomics. Milk PUFA concentration was significantly higher in the PM group than in controls, with no significant differences in dry matter intake or milk yield. PM feeding also altered ruminal microbial community composition and lipid-metabolism-related functional gene abundances, accompanied by increased glycerophosphocholine and glycerophosphoethanolamine levels. Integrated analyses linked these changes to fatty acid profiles in rumen fluid, plasma, and milk, supporting ruminal lipid remodeling as a candidate mechanism underlying the increase in milk PUFA concentrations. These findings provide evidence for the microbial and metabolic basis of the effects of PM silage on milk fatty acid composition and inform its development as a functional forage and the optimization of dairy cow feeding strategies.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Milk/chemistry/metabolism
Cattle
*Fatty Acids, Unsaturated/metabolism/analysis
Female
*Lipid Metabolism
*Silage/analysis
*Morus/chemistry
Rumen/microbiology/metabolism
Multiomics
Lactation
Animal Feed
RevDate: 2026-11-14
CmpDate: 2026-11-14
Legionella Pneumonia in the Molecular Diagnostics Era: From Intracellular Pathogenesis to Precision Management.
Diagnostics (Basel, Switzerland), 16(21):.
Legionella pneumophila is a major cause of severe community-acquired pneumonia and one of the most lethal respiratory pathogens. Nevertheless, it remains substantially underdiagnosed because conventional diagnostic assays are slow, insensitive, or limited to specific serogroups. In this narrative review, we integrate fragmented literature regarding Legionella pneumonia, covering its intracellular pathogenesis, clinical spectrum and patient management. Evidence was identified through a structured search of PubMed/MEDLINE and Europe PMC (23 September 2026) and prioritized by design, with primary clinical and mechanistic studies and systematic reviews taking precedence over narrative summaries and with society guidelines used to anchor management recommendations. We argue that this field is undergoing a diagnostic paradigm shift fueled by culture-independent molecular assays. We first review the Dot/Icm type IV secretion system and intracellular lifecycle that allow Legionella to replicate within macrophages while evading host innate immunity. We then summarize the full clinical spectrum, ranging from self-limited Pontiac fever to fatal Legionnaires' disease complicated by acute respiratory distress syndrome, as well as epidemiological characteristics including waterborne transmission and outbreak risks. The core of this review examines how metagenomic next-generation sequencing, targeted NGS, plasma microbial cell-free DNA and syndromic multiplex PCR are reshaping diagnostic workflows, and compares the strengths and limitations of these novel tools with urinary antigen testing and bacterial culture. Pooled estimates place the sensitivity of urinary antigen testing at 0.79 overall and 0.86 for serogroup 1, and that of respiratory PCR at 97.4%, with specificities of 1.00 and 0.99, respectively. We further evaluate contemporary therapeutic evidence, including fluoroquinolone versus macrolide monotherapy, combination regimens, and adjunctive corticosteroid therapy. Throughout this review, we highlight that Legionella is a pathogen whose clinical presentation, diagnostic yield and prognosis are highly dependent on host immune status, exposure setting and appropriate diagnostic stewardship. We conclude by identifying critical knowledge gaps and outlining a framework for diagnosis-driven precision antimicrobial management of this underrecognized pathogen. That framework is an evidence-informed synthesis rather than a validated pathway: no prospective study has yet shown that it improves patient outcomes, and its cost-effectiveness has not been measured.
Additional Links: PMID-43028548
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43028548,
year = {2026},
author = {Liu, L and Zhang, B and Sang, K and Lin, J and Li, M and Zhong, P and Huang, N and Lai, J and Xu, L and Chen, L and Liu, Y and Chen, S},
title = {Legionella Pneumonia in the Molecular Diagnostics Era: From Intracellular Pathogenesis to Precision Management.},
journal = {Diagnostics (Basel, Switzerland)},
volume = {16},
number = {21},
pages = {},
pmid = {43028548},
issn = {2075-4418},
abstract = {Legionella pneumophila is a major cause of severe community-acquired pneumonia and one of the most lethal respiratory pathogens. Nevertheless, it remains substantially underdiagnosed because conventional diagnostic assays are slow, insensitive, or limited to specific serogroups. In this narrative review, we integrate fragmented literature regarding Legionella pneumonia, covering its intracellular pathogenesis, clinical spectrum and patient management. Evidence was identified through a structured search of PubMed/MEDLINE and Europe PMC (23 September 2026) and prioritized by design, with primary clinical and mechanistic studies and systematic reviews taking precedence over narrative summaries and with society guidelines used to anchor management recommendations. We argue that this field is undergoing a diagnostic paradigm shift fueled by culture-independent molecular assays. We first review the Dot/Icm type IV secretion system and intracellular lifecycle that allow Legionella to replicate within macrophages while evading host innate immunity. We then summarize the full clinical spectrum, ranging from self-limited Pontiac fever to fatal Legionnaires' disease complicated by acute respiratory distress syndrome, as well as epidemiological characteristics including waterborne transmission and outbreak risks. The core of this review examines how metagenomic next-generation sequencing, targeted NGS, plasma microbial cell-free DNA and syndromic multiplex PCR are reshaping diagnostic workflows, and compares the strengths and limitations of these novel tools with urinary antigen testing and bacterial culture. Pooled estimates place the sensitivity of urinary antigen testing at 0.79 overall and 0.86 for serogroup 1, and that of respiratory PCR at 97.4%, with specificities of 1.00 and 0.99, respectively. We further evaluate contemporary therapeutic evidence, including fluoroquinolone versus macrolide monotherapy, combination regimens, and adjunctive corticosteroid therapy. Throughout this review, we highlight that Legionella is a pathogen whose clinical presentation, diagnostic yield and prognosis are highly dependent on host immune status, exposure setting and appropriate diagnostic stewardship. We conclude by identifying critical knowledge gaps and outlining a framework for diagnosis-driven precision antimicrobial management of this underrecognized pathogen. That framework is an evidence-informed synthesis rather than a validated pathway: no prospective study has yet shown that it improves patient outcomes, and its cost-effectiveness has not been measured.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Taxonomic Guilds, Action Mechanisms, and Agronomic Deployment of Maize Rhizosphere Microbiota for Yield Enhancement: Unraveling G×M×E Interactions and Translational Barriers via Multi Omics.
Plants (Basel, Switzerland), 15(21):.
Maize (Zea mays L.) productivity is increasingly constrained by edaphic stressors-including soil salinization, drought, and acidification-posing mounting threats to global food security. Plant growth-promoting microorganisms (PGPM) offer a sustainable alternative to synthetic fertilizers, yet despite their pronounced performance under controlled laboratory settings, their beneficial effects often prove inconsistent in open-field environments, largely due to spatial heterogeneity and genotype-specific responses. Herein, we present a comprehensive synthesis of the core microbial guilds that have been robustly associated with maize yield enhancement. The mechanisms underlying these effects are examined across several dimensions, including biological nitrogen fixation, phosphate and potassium solubilization, phytohormone production, ethylene regulation, stress tolerance, and rhizosphere colonization. Genomic, transcriptomic, metagenomic, and metabolomic data are integrated to construct an interaction network between maize and rhizosphere microbes. We identify the genotype-by-microbiome-by-environment (G×M×E) interplay as the principal bottleneck hampering field-scale translational success. To address this issue, three priorities are proposed: (i) designing synthetic microbial consortia, (ii) using multi-omics to examine mechanisms, and (iii) developing tools for site-specific deployment. In conclusion, these sections provide a framework for developing more reliable microbial inoculants for maize.
Additional Links: PMID-43028980
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43028980,
year = {2026},
author = {Lei, J and Lv, N and Liu, P and Liu, C and Peng, G},
title = {Taxonomic Guilds, Action Mechanisms, and Agronomic Deployment of Maize Rhizosphere Microbiota for Yield Enhancement: Unraveling G×M×E Interactions and Translational Barriers via Multi Omics.},
journal = {Plants (Basel, Switzerland)},
volume = {15},
number = {21},
pages = {},
pmid = {43028980},
issn = {2223-7747},
support = {CSTB2025TIAD-LYKJXZLHGGXO004//Chongqing Key Special Project of Technological Innovation and Application Development/ ; NW-CSTB2025NYKJGGXMLSGXZYLXMX0001//the Agricultural Science and Technology R&D Program of Chongqing/ ; WSWZZ 2020004//Chongqing Entomopathogenic Fungi Strain Resource Bank/ ; },
abstract = {Maize (Zea mays L.) productivity is increasingly constrained by edaphic stressors-including soil salinization, drought, and acidification-posing mounting threats to global food security. Plant growth-promoting microorganisms (PGPM) offer a sustainable alternative to synthetic fertilizers, yet despite their pronounced performance under controlled laboratory settings, their beneficial effects often prove inconsistent in open-field environments, largely due to spatial heterogeneity and genotype-specific responses. Herein, we present a comprehensive synthesis of the core microbial guilds that have been robustly associated with maize yield enhancement. The mechanisms underlying these effects are examined across several dimensions, including biological nitrogen fixation, phosphate and potassium solubilization, phytohormone production, ethylene regulation, stress tolerance, and rhizosphere colonization. Genomic, transcriptomic, metagenomic, and metabolomic data are integrated to construct an interaction network between maize and rhizosphere microbes. We identify the genotype-by-microbiome-by-environment (G×M×E) interplay as the principal bottleneck hampering field-scale translational success. To address this issue, three priorities are proposed: (i) designing synthetic microbial consortia, (ii) using multi-omics to examine mechanisms, and (iii) developing tools for site-specific deployment. In conclusion, these sections provide a framework for developing more reliable microbial inoculants for maize.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Density-Dependent Changes in Root Exudates, Rhizosphere Microbes, and Soil Metabolites of Solanum rostratum.
Plants (Basel, Switzerland), 15(21):.
Intraspecific competition shapes high-density monodominant communities of invasive plants, but how density is associated with rhizosphere microbial reassembly and soil metabolite accumulation via root exudates remains unclear. We established four planting densities (1, 3, 5, and 10 plants per pot) and integrated growth and photosynthetic physiology, root exudate metabolomics, rhizosphere metagenomics, and soil metabolomics using Mantel tests and variable-level correlation analysis. Increasing density reduced all traits except plant height, which showed a slight, non-significant increase followed by a significant decrease. Root exudates shifted from phenolic acid/terpenoid dominance at low density to sugar/peptide dominance at high density. Low-density-enriched bacterial genera were positively correlated with biomass, whereas high-density-enriched genera (e.g., Ilumatobacter, Allosphingosinicella) were negatively correlated. Soil metabolites such as Glutarate Semialdehyde and Sutherlandin D were negatively correlated with biomass, root traits, and photosynthesis. Mantel tests showed strong correlations between plant phenotypes and root exudates (r = 0.80) and between exudates and soil metabolites (r = 0.83). Variable-level analysis linked phenolic acid exudates to putatively growth-associated bacteria (e.g., Flavitalea, Blastococcus) and sugar/peptide exudates to putatively stress-adapted bacteria (e.g., Allosphingosinicella, Ilumatobacter), which were positively correlated with Glutarate Semialdehyde and Sutherlandin D accumulation. These results indicate that density-associated exudate changes synchronized with rhizosphere microbial reassembly and soil metabolite accumulation, forming an "exudate-microbe-metabolite" cascade. This cascade may buffer against high-density intraspecific competition and help explain monodominant community formation in invaded ranges.
Additional Links: PMID-43029048
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43029048,
year = {2026},
author = {Wu, Y and Wang, F and Liu, S and Dai, T and Deng, J and Jiang, L and Yimingniyazi, A and Wang, S},
title = {Density-Dependent Changes in Root Exudates, Rhizosphere Microbes, and Soil Metabolites of Solanum rostratum.},
journal = {Plants (Basel, Switzerland)},
volume = {15},
number = {21},
pages = {},
pmid = {43029048},
issn = {2223-7747},
support = {32460684//National Natural Science Foundation of China/ ; 2023D01B37//Natural Science Foundation of the Xinjiang Uygur Autonomous Region/ ; XKLEAEEEO-09//Xinjiang Key Laboratory for Ecological Adaptation and Evolution of Extreme Environment Organisms/ ; //Forage Germplasm Resources Survey in Xinjiang/ ; },
abstract = {Intraspecific competition shapes high-density monodominant communities of invasive plants, but how density is associated with rhizosphere microbial reassembly and soil metabolite accumulation via root exudates remains unclear. We established four planting densities (1, 3, 5, and 10 plants per pot) and integrated growth and photosynthetic physiology, root exudate metabolomics, rhizosphere metagenomics, and soil metabolomics using Mantel tests and variable-level correlation analysis. Increasing density reduced all traits except plant height, which showed a slight, non-significant increase followed by a significant decrease. Root exudates shifted from phenolic acid/terpenoid dominance at low density to sugar/peptide dominance at high density. Low-density-enriched bacterial genera were positively correlated with biomass, whereas high-density-enriched genera (e.g., Ilumatobacter, Allosphingosinicella) were negatively correlated. Soil metabolites such as Glutarate Semialdehyde and Sutherlandin D were negatively correlated with biomass, root traits, and photosynthesis. Mantel tests showed strong correlations between plant phenotypes and root exudates (r = 0.80) and between exudates and soil metabolites (r = 0.83). Variable-level analysis linked phenolic acid exudates to putatively growth-associated bacteria (e.g., Flavitalea, Blastococcus) and sugar/peptide exudates to putatively stress-adapted bacteria (e.g., Allosphingosinicella, Ilumatobacter), which were positively correlated with Glutarate Semialdehyde and Sutherlandin D accumulation. These results indicate that density-associated exudate changes synchronized with rhizosphere microbial reassembly and soil metabolite accumulation, forming an "exudate-microbe-metabolite" cascade. This cascade may buffer against high-density intraspecific competition and help explain monodominant community formation in invaded ranges.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Tuberculosis-Associated Hemophagocytic Lymphohistiocytosis with Bone Marrow Involvement: Case Report and Literature Review.
Journal of clinical medicine, 15(21):.
Tuberculosis is a rare but fatal trigger of secondary hemophagocytic lymphohistiocytosis (HLH). Its diagnosis is challenging because clinical manifestations overlap with HLH, and immunological tests may be falsely negative in HLH-related immune dysfunction. We report a case of a 34-year-old male with fever and pancytopenia. The patient's diagnostic course was protracted and misleading: all conventional tuberculosis immunological tests were negative, and he initially did not meet HLH-2004 criteria. The diagnosis of TB-HLH was eventually established by a second bone marrow biopsy and metagenomic next-generation sequencing. We also reviewed 66 TB-HLH cases, revealing frequent negative immunological test results and showing that timely anti-tuberculosis therapy was associated with reduced mortality (78.3% vs. 15.0%, p < 0.001). This case and our review underscore the importance of early bone marrow biopsy and molecular testing in unexplained HLH, particularly when tuberculosis tests are negative. Given the high mortality and frequent diagnostic delay, clinicians should maintain a high index of suspicion and pursue aggressive diagnostic workup for tuberculosis in HLH patients without a clear trigger. Furthermore, we emphasize the need to raise awareness of TB-HLH and to interpret immunological test results with caution in HLH patients, as negative findings should not be used to exclude tuberculosis.
Additional Links: PMID-43030384
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43030384,
year = {2026},
author = {You, Y and Li, H and Pei, X and Wu, D and Lv, X and Wu, Y},
title = {Tuberculosis-Associated Hemophagocytic Lymphohistiocytosis with Bone Marrow Involvement: Case Report and Literature Review.},
journal = {Journal of clinical medicine},
volume = {15},
number = {21},
pages = {},
pmid = {43030384},
issn = {2077-0383},
abstract = {Tuberculosis is a rare but fatal trigger of secondary hemophagocytic lymphohistiocytosis (HLH). Its diagnosis is challenging because clinical manifestations overlap with HLH, and immunological tests may be falsely negative in HLH-related immune dysfunction. We report a case of a 34-year-old male with fever and pancytopenia. The patient's diagnostic course was protracted and misleading: all conventional tuberculosis immunological tests were negative, and he initially did not meet HLH-2004 criteria. The diagnosis of TB-HLH was eventually established by a second bone marrow biopsy and metagenomic next-generation sequencing. We also reviewed 66 TB-HLH cases, revealing frequent negative immunological test results and showing that timely anti-tuberculosis therapy was associated with reduced mortality (78.3% vs. 15.0%, p < 0.001). This case and our review underscore the importance of early bone marrow biopsy and molecular testing in unexplained HLH, particularly when tuberculosis tests are negative. Given the high mortality and frequent diagnostic delay, clinicians should maintain a high index of suspicion and pursue aggressive diagnostic workup for tuberculosis in HLH patients without a clear trigger. Furthermore, we emphasize the need to raise awareness of TB-HLH and to interpret immunological test results with caution in HLH patients, as negative findings should not be used to exclude tuberculosis.},
}
RevDate: 2026-11-14
proMGE2-a planetary-scale resource of mobile genetic elements in microbial (meta)genomes.
Nucleic acids research pii:9051884 [Epub ahead of print].
proMGE is a resource of prokaryotic mobile genetic elements (MGEs) that systematically catalogues MGEs from prokaryotes, their intercellular spread, antimicrobial resistance gene carriage, and MGE nesting interactions. We present proMGE2 featuring a 6X expansion, a total of >16.4 million MGEs from 670k high-quality isolate genomes, and 180k metagenome-assembled genomes (MAGs), spanning diverse MGE categories: transposons and insertion sequences, phages, phage-like elements, conjugative elements, mobility islands, and integrons. proMGE2 includes MGEs from MAGs derived from environmental microbiomes, encompassing ~50k samples across several major habitat types. proMGE2 provides a standardized MGE reference collection of >5 million non-redundant MGE sequences drawn from our curated database, and enables identification of MGEs in user-provided genomes. proMGE2 captures MGE diversity and their functional potential at unprecedented scale to support future research of mobile genetic elements and transfer of genetic material in microbes. The database is available at http://promge.embl.de/.
Additional Links: PMID-43031543
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43031543,
year = {2026},
author = {Grekova, A and Schudoma, C and Letunic, I and Kim, CY and Podlesny, D and Fullam, A and Schmidt, TSB and Orakov, A and Mende, D and Schiller, J and Erbstein, KOE and Bartmanski, B and Zimmermann-Kogadeeva, M and Zimmermann, M and Kuhn, M and Bork, P and Khedkar, S},
title = {proMGE2-a planetary-scale resource of mobile genetic elements in microbial (meta)genomes.},
journal = {Nucleic acids research},
volume = {},
number = {},
pages = {},
doi = {10.1093/nar/gkag1163},
pmid = {43031543},
issn = {1362-4962},
support = {//Ministry of Science, Research and the Arts Baden-Württemberg (MWK)/ ; W-de.NBI-014//Federal Ministry of Research, Technology and Space in the frame of de.NBI & ELIXIR-DE/ ; //European Molecular Biology Laboratory/ ; 555128266//Deutsche Forschungsgemeinschaft/ ; 460129525//Deutsche Forschungsgemeinschaft/ ; //EMBL Planetary Biology Transversal Theme/ ; },
abstract = {proMGE is a resource of prokaryotic mobile genetic elements (MGEs) that systematically catalogues MGEs from prokaryotes, their intercellular spread, antimicrobial resistance gene carriage, and MGE nesting interactions. We present proMGE2 featuring a 6X expansion, a total of >16.4 million MGEs from 670k high-quality isolate genomes, and 180k metagenome-assembled genomes (MAGs), spanning diverse MGE categories: transposons and insertion sequences, phages, phage-like elements, conjugative elements, mobility islands, and integrons. proMGE2 includes MGEs from MAGs derived from environmental microbiomes, encompassing ~50k samples across several major habitat types. proMGE2 provides a standardized MGE reference collection of >5 million non-redundant MGE sequences drawn from our curated database, and enables identification of MGEs in user-provided genomes. proMGE2 captures MGE diversity and their functional potential at unprecedented scale to support future research of mobile genetic elements and transfer of genetic material in microbes. The database is available at http://promge.embl.de/.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
Variation in rhizosphere microbial communities and nitrogen- and phosphorus-cycling functional genes among rubber plantations in Yunnan, China.
Frontiers in microbiology, 17:1977925.
INTRODUCTION: Rhizosphere microorganisms are important participants in soil nutrient transformation. However, variation in rhizosphere microbial community composition and nitrogen (N)- and phosphorus (P)-cycling functional gene profiles among rubber plantations in Yunnan Province remains unclear.
METHODS: Metagenomic sequencing was used to compare rhizosphere microbial community composition and N- and P-cycling functional gene profiles among the rubber plantations located in five major natural rubber-producing areas in Yunnan Province, China: Gengma (MD), Ximeng (XM), Mengla (ML), Jinghong (JH), and Jiangcheng (JC).
RESULTS: Bacterial and fungal community composition and N- and P-cycling functional gene profiles differed significantly among the five rubber plantations. JC exhibited higher fungal diversity and richness, but its bacterial diversity was significantly lower than that in MD. The bacterial communities were dominated by Pseudomonadota, Acidobacteriota, and Actinomycetota, and the relative abundance of Acidobacteria was significantly enriched in JC compared to JH. Furthermore, Mucoromycota, Ascomycota, and Basidiomycota were the dominant phyla in the fungal community. At the functional level, compared to JC, key genes associated with N cycling were significantly enriched in ML, including N assimilation (glnA, gltB, gltD), dissimilatory nitrate reduction to ammonium (narG, narH, nirB), nitrification (nxrA, nxrB), and denitrification (napA, napB). Furthermore, the nosZ gene was also significantly enriched in MD compared to JC. Regarding the P cycle, compared to JC, genes associated with high-affinity phosphate transporters (pstA, pstB, pstC) and polyphosphate degradation (ndk) were significantly enriched in MD, whereas the organic phosphorus mineralization-related gene phnX exhibited the opposite trend. Meanwhile, compared to MD, the polyphosphate degradation-related gene ppk2 was significantly enriched in ML, whereas the inorganic phosphorus solubilization-related gene pqqC exhibited the opposite pattern. Hierarchical partitioning identified soil pH, total phosphorus (TP), and total potassium (TK) as the primary factors significantly associated with variations in bacterial community composition and P-cycling functional gene profiles.
CONCLUSION: In summary, the composition of rhizosphere bacterial and fungal communities, as well as the functional gene profiles related to N and P cycling, differed significantly among the investigated rubber plantations. These variations were closely associated with soil physicochemical factors. These findings may provide a microbiological basis for further developing site-specific nutrient management strategies in rubber plantations.
Additional Links: PMID-43031719
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43031719,
year = {2026},
author = {Geng, S and Yang, J and Shi, X and Zhang, Q and Yang, C},
title = {Variation in rhizosphere microbial communities and nitrogen- and phosphorus-cycling functional genes among rubber plantations in Yunnan, China.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1977925},
pmid = {43031719},
issn = {1664-302X},
abstract = {INTRODUCTION: Rhizosphere microorganisms are important participants in soil nutrient transformation. However, variation in rhizosphere microbial community composition and nitrogen (N)- and phosphorus (P)-cycling functional gene profiles among rubber plantations in Yunnan Province remains unclear.
METHODS: Metagenomic sequencing was used to compare rhizosphere microbial community composition and N- and P-cycling functional gene profiles among the rubber plantations located in five major natural rubber-producing areas in Yunnan Province, China: Gengma (MD), Ximeng (XM), Mengla (ML), Jinghong (JH), and Jiangcheng (JC).
RESULTS: Bacterial and fungal community composition and N- and P-cycling functional gene profiles differed significantly among the five rubber plantations. JC exhibited higher fungal diversity and richness, but its bacterial diversity was significantly lower than that in MD. The bacterial communities were dominated by Pseudomonadota, Acidobacteriota, and Actinomycetota, and the relative abundance of Acidobacteria was significantly enriched in JC compared to JH. Furthermore, Mucoromycota, Ascomycota, and Basidiomycota were the dominant phyla in the fungal community. At the functional level, compared to JC, key genes associated with N cycling were significantly enriched in ML, including N assimilation (glnA, gltB, gltD), dissimilatory nitrate reduction to ammonium (narG, narH, nirB), nitrification (nxrA, nxrB), and denitrification (napA, napB). Furthermore, the nosZ gene was also significantly enriched in MD compared to JC. Regarding the P cycle, compared to JC, genes associated with high-affinity phosphate transporters (pstA, pstB, pstC) and polyphosphate degradation (ndk) were significantly enriched in MD, whereas the organic phosphorus mineralization-related gene phnX exhibited the opposite trend. Meanwhile, compared to MD, the polyphosphate degradation-related gene ppk2 was significantly enriched in ML, whereas the inorganic phosphorus solubilization-related gene pqqC exhibited the opposite pattern. Hierarchical partitioning identified soil pH, total phosphorus (TP), and total potassium (TK) as the primary factors significantly associated with variations in bacterial community composition and P-cycling functional gene profiles.
CONCLUSION: In summary, the composition of rhizosphere bacterial and fungal communities, as well as the functional gene profiles related to N and P cycling, differed significantly among the investigated rubber plantations. These variations were closely associated with soil physicochemical factors. These findings may provide a microbiological basis for further developing site-specific nutrient management strategies in rubber plantations.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
The Temperature Sensitivity of Soil CO2 Release Is Modified by Virus-Bacteria Interactions.
Research (Washington, D.C.), 9:1479.
The temperature sensitivity (Q 10) of soil carbon (C) release is key to predicting C-climate feedback, yet omitting microbial interactions, especially with soil viruses, introduces substantial uncertainty. To address this, we performed 30- and 365-d incubations with metagenomics and metatranscriptomics profiling. We found that soil viruses markedly decreased Q 10 during 30-d incubation. Over 365-d incubation, however, their effects on microbial metabolism and Q 10 were temperature-dependent. The relative abundance of pathways related to viral replication, including those for lytic cycle initiation (e.g., propanoate metabolism decreased by 15.24%) and the energy supply (e.g., lipoic acid metabolism decreased by 20.95%), became less abundant as temperature rose from 15 to 35 °C. Metatranscriptomics data confirmed this trend, with up-regulation of metabolic functions related to viral infection and replication at 15 °C. Thus, temperature-driven variations in viral activity underlie the contrasting roles of soil viruses in regulating Q 10. This study provides the first evidence of temperature-dependent viral regulation of microbial metabolism and Q 10, highlighting the need to incorporate soil viruses into C-climate models.
Additional Links: PMID-43031837
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43031837,
year = {2026},
author = {Xu, J and Tong, D and Shi, J and Xie, J and Zhang, N and Dai, Z and Zhang, S and Dahlgren, RA and Ma, B},
title = {The Temperature Sensitivity of Soil CO2 Release Is Modified by Virus-Bacteria Interactions.},
journal = {Research (Washington, D.C.)},
volume = {9},
number = {},
pages = {1479},
pmid = {43031837},
issn = {2639-5274},
abstract = {The temperature sensitivity (Q 10) of soil carbon (C) release is key to predicting C-climate feedback, yet omitting microbial interactions, especially with soil viruses, introduces substantial uncertainty. To address this, we performed 30- and 365-d incubations with metagenomics and metatranscriptomics profiling. We found that soil viruses markedly decreased Q 10 during 30-d incubation. Over 365-d incubation, however, their effects on microbial metabolism and Q 10 were temperature-dependent. The relative abundance of pathways related to viral replication, including those for lytic cycle initiation (e.g., propanoate metabolism decreased by 15.24%) and the energy supply (e.g., lipoic acid metabolism decreased by 20.95%), became less abundant as temperature rose from 15 to 35 °C. Metatranscriptomics data confirmed this trend, with up-regulation of metabolic functions related to viral infection and replication at 15 °C. Thus, temperature-driven variations in viral activity underlie the contrasting roles of soil viruses in regulating Q 10. This study provides the first evidence of temperature-dependent viral regulation of microbial metabolism and Q 10, highlighting the need to incorporate soil viruses into C-climate models.},
}
RevDate: 2026-11-14
CmpDate: 2026-11-14
A fatal threat: Pneumocystis jirovecii pneumonia with rapidly progressive interstitial lung disease in anti-MDA5-positive dermatomyositis-a case report.
Frontiers in medicine, 13:1977161.
Pneumocystis jirovecii pneumonia (PJP) is a respiratory infection caused by Pneumocystis organisms, representing an opportunistic fungal infection. The clinical occurrence of PJP combined with rapidly progressive interstitial lung disease (RP-ILD) in dermatomyositis (DM) have also been reported occasionally, but patients with this combination of conditions experience rapid disease progression and high mortality rates. We reported a 70-year-old female patient with dyspnea and fever, detailing her laboratory tests, computed tomography (CT) findings and bronchoalveolar lavage fluid findings. The patient's main clinical manifestations were dyspnea and fever. CT scans revealed scattered reticular opacities and ground-glass opacities in both lungs and metagenomic next-generation sequencing (mNGS) of the bronchoalveolar lavage fluid detected Pneumocystis jirovecii. Laboratory tests revealed positive anti-MDA5 antibodies during hospitalization. The patient was treated with high-flow oxygen therapy and methylprednisolone upon admission. Subsequently, the patient received mechanical ventilator support and was administered trimethoprim-sulfamethoxazole (TMP-SMX), plasma exchanges and immunoglobulin treatment. The patient's fever subsided, but respiratory failure and interstitial lung disease continued to progressively worsen. Ground-glass opacities, mixed ground-glass opacities, consolidations and reticular opacities were significantly more pronounced than on admission, indicating RP-ILD changes post-treatment. The patient succumbed after 13 days of treatment. PJP complicated by rapidly progressive interstitial lung disease can be fatal in patients, emphasizing the critical importance of early identification of PJP and prompt initiation of anti-PJP treatment. It is crucial to recognize that some MDA5+ DM patients may not present with the typical rash, and in certain cases, RP-ILD may be the primary manifestation. Therefore, early screening for anti-MDA5 antibodies is essential for patients presenting with unexplained early-stage RP-ILD.
Additional Links: PMID-43031840
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43031840,
year = {2026},
author = {Li, X and Xu, S and Feng, Y and Wang, R and Wang, X},
title = {A fatal threat: Pneumocystis jirovecii pneumonia with rapidly progressive interstitial lung disease in anti-MDA5-positive dermatomyositis-a case report.},
journal = {Frontiers in medicine},
volume = {13},
number = {},
pages = {1977161},
pmid = {43031840},
issn = {2296-858X},
abstract = {Pneumocystis jirovecii pneumonia (PJP) is a respiratory infection caused by Pneumocystis organisms, representing an opportunistic fungal infection. The clinical occurrence of PJP combined with rapidly progressive interstitial lung disease (RP-ILD) in dermatomyositis (DM) have also been reported occasionally, but patients with this combination of conditions experience rapid disease progression and high mortality rates. We reported a 70-year-old female patient with dyspnea and fever, detailing her laboratory tests, computed tomography (CT) findings and bronchoalveolar lavage fluid findings. The patient's main clinical manifestations were dyspnea and fever. CT scans revealed scattered reticular opacities and ground-glass opacities in both lungs and metagenomic next-generation sequencing (mNGS) of the bronchoalveolar lavage fluid detected Pneumocystis jirovecii. Laboratory tests revealed positive anti-MDA5 antibodies during hospitalization. The patient was treated with high-flow oxygen therapy and methylprednisolone upon admission. Subsequently, the patient received mechanical ventilator support and was administered trimethoprim-sulfamethoxazole (TMP-SMX), plasma exchanges and immunoglobulin treatment. The patient's fever subsided, but respiratory failure and interstitial lung disease continued to progressively worsen. Ground-glass opacities, mixed ground-glass opacities, consolidations and reticular opacities were significantly more pronounced than on admission, indicating RP-ILD changes post-treatment. The patient succumbed after 13 days of treatment. PJP complicated by rapidly progressive interstitial lung disease can be fatal in patients, emphasizing the critical importance of early identification of PJP and prompt initiation of anti-PJP treatment. It is crucial to recognize that some MDA5+ DM patients may not present with the typical rash, and in certain cases, RP-ILD may be the primary manifestation. Therefore, early screening for anti-MDA5 antibodies is essential for patients presenting with unexplained early-stage RP-ILD.},
}
RevDate: 2026-05-18
CmpDate: 2026-05-18
The Gut Feeling Behind Autoimmune Encephalitis.
Epilepsy currents, 26(3):231-233.
OBJECTIVE: Autoimmune encephalitis is a cause of brain inflammation characterized by auto-antibodies, which target cell surface neuronal proteins and lead to neuronal dysfunction. The most common form is associated with auto-antibodies to leucine-rich glioma-inactivated 1 (LGI1) protein, the presentation of which includes frequent focal seizures. The exact cause of these auto-antibodies remains unknown, but established predispositions include overrepresented human leukocyte antigen (HLA) alleles. Yet, these HLA alleles are themselves common in the healthy ancestry-matched population. One potential etiological hypothesis is that an environmental trigger, such as the gut microbiome, interacts with a genetically predisposed individual.
METHODS: To investigate this, we studied 42 patients with LGI1-antibody encephalitis (LGI1-Ab-E) and 27 familial/environmentally matched controls and performed metagenomic shotgun sequencing, to describe the compositional and functional differences in the gut microbiome.
RESULTS: We observed that LGI1-Ab-E gut microbiomes exhibited a significant reduction in the ratio of Firmicutes (or Bacillota) and Bacteroidetes phyla, which is associated with the dosage of HLA susceptibility allele count in patients with LGI1-Ab-E. Furthermore, we identified differences in functional gene profiles in the gut microbiome that led to a reduction of neuroinflammatory protective short-chain fatty acids (SCFAs) in LGI1-Ab-E patients.
SIGNIFICANCE: Taken together, our results suggest that a compositional shift in the gut microbiome of LGI1-Ab-E associates with a neuroinflammatory state, possibly through the reduction of SCFA production. Our study highlights the potential of the gut microbiome to explain some of the complex condition and unravel etiological questions. Validation studies with greater sample sizes are recommended.
Additional Links: PMID-41509546
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid41509546,
year = {2026},
author = {Steriade, C},
title = {The Gut Feeling Behind Autoimmune Encephalitis.},
journal = {Epilepsy currents},
volume = {26},
number = {3},
pages = {231-233},
pmid = {41509546},
issn = {1535-7597},
abstract = {OBJECTIVE: Autoimmune encephalitis is a cause of brain inflammation characterized by auto-antibodies, which target cell surface neuronal proteins and lead to neuronal dysfunction. The most common form is associated with auto-antibodies to leucine-rich glioma-inactivated 1 (LGI1) protein, the presentation of which includes frequent focal seizures. The exact cause of these auto-antibodies remains unknown, but established predispositions include overrepresented human leukocyte antigen (HLA) alleles. Yet, these HLA alleles are themselves common in the healthy ancestry-matched population. One potential etiological hypothesis is that an environmental trigger, such as the gut microbiome, interacts with a genetically predisposed individual.
METHODS: To investigate this, we studied 42 patients with LGI1-antibody encephalitis (LGI1-Ab-E) and 27 familial/environmentally matched controls and performed metagenomic shotgun sequencing, to describe the compositional and functional differences in the gut microbiome.
RESULTS: We observed that LGI1-Ab-E gut microbiomes exhibited a significant reduction in the ratio of Firmicutes (or Bacillota) and Bacteroidetes phyla, which is associated with the dosage of HLA susceptibility allele count in patients with LGI1-Ab-E. Furthermore, we identified differences in functional gene profiles in the gut microbiome that led to a reduction of neuroinflammatory protective short-chain fatty acids (SCFAs) in LGI1-Ab-E patients.
SIGNIFICANCE: Taken together, our results suggest that a compositional shift in the gut microbiome of LGI1-Ab-E associates with a neuroinflammatory state, possibly through the reduction of SCFA production. Our study highlights the potential of the gut microbiome to explain some of the complex condition and unravel etiological questions. Validation studies with greater sample sizes are recommended.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Metagenomic next-generation sequencing improves pathogen detection in culture-negative infections: a diagnostic performance study.
Frontiers in microbiology, 17:1971380.
The conventional microbiological diagnosis is frequently insensitive, time consuming and targeted. Patients with underlying immune suppression and those who have recently been on antibiotics often have culture-negative infections. In recent years, metagenomic next-generation sequencing (mNGS) has emerged as a novel, pathogen-agnostic diagnostic approach. A total of 136 clinical specimens were submitted for metagenomic next-generation sequencing (mNGS) between August 2022 and January 2024. Of these samples, 86 from 73 patients were included in this retrospective study. Thirteen patients had more than one specimen included in the study. All specimens were processed using conventional microbiological methods and analyzed using mNGS technology, either by using Illumina next-generation sequencing technology or Oxford Nanopore long-read technology. The diagnostic performance characteristics, including sensitivity, specificity, positive predictive value (PPV), and negative predictive value (NPV), were calculated for both methods. The agreement between the two methods was evaluated using Cohen's kappa coefficient. The paired comparisons were made using the McNemar test. The areas under the ROC curves for both methods were calculated, and the multivariable logistic regression analysis was used to identify the variables that were independently associated with mNGS positivity. Here, conventional microbiology identified pathogens in 16.3% of samples, whereas mNGS identified pathogens in 45.3% of samples. The sensitivity and specificity of mNGS were 57.1 and 84.2%, respectively. By McNemar testing, there was significant detection of pathogens by mNGS (p = 0.0015). The agreement between two methods was poor (kappa = 0.20). The ROC curve for mNGS results showed moderate diagnostic accuracy with an AUC of 0.71. By multivariable logistic regression, immunocompromised status (OR 2.8, 95% CI 1.3-6.0, p = 0.018) and positive imaging findings (OR 3.1, 95% CI 1.4-6.9, p = 0.010) were independently associated with mNGS positivity. mNGS is able to identify more pathogens in culture-negative cases than conventional methods. There is scope for mNGS to be used in a complementary fashion, whilst relying on conventional methods as the primary tool for diagnosis of infections. Notably, the greatest additional information will come from the use of mNGS in immunocompromised patients.
Additional Links: PMID-43017131
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43017131,
year = {2026},
author = {Tayeb, HT and Abouelhoda, M and Alqasabi, A and Magrashi, A and Nafe, G and Alzahrani, T and Alzeer, S and Bakheet, R},
title = {Metagenomic next-generation sequencing improves pathogen detection in culture-negative infections: a diagnostic performance study.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1971380},
pmid = {43017131},
issn = {1664-302X},
abstract = {The conventional microbiological diagnosis is frequently insensitive, time consuming and targeted. Patients with underlying immune suppression and those who have recently been on antibiotics often have culture-negative infections. In recent years, metagenomic next-generation sequencing (mNGS) has emerged as a novel, pathogen-agnostic diagnostic approach. A total of 136 clinical specimens were submitted for metagenomic next-generation sequencing (mNGS) between August 2022 and January 2024. Of these samples, 86 from 73 patients were included in this retrospective study. Thirteen patients had more than one specimen included in the study. All specimens were processed using conventional microbiological methods and analyzed using mNGS technology, either by using Illumina next-generation sequencing technology or Oxford Nanopore long-read technology. The diagnostic performance characteristics, including sensitivity, specificity, positive predictive value (PPV), and negative predictive value (NPV), were calculated for both methods. The agreement between the two methods was evaluated using Cohen's kappa coefficient. The paired comparisons were made using the McNemar test. The areas under the ROC curves for both methods were calculated, and the multivariable logistic regression analysis was used to identify the variables that were independently associated with mNGS positivity. Here, conventional microbiology identified pathogens in 16.3% of samples, whereas mNGS identified pathogens in 45.3% of samples. The sensitivity and specificity of mNGS were 57.1 and 84.2%, respectively. By McNemar testing, there was significant detection of pathogens by mNGS (p = 0.0015). The agreement between two methods was poor (kappa = 0.20). The ROC curve for mNGS results showed moderate diagnostic accuracy with an AUC of 0.71. By multivariable logistic regression, immunocompromised status (OR 2.8, 95% CI 1.3-6.0, p = 0.018) and positive imaging findings (OR 3.1, 95% CI 1.4-6.9, p = 0.010) were independently associated with mNGS positivity. mNGS is able to identify more pathogens in culture-negative cases than conventional methods. There is scope for mNGS to be used in a complementary fashion, whilst relying on conventional methods as the primary tool for diagnosis of infections. Notably, the greatest additional information will come from the use of mNGS in immunocompromised patients.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Integrated gut microbiome and fecal metabolome profiling identifies taurine-associated microbial dysbiosis across the Alzheimer's disease clinical spectrum.
Frontiers in aging neuroscience, 18:1981076.
BACKGROUND: Gut microbiome alterations have been associated with Alzheimer's disease (AD), but how microbial composition, fecal metabolites, and cognitive function relate across the cognitive impairment spectrum-from subjective cognitive decline (SCD) through mild cognitive impairment (MCI) to AD-remains unclear. We integrated microbiome and metabolome profiles across this clinical spectrum.
METHODS: Fecal samples from 84 participants [10 healthy controls (HC), 22 SCD, 23 MCI, 29 AD] underwent 16S rRNA V3-V4 sequencing and untargeted LC-MS metabolomics. Profiles were integrated via Procrustes analysis, sparse canonical correlation, MIMOSA2 community-metabolism modeling, and mediation analysis. Differential abundance was assessed by Kruskal-Wallis, ANCOM-BC2, and LinDA.
RESULTS: Microbial composition differed across groups (ANOSIM R = 0.111, p = 0.001; Adonis R [2] = 0.070, p = 0.002). Cross-method validation confirmed 8 differentially abundant genera in AD vs. HC with 100% directional concordance between ANCOM-BC2 and LinDA, including depletion of butyrate-producing Lachnospiraceae (Blautia, Dorea, Fusicatenibacter) and enrichment of opportunistic pathobionts (Klebsiella, Escherichia-Shigella). Taurine showed the highest MIMOSA2 community-prediction R [2] (0.129; n = 62) and was positively associated with MMSE scores (r = 0.45, BH-FDR Q = 0.0005). A multi-omic random forest model (5 genera + 5 metabolites + age + sex) discriminated AD-clinical spectrum participants from HC with AUC = 0.86 (95% CI 0.82-0.90), although pairwise differences from the age + sex baseline (AUC = 0.75) did not reach statistical significance. Statistical mediation analysis was consistent with a Blautia → taurine → MMSE statistical mediation [ACME = 0.21, p = 0.008; 54% proportion mediated, bootstrap 95% CI (9%, 58%)].
CONCLUSION: Integrated multi-omic profiling reveals coordinated microbiome-metabolome dysregulation across the AD clinical spectrum, with taurine as a candidate microbial-metabolic node linking gut dysbiosis to cognitive performance. Microbial and metabolic alterations are detectable at the SCD stage, before objective cognitive impairment. These findings warrant validation in larger, age-matched cohorts and higher-resolution metagenomic studies.
Additional Links: PMID-43017153
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43017153,
year = {2026},
author = {Feng, Y and Sha, Y and Xu, L and Tian, J and Gao, Y and Zhou, R and Liu, G},
title = {Integrated gut microbiome and fecal metabolome profiling identifies taurine-associated microbial dysbiosis across the Alzheimer's disease clinical spectrum.},
journal = {Frontiers in aging neuroscience},
volume = {18},
number = {},
pages = {1981076},
pmid = {43017153},
issn = {1663-4365},
abstract = {BACKGROUND: Gut microbiome alterations have been associated with Alzheimer's disease (AD), but how microbial composition, fecal metabolites, and cognitive function relate across the cognitive impairment spectrum-from subjective cognitive decline (SCD) through mild cognitive impairment (MCI) to AD-remains unclear. We integrated microbiome and metabolome profiles across this clinical spectrum.
METHODS: Fecal samples from 84 participants [10 healthy controls (HC), 22 SCD, 23 MCI, 29 AD] underwent 16S rRNA V3-V4 sequencing and untargeted LC-MS metabolomics. Profiles were integrated via Procrustes analysis, sparse canonical correlation, MIMOSA2 community-metabolism modeling, and mediation analysis. Differential abundance was assessed by Kruskal-Wallis, ANCOM-BC2, and LinDA.
RESULTS: Microbial composition differed across groups (ANOSIM R = 0.111, p = 0.001; Adonis R [2] = 0.070, p = 0.002). Cross-method validation confirmed 8 differentially abundant genera in AD vs. HC with 100% directional concordance between ANCOM-BC2 and LinDA, including depletion of butyrate-producing Lachnospiraceae (Blautia, Dorea, Fusicatenibacter) and enrichment of opportunistic pathobionts (Klebsiella, Escherichia-Shigella). Taurine showed the highest MIMOSA2 community-prediction R [2] (0.129; n = 62) and was positively associated with MMSE scores (r = 0.45, BH-FDR Q = 0.0005). A multi-omic random forest model (5 genera + 5 metabolites + age + sex) discriminated AD-clinical spectrum participants from HC with AUC = 0.86 (95% CI 0.82-0.90), although pairwise differences from the age + sex baseline (AUC = 0.75) did not reach statistical significance. Statistical mediation analysis was consistent with a Blautia → taurine → MMSE statistical mediation [ACME = 0.21, p = 0.008; 54% proportion mediated, bootstrap 95% CI (9%, 58%)].
CONCLUSION: Integrated multi-omic profiling reveals coordinated microbiome-metabolome dysregulation across the AD clinical spectrum, with taurine as a candidate microbial-metabolic node linking gut dysbiosis to cognitive performance. Microbial and metabolic alterations are detectable at the SCD stage, before objective cognitive impairment. These findings warrant validation in larger, age-matched cohorts and higher-resolution metagenomic studies.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Improving Talaromycosis Infection Diagnosis and Prognosis with Metagenomic Next-Generation Sequencing.
Infection and drug resistance, 19:606899.
BACKGROUND: Early diagnosis and timely initiation of appropriate antifungal treatment are critical for improving the prognosis of talaromycosis infection. We aimed to test whether metagenomic next-generation sequencing (mNGS), versus conventional microbiological testing (CMT), improves talaromycosis diagnosis and management in HIV-negative patients via higher sensitivity and faster detection.
METHODS: We retrospectively investigated the clinical data of patients with Talaromyces marneffei (T. marneffei) infection in six hospitals in Fujian, China, between July 2015 and June 2024. The diagnostic performance and clinical impact of mNGS were compared with those of CMT.
RESULTS: Seventy-four patients were included, with 38 cases in the mNGS group and 36 cases for CMT group, based on the method that established the definitive diagnosis. The median duration from hospitalization to confirmed diagnosis was significantly shorter in the mNGS group than in the CMT group (5.0 vs 12.0 days, p < 0.0001). The turnround time for mNGS was shorter than that of CMT (1.0 days vs 6.5 days, p < 0.0001). The mNGS detected more mycobacteria and other fungal infection co-infections than CMT (36.8% vs 13.9%, p = 0.033; 26.3% vs 5.56%, p = 0.025). The mNGS group has a trend of lower mortality compared with the CMT group (18.4% vs 38.9%, p = 0.072). In addition, the length of stay was significantly shorter in the mNGS group than in the CMT group (14.5 days vs 21.0 days, p = 0.007).
CONCLUSION: mNGS facilitates rapid diagnosis and improves administration of targeted antimicrobial therapy, thus improving the prognosis of T. marneffei infection. The clinical utility of mNGS for talaromycosis in at-risk populations should be investigated in prospective and large studies.
Additional Links: PMID-43017160
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43017160,
year = {2026},
author = {Zhang, L and He, X and Xu, L and Xiao, J and Lin, J and Song, B and Chen, L and Chen, G and Liu, K},
title = {Improving Talaromycosis Infection Diagnosis and Prognosis with Metagenomic Next-Generation Sequencing.},
journal = {Infection and drug resistance},
volume = {19},
number = {},
pages = {606899},
pmid = {43017160},
issn = {1178-6973},
abstract = {BACKGROUND: Early diagnosis and timely initiation of appropriate antifungal treatment are critical for improving the prognosis of talaromycosis infection. We aimed to test whether metagenomic next-generation sequencing (mNGS), versus conventional microbiological testing (CMT), improves talaromycosis diagnosis and management in HIV-negative patients via higher sensitivity and faster detection.
METHODS: We retrospectively investigated the clinical data of patients with Talaromyces marneffei (T. marneffei) infection in six hospitals in Fujian, China, between July 2015 and June 2024. The diagnostic performance and clinical impact of mNGS were compared with those of CMT.
RESULTS: Seventy-four patients were included, with 38 cases in the mNGS group and 36 cases for CMT group, based on the method that established the definitive diagnosis. The median duration from hospitalization to confirmed diagnosis was significantly shorter in the mNGS group than in the CMT group (5.0 vs 12.0 days, p < 0.0001). The turnround time for mNGS was shorter than that of CMT (1.0 days vs 6.5 days, p < 0.0001). The mNGS detected more mycobacteria and other fungal infection co-infections than CMT (36.8% vs 13.9%, p = 0.033; 26.3% vs 5.56%, p = 0.025). The mNGS group has a trend of lower mortality compared with the CMT group (18.4% vs 38.9%, p = 0.072). In addition, the length of stay was significantly shorter in the mNGS group than in the CMT group (14.5 days vs 21.0 days, p = 0.007).
CONCLUSION: mNGS facilitates rapid diagnosis and improves administration of targeted antimicrobial therapy, thus improving the prognosis of T. marneffei infection. The clinical utility of mNGS for talaromycosis in at-risk populations should be investigated in prospective and large studies.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Effects of different concentrations of exogenous methyl jasmonate on tuber metabolism and rhizosphere bacterial community dynamics in Polygonatum cyrtonema Hua.
Frontiers in plant science, 17:1950448.
Polygonatum cyrtonema Hua is a traditional medicinal and edible plant valued for its tubers, which are rich in bioactive components such as polysaccharides, flavonoids, and saponins. The phytohormone methyl jasmonate (MeJA) regulates plant secondary metabolism and stress responses, promoting the accumulation of medicinal compounds in P. cyrtonema. However, the temporal dynamics of the rhizosphere microbiome and its association with the tuber metabolism under different MeJA concentrations remain poorly understood. In this study, three-year-old P. cyrtonema plants were treated with rhizosphere sprays of 200 μmol L[-1] and 400 μmol L[-1] MeJA. Plants were sampled at 7, 14, 21, and 28 days post-treatment to evaluate tuber physiology and metabolite profiles, and metagenomic sequencing was used to characterize the structure and function of the rhizosphere bacterial community. Application of 200 μmol L[-1] MeJA significantly enhanced tuber peroxidase (POD) activity, which peaked at 2.29 times that of the control on day 14 (P < 0.05), and increased total flavonoid and saponin contents by 22.58% and 13.85%, respectively (P < 0.05). In contrast, 400 μmol L[-1] MeJA mainly inhibited antioxidant enzyme activities and delayed the induction of secondary metabolism. Metagenomic analysis revealed that the two MeJA concentrations shared the same dominant bacterial phyla but showed concentration-dependent differences in relative abundance dynamics, alpha diversity trajectories, and temporal patterns of community structure. The 200 μmol L[-1] MeJA group exhibited a directional temporal shift (PERMANOVA R[2] = 0.79, P = 0.003) and continuously rising alpha diversity, whereas the 400 μmol L[-1] treatment showed nonspecific stress fluctuations, with reduced alpha diversity, a weaker temporal gradient in community structure, and increased community homogenization. At the functional level, the 200 μmol L[-1] MeJA group had higher relative abundances of microbial genes associated with nitrogen, carbon, and secondary metabolic pathways. In contrast, the 400 μmol L[-1] MeJA group showed disrupted coordination among functional genes and persistently higher abundances of genes involved in antioxidant pathways. Correlation analysis identified associations between specific phyla and host traits: Nitrospirota was positively correlated with POD activity (r = 0.48, P < 0.05); Methylomirabilota was positively correlated with total flavonoid and saponin contents (r ≥ 0.41, P < 0.05); and Verrucomicrobiota was negatively correlated with these components (r < -0.55, P < 0.01). These findings suggest that 200 μmol L[-1] MeJA promotes the accumulation of bioactive metabolites in P. cyrtonema through two proposed ways: the direct activation of host metabolism and the indirect association with shifts in potentially beneficial rhizosphere microbial taxa. This study provides a practical foundation for selective MeJA concentrations in medicinal plant cultivation.
Additional Links: PMID-43017163
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43017163,
year = {2026},
author = {Liu, M and Yan, L and Lai, J and Xu, X and Cheng, S},
title = {Effects of different concentrations of exogenous methyl jasmonate on tuber metabolism and rhizosphere bacterial community dynamics in Polygonatum cyrtonema Hua.},
journal = {Frontiers in plant science},
volume = {17},
number = {},
pages = {1950448},
pmid = {43017163},
issn = {1664-462X},
abstract = {Polygonatum cyrtonema Hua is a traditional medicinal and edible plant valued for its tubers, which are rich in bioactive components such as polysaccharides, flavonoids, and saponins. The phytohormone methyl jasmonate (MeJA) regulates plant secondary metabolism and stress responses, promoting the accumulation of medicinal compounds in P. cyrtonema. However, the temporal dynamics of the rhizosphere microbiome and its association with the tuber metabolism under different MeJA concentrations remain poorly understood. In this study, three-year-old P. cyrtonema plants were treated with rhizosphere sprays of 200 μmol L[-1] and 400 μmol L[-1] MeJA. Plants were sampled at 7, 14, 21, and 28 days post-treatment to evaluate tuber physiology and metabolite profiles, and metagenomic sequencing was used to characterize the structure and function of the rhizosphere bacterial community. Application of 200 μmol L[-1] MeJA significantly enhanced tuber peroxidase (POD) activity, which peaked at 2.29 times that of the control on day 14 (P < 0.05), and increased total flavonoid and saponin contents by 22.58% and 13.85%, respectively (P < 0.05). In contrast, 400 μmol L[-1] MeJA mainly inhibited antioxidant enzyme activities and delayed the induction of secondary metabolism. Metagenomic analysis revealed that the two MeJA concentrations shared the same dominant bacterial phyla but showed concentration-dependent differences in relative abundance dynamics, alpha diversity trajectories, and temporal patterns of community structure. The 200 μmol L[-1] MeJA group exhibited a directional temporal shift (PERMANOVA R[2] = 0.79, P = 0.003) and continuously rising alpha diversity, whereas the 400 μmol L[-1] treatment showed nonspecific stress fluctuations, with reduced alpha diversity, a weaker temporal gradient in community structure, and increased community homogenization. At the functional level, the 200 μmol L[-1] MeJA group had higher relative abundances of microbial genes associated with nitrogen, carbon, and secondary metabolic pathways. In contrast, the 400 μmol L[-1] MeJA group showed disrupted coordination among functional genes and persistently higher abundances of genes involved in antioxidant pathways. Correlation analysis identified associations between specific phyla and host traits: Nitrospirota was positively correlated with POD activity (r = 0.48, P < 0.05); Methylomirabilota was positively correlated with total flavonoid and saponin contents (r ≥ 0.41, P < 0.05); and Verrucomicrobiota was negatively correlated with these components (r < -0.55, P < 0.01). These findings suggest that 200 μmol L[-1] MeJA promotes the accumulation of bioactive metabolites in P. cyrtonema through two proposed ways: the direct activation of host metabolism and the indirect association with shifts in potentially beneficial rhizosphere microbial taxa. This study provides a practical foundation for selective MeJA concentrations in medicinal plant cultivation.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Microbiome-targeted strategies for Alzheimer's disease: multi-omics signatures, mechanistic links, and intervention evidence.
Frontiers in microbiology, 17:1828118.
Alzheimer's disease (AD) is increasingly studied in relation to gut microbial mechanisms because microbial metabolism, mucosal barrier function, and peripheral immune activation may influence neuroinflammation and host metabolic stress. This review evaluates microbiome-linked evidence in AD with emphasis on studies that combine metagenomics, metabolomics, inflammatory markers, neuroimaging, cerebrospinal fluid or blood biomarkers, and cognitive phenotyping. Current data support several recurring but not yet causal axes, including bile acid transformation, short-chain fatty acid (SCFA) production, trimethylamine N-oxide (TMAO), bacterial product-related inflammatory signaling, and barrier dysfunction. We also review microbiome-targeted interventions, including diet and prebiotics, strain-specified probiotics and synbiotics, oligosaccharide-based agents, gut-restricted antimicrobials, and fecal microbiota transplantation (FMT). The strength of evidence differs substantially across these strategies: sodium oligomannate has been tested in a phase 3 trial, whereas many probiotic, antimicrobial, and FMT approaches remain early clinical, preclinical, or mechanism-generating. Future trials should define a priori microbial, metabolite, immune, and brain biomarker endpoints; control diet, medication, and comorbidity confounding; and validate candidate signatures across independent cohorts before using microbiome profiles for screening or treatment selection.
Additional Links: PMID-43017556
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43017556,
year = {2026},
author = {Huang, X and Zhang, X and Zhu, Q},
title = {Microbiome-targeted strategies for Alzheimer's disease: multi-omics signatures, mechanistic links, and intervention evidence.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1828118},
pmid = {43017556},
issn = {1664-302X},
abstract = {Alzheimer's disease (AD) is increasingly studied in relation to gut microbial mechanisms because microbial metabolism, mucosal barrier function, and peripheral immune activation may influence neuroinflammation and host metabolic stress. This review evaluates microbiome-linked evidence in AD with emphasis on studies that combine metagenomics, metabolomics, inflammatory markers, neuroimaging, cerebrospinal fluid or blood biomarkers, and cognitive phenotyping. Current data support several recurring but not yet causal axes, including bile acid transformation, short-chain fatty acid (SCFA) production, trimethylamine N-oxide (TMAO), bacterial product-related inflammatory signaling, and barrier dysfunction. We also review microbiome-targeted interventions, including diet and prebiotics, strain-specified probiotics and synbiotics, oligosaccharide-based agents, gut-restricted antimicrobials, and fecal microbiota transplantation (FMT). The strength of evidence differs substantially across these strategies: sodium oligomannate has been tested in a phase 3 trial, whereas many probiotic, antimicrobial, and FMT approaches remain early clinical, preclinical, or mechanism-generating. Future trials should define a priori microbial, metabolite, immune, and brain biomarker endpoints; control diet, medication, and comorbidity confounding; and validate candidate signatures across independent cohorts before using microbiome profiles for screening or treatment selection.},
}
RevDate: 2026-11-12
Near-complete genome of Tetraparvovirus sp. specimen WTD-TP1-22 identified in white-tailed deer (Odocoileus virginianus).
Microbiology resource announcements [Epub ahead of print].
Tetraparvovirus sp. specimen WTD-TP1-22 was recovered from a wild white-tailed deer (Odocoileus virginianus) nasal swab in Pennsylvania. The near-complete genome is 4,980 bp with a guanine-cytosine content of 47.9%, and this virus shares 95.91% nucleotide identity with isolate tetra-CA1 (GenBank NC_031670.1) from mule deer.
Additional Links: PMID-43018149
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43018149,
year = {2026},
author = {Cluett, H and Chandler, JC and Bisha, B},
title = {Near-complete genome of Tetraparvovirus sp. specimen WTD-TP1-22 identified in white-tailed deer (Odocoileus virginianus).},
journal = {Microbiology resource announcements},
volume = {},
number = {},
pages = {e0078226},
doi = {10.1128/mra.00782-26},
pmid = {43018149},
issn = {2576-098X},
abstract = {Tetraparvovirus sp. specimen WTD-TP1-22 was recovered from a wild white-tailed deer (Odocoileus virginianus) nasal swab in Pennsylvania. The near-complete genome is 4,980 bp with a guanine-cytosine content of 47.9%, and this virus shares 95.91% nucleotide identity with isolate tetra-CA1 (GenBank NC_031670.1) from mule deer.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-12
ArchaeaHQ: a curated reference database of archaeal genomes.
Microbial genomics, 12(11):.
Archaea have proven to be major players in biogeochemical cycles across diverse ecosystems, yet we still see an underrepresentation of archaeal genomes in the datasets used by popular computational biology tools. Here, we present ArchaeaHQ, a quality-controlled, systematically curated reference database of 25,584 archaeal genomes compiled initially from 43,045 assemblies from all four archaeal kingdoms retrieved from NCBI: Methanobacteriati (Euryarchaeota), Thermoproteati (TACK), Nanobdellati (DPANN) and Promethearchaeati (Asgard). All genomes in the database passed standardized quality control, requiring ≥70% completeness and ≤10% contamination. A total of 44.0% of genomes in ArchaeaHQ achieved ≥90% completeness, while 92.7% exhibited ≤5% contamination. ArchaeaHQ comprises 18,701 metagenome-assembled genomes (MAGs; 73.1%) and 6,883 isolate genomes (26.9%). Approximately 76% of MAGs are assigned to 17 ecologically meaningful categories based on sampling origin, and around 65% of genomes include geographic metadata. ArchaeaHQ is available at https://doi.org/10.6084/m9.figshare.32266599 for download and at https://vee-lab.eu/archaeahq for online browsing. This resource was designed to provide an analysis-ready reference set for metagenomic, biogeochemical and ecological studies, comparative genomics analyses and the development of archaeal-specific bioinformatic tools.
Additional Links: PMID-43018641
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43018641,
year = {2026},
author = {Bespiatykh, D and Leão, P},
title = {ArchaeaHQ: a curated reference database of archaeal genomes.},
journal = {Microbial genomics},
volume = {12},
number = {11},
pages = {},
pmid = {43018641},
issn = {2057-5858},
mesh = {*Genome, Archaeal ; *Archaea/genetics/classification ; *Databases, Genetic ; Metagenome ; Metagenomics ; Biocuration ; Computational Biology/methods ; Phylogeny ; },
abstract = {Archaea have proven to be major players in biogeochemical cycles across diverse ecosystems, yet we still see an underrepresentation of archaeal genomes in the datasets used by popular computational biology tools. Here, we present ArchaeaHQ, a quality-controlled, systematically curated reference database of 25,584 archaeal genomes compiled initially from 43,045 assemblies from all four archaeal kingdoms retrieved from NCBI: Methanobacteriati (Euryarchaeota), Thermoproteati (TACK), Nanobdellati (DPANN) and Promethearchaeati (Asgard). All genomes in the database passed standardized quality control, requiring ≥70% completeness and ≤10% contamination. A total of 44.0% of genomes in ArchaeaHQ achieved ≥90% completeness, while 92.7% exhibited ≤5% contamination. ArchaeaHQ comprises 18,701 metagenome-assembled genomes (MAGs; 73.1%) and 6,883 isolate genomes (26.9%). Approximately 76% of MAGs are assigned to 17 ecologically meaningful categories based on sampling origin, and around 65% of genomes include geographic metadata. ArchaeaHQ is available at https://doi.org/10.6084/m9.figshare.32266599 for download and at https://vee-lab.eu/archaeahq for online browsing. This resource was designed to provide an analysis-ready reference set for metagenomic, biogeochemical and ecological studies, comparative genomics analyses and the development of archaeal-specific bioinformatic tools.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Genome, Archaeal
*Archaea/genetics/classification
*Databases, Genetic
Metagenome
Metagenomics
Biocuration
Computational Biology/methods
Phylogeny
RevDate: 2026-11-12
Immunologically optimized Zmp1 peptides reveal a translational serological biomarker platform for tuberculosis diagnosis across disease manifestations.
European journal of clinical microbiology & infectious diseases : official publication of the European Society of Clinical Microbiology [Epub ahead of print].
Tuberculosis (TB) diagnosis remains challenging, particularly for extrapulmonary TB (EPTB), where invasive sampling, low bacillary burden, and limited sensitivity of nucleic acid-based tests in peripheral specimens hinder timely detection. Here, we report an immunology-driven strategy for biomarker discovery and development of a peptide-based serological assay targeting Mycobacterium tuberculosis zinc metalloprotease-1 (Zmp1). Based on the principle that antigenic regions containing overlapping B-cell and CD4[+]T-cell epitopes can promote strong antibody responses through linked recognition and cognate T-cell help, we used an immunoinformatics pipeline to identify two nested immunodominant Zmp1 peptide regions, Mtb-Zp-NT and Mtb-Zp-CT, enriched for overlapping B- and T-cell epitopes.Their diagnostic potential was evaluated using ELISA-based serological assays. A blinded pilot study (N = 137) demonstrated clear discrimination between active TB and TB-recovered individuals. An expanded validation cohort (N = 875), identified from screening approximately 6,086 individuals, included pulmonary TB (PTB), EPTB, TB-recovered individuals, household contacts, non-specific infection/inflammation, and healthy controls. Receiver operating characteristic analyses, supported by DeLong and bootstrap comparisons, demonstrated superior diagnostic performance of the peptide assays compared with Mtb-FL-Zmp1. Mtb-Zp-CT showed the highest accuracy (AUC = 0.93), with > 93% specificity and ~ 76% sensitivity.These findings establish immunologically optimized Zmp1 peptides as promising serological biomarkers for TB, including EPTB. Although further optimization may be required to meet WHO target product profiles, their high specificity supports potential use as an adjunctive diagnostic, particularly for EPTB, and as components of future multi-antigen panels. More broadly, this study demonstrates how mechanistically informed epitope selection can facilitate translation of pathogen-specific immune signatures into minimally invasive diagnostic platforms.
Additional Links: PMID-43018675
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43018675,
year = {2026},
author = {Zade, OS and Yandrapally, S and Choudari, K and Gaikwad, AV and Panda, R and Neela, VSK and Prudhula, DK and Eedara, VVR and Ansari, MS and Chandrashekhar, C and Sriram, D and Mohareer, K and Valluri, VL and Somvanshi, PR and Banerjee, S},
title = {Immunologically optimized Zmp1 peptides reveal a translational serological biomarker platform for tuberculosis diagnosis across disease manifestations.},
journal = {European journal of clinical microbiology & infectious diseases : official publication of the European Society of Clinical Microbiology},
volume = {},
number = {},
pages = {},
pmid = {43018675},
issn = {1435-4373},
support = {(5/8/5/21/ITRC/Diag/2021/ECD-1).//Indian Council of Medical Research/ ; (5/8/5/21/ITRC/Diag/2021/ECD-1).//Indian Council of Medical Research/ ; },
abstract = {Tuberculosis (TB) diagnosis remains challenging, particularly for extrapulmonary TB (EPTB), where invasive sampling, low bacillary burden, and limited sensitivity of nucleic acid-based tests in peripheral specimens hinder timely detection. Here, we report an immunology-driven strategy for biomarker discovery and development of a peptide-based serological assay targeting Mycobacterium tuberculosis zinc metalloprotease-1 (Zmp1). Based on the principle that antigenic regions containing overlapping B-cell and CD4[+]T-cell epitopes can promote strong antibody responses through linked recognition and cognate T-cell help, we used an immunoinformatics pipeline to identify two nested immunodominant Zmp1 peptide regions, Mtb-Zp-NT and Mtb-Zp-CT, enriched for overlapping B- and T-cell epitopes.Their diagnostic potential was evaluated using ELISA-based serological assays. A blinded pilot study (N = 137) demonstrated clear discrimination between active TB and TB-recovered individuals. An expanded validation cohort (N = 875), identified from screening approximately 6,086 individuals, included pulmonary TB (PTB), EPTB, TB-recovered individuals, household contacts, non-specific infection/inflammation, and healthy controls. Receiver operating characteristic analyses, supported by DeLong and bootstrap comparisons, demonstrated superior diagnostic performance of the peptide assays compared with Mtb-FL-Zmp1. Mtb-Zp-CT showed the highest accuracy (AUC = 0.93), with > 93% specificity and ~ 76% sensitivity.These findings establish immunologically optimized Zmp1 peptides as promising serological biomarkers for TB, including EPTB. Although further optimization may be required to meet WHO target product profiles, their high specificity supports potential use as an adjunctive diagnostic, particularly for EPTB, and as components of future multi-antigen panels. More broadly, this study demonstrates how mechanistically informed epitope selection can facilitate translation of pathogen-specific immune signatures into minimally invasive diagnostic platforms.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Optimizing tissue lysis and DNA extraction protocols to enhance bacterial diversity profiling in the Drosophila melanogaster gut microbiome.
Scientific reports, 16:34962.
UNLABELLED: The gut microbiota is a dynamic community that influences host metabolism, immunity, and overall health. Accurate characterization of this community requires robust and reproducible DNA extraction methods; however, technical biases introduced during tissue lysis and DNA isolation remain major challenges in microbiome research, particularly in animal model systems. In this study, we compared two lysis methods (manual pestle homogenization and bead-beating) and two commercial DNA extraction kits (Qiagen and Zymo) to evaluate their impact on microbiota profiling in a microbial community standard (MCS) and Drosophila melanogaster gut samples, a tractable model for host-microbe interactions. Full-length 16S rRNA sequencing was performed using Oxford Nanopore Technologies (ONT), followed by taxonomic classification with EPI2ME and downstream diversity analyses using standard pipelines implemented through a custom script. Our data revealed that extraction and lysis methods influence microbial composition, with some protocols resulting in additional species-level assignments in MCS samples, likely reflecting limitations of sequencing technology and taxonomic misidentifications. Pestle homogenization, combined with the Qiagen kit, recovered the highest number of detected taxa and showed a more balanced representation of both Gram-positive and Gram-negative bacteria across the Drosophila gut samples. These findings highlight that extraction methodology influences microbial diversity estimates in Drosophila gut samples, although sequencing approach and taxonomic classification accuracy may also contribute to the observed differences. Our results emphasize that the choice of DNA extraction protocol should be guided by the biological question being addressed and highlight the need for standardized protocols to ensure reproducibility across microbiome studies, particularly those using model systems.
SUPPLEMENTARY INFORMATION: The online version contains supplementary material available at https://doi.org/10.1038/s41598-026-63122-w.
Additional Links: PMID-43019118
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43019118,
year = {2026},
author = {Quiñones-Sanchez, CL and Bilbao-Del Valle, JL and Urdaneta-Colon, MA and Santiago-Rodriguez, TM and Rodriguez-Fernandez, IA},
title = {Optimizing tissue lysis and DNA extraction protocols to enhance bacterial diversity profiling in the Drosophila melanogaster gut microbiome.},
journal = {Scientific reports},
volume = {16},
number = {},
pages = {34962},
pmid = {43019118},
issn = {2045-2322},
support = {P20 GM103642/GM/NIGMS NIH HHS/United States ; R25 HG012702/HG/NHGRI NIH HHS/United States ; },
mesh = {Animals ; *Drosophila melanogaster/microbiology ; RNA, Ribosomal, 16S/genetics ; *DNA, Bacterial/isolation & purification/genetics ; *Gastrointestinal Microbiome/genetics ; *Bacteria/genetics/classification ; Biodiversity ; Sequence Analysis, DNA ; },
abstract = {UNLABELLED: The gut microbiota is a dynamic community that influences host metabolism, immunity, and overall health. Accurate characterization of this community requires robust and reproducible DNA extraction methods; however, technical biases introduced during tissue lysis and DNA isolation remain major challenges in microbiome research, particularly in animal model systems. In this study, we compared two lysis methods (manual pestle homogenization and bead-beating) and two commercial DNA extraction kits (Qiagen and Zymo) to evaluate their impact on microbiota profiling in a microbial community standard (MCS) and Drosophila melanogaster gut samples, a tractable model for host-microbe interactions. Full-length 16S rRNA sequencing was performed using Oxford Nanopore Technologies (ONT), followed by taxonomic classification with EPI2ME and downstream diversity analyses using standard pipelines implemented through a custom script. Our data revealed that extraction and lysis methods influence microbial composition, with some protocols resulting in additional species-level assignments in MCS samples, likely reflecting limitations of sequencing technology and taxonomic misidentifications. Pestle homogenization, combined with the Qiagen kit, recovered the highest number of detected taxa and showed a more balanced representation of both Gram-positive and Gram-negative bacteria across the Drosophila gut samples. These findings highlight that extraction methodology influences microbial diversity estimates in Drosophila gut samples, although sequencing approach and taxonomic classification accuracy may also contribute to the observed differences. Our results emphasize that the choice of DNA extraction protocol should be guided by the biological question being addressed and highlight the need for standardized protocols to ensure reproducibility across microbiome studies, particularly those using model systems.
SUPPLEMENTARY INFORMATION: The online version contains supplementary material available at https://doi.org/10.1038/s41598-026-63122-w.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Animals
*Drosophila melanogaster/microbiology
RNA, Ribosomal, 16S/genetics
*DNA, Bacterial/isolation & purification/genetics
*Gastrointestinal Microbiome/genetics
*Bacteria/genetics/classification
Biodiversity
Sequence Analysis, DNA
RevDate: 2026-11-12
Nitrogen availability modulates soil microbial carbon fixation in wetland through community and metabolic remodelling.
Water research, 309(Pt C):127239 pii:S0043-1354(26)01910-X [Epub ahead of print].
Excess agricultural nitrogen (N) from farm drainage and runoff causes wetland water N pollution and destabilizes soil carbon (C) sinks, however, how this N enrichment regulates wetland soil microbial autotrophic C fixation remains unclear. Here, we combined field experiments, [13]CO2 continuous pulse labeling, DNA-stable isotope probing and metagenomics to investigate the dose-dependent effects of N input on soil microbial community assembly, co-occurrence networks, C-fixation pathways, and soil physicochemical properties in Songnen Plain saline-alkali wetlands of China. The results indicate that N input exerts a threshold regulation on microbial C fixation, with distinct depth-dependent responses. Intermediate N input (N2, 10 g N m[-2] yr[-1]) alleviates soil microbial N limitation, elevates RubisCO and hydrolase activities, optimizes soil CN-phosphorus stoichiometry, and promotes the accumulation of mineral-associated organic C. Such intermediate N input remodels microbial community structure, increases microbial diversity and network stability, and enriches key C-fixing taxa including Cyanobacteriota and Alphaproteobacteria, thereby upregulating six core autotrophic C-fixation pathways. In contrast, excessive N (N4, 24 g N m[-2] yr[-1]) input induces soil acidification and nutrient imbalance, disrupts microbial network stability, declines functional C-fixing microbiota and suppresses C-fixation gene expression, ultimately reducing soil CO2 assimilation and organic C accumulation. Structural equation modelling and random forest analyses further reveal soil microbial biomass N and RubisCO activity are the primary drivers of microbial C fixation, mediated by soil microbial C metabolic pathways and community diversity. This work reveals mechanisms of wetland soil C fixation under agricultural N pollution and informs wetland nutrient management and C-sink conservation.
Additional Links: PMID-43020003
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43020003,
year = {2026},
author = {Luo, S and Song, Y and Zhao, Y and Zhang, H and Ren, J and Zhang, J and Song, C},
title = {Nitrogen availability modulates soil microbial carbon fixation in wetland through community and metabolic remodelling.},
journal = {Water research},
volume = {309},
number = {Pt C},
pages = {127239},
doi = {10.1016/j.watres.2026.127239},
pmid = {43020003},
issn = {1879-2448},
abstract = {Excess agricultural nitrogen (N) from farm drainage and runoff causes wetland water N pollution and destabilizes soil carbon (C) sinks, however, how this N enrichment regulates wetland soil microbial autotrophic C fixation remains unclear. Here, we combined field experiments, [13]CO2 continuous pulse labeling, DNA-stable isotope probing and metagenomics to investigate the dose-dependent effects of N input on soil microbial community assembly, co-occurrence networks, C-fixation pathways, and soil physicochemical properties in Songnen Plain saline-alkali wetlands of China. The results indicate that N input exerts a threshold regulation on microbial C fixation, with distinct depth-dependent responses. Intermediate N input (N2, 10 g N m[-2] yr[-1]) alleviates soil microbial N limitation, elevates RubisCO and hydrolase activities, optimizes soil CN-phosphorus stoichiometry, and promotes the accumulation of mineral-associated organic C. Such intermediate N input remodels microbial community structure, increases microbial diversity and network stability, and enriches key C-fixing taxa including Cyanobacteriota and Alphaproteobacteria, thereby upregulating six core autotrophic C-fixation pathways. In contrast, excessive N (N4, 24 g N m[-2] yr[-1]) input induces soil acidification and nutrient imbalance, disrupts microbial network stability, declines functional C-fixing microbiota and suppresses C-fixation gene expression, ultimately reducing soil CO2 assimilation and organic C accumulation. Structural equation modelling and random forest analyses further reveal soil microbial biomass N and RubisCO activity are the primary drivers of microbial C fixation, mediated by soil microbial C metabolic pathways and community diversity. This work reveals mechanisms of wetland soil C fixation under agricultural N pollution and informs wetland nutrient management and C-sink conservation.},
}
RevDate: 2026-11-12
Microbial-driven cryptic sulfur recycling in volcanic barrier lakes: seasonal dynamics of biogenic sulfide.
Environmental research pii:S0013-9351(26)02496-5 [Epub ahead of print].
Seasonal hydrological fluctuations shape biogeochemical cycling of carbon-nitrogen-sulfur (C-N-S) and sustain aquatic ecosystem functioning, wherein biogenic sulfide (BioS) acts as a critical cryptic regulator. Microbial sulfur cycling is well-recognized, yet the functional importance of BioS - re-oxidized intermediates of dissimilatory sulfate reduction fueling internal sulfur recycling - remains poorly constrained. Using Wudalianchi as a model system, we examined seasonal sulfate dynamics across summer, autumn and late winter. We integrated isotopic tracing, functional-gene profiling and metagenomics alongside statistical screening, machine-learning driver identification, and isotopic quantitative modelling. Microbial community dynamics were primarily governed by seasonal changes in temperature (63 %), sulfate concentration (37 %), and sulfur and oxygen isotopes (δ[18]OSO4, 49%; δ[34]SSO4, 45%). Sulfate sources shifted from precipitation-dominated inputs in summer (∼73 %) toward sediment-derived sulfide oxidation in autumn and winter (∼74-83 %), concurrent with elevated BioS fractions (up to ∼20 %). This seasonal transition was controlled by turnover of key bacterial taxa (Microcystis, Limnohabitans, Polynucleobacter) and topological rewiring of microbial co-occurrence networks. Summer-dominant Microcystis may modulate BioS via oxygenic Fe(II) oxidation, whereas autumn-enriched Limnohabitans has genomic potential for sulfide re-oxidation via the SOX system. In winter, sulfur-metabolizing and disproportionating consortia sustained low-magnitude but persistent cryptic sulfur recycling. BioS forms an essential cryptic cycling loop that stabilizes sulfur availability, modulates isotopic signatures, and sustains seasonal C-N-S cycling under limited external inputs. This work elucidates the underrecognized BioS-mediated mechanism connecting seasonal hydrological variability to internal nutrient cycling. It advances mechanistic insights into freshwater sulfur biogeochemistry with broader implications for aquatic management under global environmental change.
Additional Links: PMID-43020410
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43020410,
year = {2026},
author = {Zou, J and Zhang, C and Fang, Z},
title = {Microbial-driven cryptic sulfur recycling in volcanic barrier lakes: seasonal dynamics of biogenic sulfide.},
journal = {Environmental research},
volume = {},
number = {},
pages = {126159},
doi = {10.1016/j.envres.2026.126159},
pmid = {43020410},
issn = {1096-0953},
abstract = {Seasonal hydrological fluctuations shape biogeochemical cycling of carbon-nitrogen-sulfur (C-N-S) and sustain aquatic ecosystem functioning, wherein biogenic sulfide (BioS) acts as a critical cryptic regulator. Microbial sulfur cycling is well-recognized, yet the functional importance of BioS - re-oxidized intermediates of dissimilatory sulfate reduction fueling internal sulfur recycling - remains poorly constrained. Using Wudalianchi as a model system, we examined seasonal sulfate dynamics across summer, autumn and late winter. We integrated isotopic tracing, functional-gene profiling and metagenomics alongside statistical screening, machine-learning driver identification, and isotopic quantitative modelling. Microbial community dynamics were primarily governed by seasonal changes in temperature (63 %), sulfate concentration (37 %), and sulfur and oxygen isotopes (δ[18]OSO4, 49%; δ[34]SSO4, 45%). Sulfate sources shifted from precipitation-dominated inputs in summer (∼73 %) toward sediment-derived sulfide oxidation in autumn and winter (∼74-83 %), concurrent with elevated BioS fractions (up to ∼20 %). This seasonal transition was controlled by turnover of key bacterial taxa (Microcystis, Limnohabitans, Polynucleobacter) and topological rewiring of microbial co-occurrence networks. Summer-dominant Microcystis may modulate BioS via oxygenic Fe(II) oxidation, whereas autumn-enriched Limnohabitans has genomic potential for sulfide re-oxidation via the SOX system. In winter, sulfur-metabolizing and disproportionating consortia sustained low-magnitude but persistent cryptic sulfur recycling. BioS forms an essential cryptic cycling loop that stabilizes sulfur availability, modulates isotopic signatures, and sustains seasonal C-N-S cycling under limited external inputs. This work elucidates the underrecognized BioS-mediated mechanism connecting seasonal hydrological variability to internal nutrient cycling. It advances mechanistic insights into freshwater sulfur biogeochemistry with broader implications for aquatic management under global environmental change.},
}
RevDate: 2026-11-12
Metagenome-assembled genomes from lab-scale phosphorus-removal bioreactor.
Microbiology resource announcements [Epub ahead of print].
We report the metagenome of a microbial community enriched in phosphate-accumulating organisms (PAOs) from a laboratory batch reactor. A total of 23 metagenome-assembled genomes of the community members were obtained, including putative PAOs, glycogen-accumulating microorganisms, and nitrifiers. These data will contribute to the study of the metabolic capabilities of phosphate-removing communities.
Additional Links: PMID-43020474
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43020474,
year = {2026},
author = {Gruzdev, EV and Dorofeev, AG and Pelevina, AV and Beletsky, AV and Pimenov, NV and Ravin, NV and Mardanov, AV},
title = {Metagenome-assembled genomes from lab-scale phosphorus-removal bioreactor.},
journal = {Microbiology resource announcements},
volume = {},
number = {},
pages = {e0107126},
doi = {10.1128/mra.01071-26},
pmid = {43020474},
issn = {2576-098X},
abstract = {We report the metagenome of a microbial community enriched in phosphate-accumulating organisms (PAOs) from a laboratory batch reactor. A total of 23 metagenome-assembled genomes of the community members were obtained, including putative PAOs, glycogen-accumulating microorganisms, and nitrifiers. These data will contribute to the study of the metabolic capabilities of phosphate-removing communities.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
Vaginal resistome redundancy is associated with microbial metabolic potential and host glycomic features in endometriosis.
NPJ biofilms and microbiomes, 12(1):.
The vaginal microbiome plays a critical role in reproductive health, yet most studies remain descriptive and functional features connecting microbial communities to host physiology remain poorly understood. In endometriosis, alterations in the vaginal microbiome and host glycosylation have been reported, yet ecological properties connecting these systems remain unclear. We investigated whether vaginal resistome functional redundancy (FR) represents a functional link between microbial community structure, microbial metabolic capacity, and host glycomic variation across endometriosis severity. Shotgun metagenomic sequencing was performed on high vaginal swabs from women with healthy controls, minimal/mild, and moderate/severe endometriosis (n = 40). While total antibiotic resistance genes (ARG) abundance remained relatively stable across disease groups, resistome FR increased significantly in moderate/severe endometriosis, reflecting a transition from low-diversity, Lactobacillus-dominated communities toward more taxonomically diverse, ARG-harbouring taxa. Increased redundancy was associated with higher microbial alpha diversity and enrichment of glycan-related metabolic pathways, particularly UDP-N-acetyl-D-glucosamine biosynthesis, a precursor pathway for glycan assembly. These microbial functional features correlated with host glycomic signatures in serum and urine. Mediation and path modelling analyses supported associations among resistome FR, predicted UDP-sugar pathway potential, host glycomic variation, and endocrine-related traits. Our findings identify resistome FR as a quantifiable ecological property of the vaginal microbiome that is associated with microbial metabolic capacity and host glycomic variation in endometriosis. Rather than acting as a direct driver, FR may serve as a marker of microbiome organisation that relates to host-associated phenotypes, providing a framework for understanding microbiome-glycome interactions in reproductive tract disease.
Additional Links: PMID-43020834
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43020834,
year = {2026},
author = {Pan, Z and Glover, L and Reidy, F and McAuliffe, FM and Wingfield, M and van Sinderen, D and Saldova, R and MacSharry, J},
title = {Vaginal resistome redundancy is associated with microbial metabolic potential and host glycomic features in endometriosis.},
journal = {NPJ biofilms and microbiomes},
volume = {12},
number = {1},
pages = {},
pmid = {43020834},
issn = {2055-5008},
mesh = {Female ; Humans ; *Endometriosis/microbiology/metabolism ; *Vagina/microbiology ; *Microbiota ; *Glycomics ; Metagenomics/methods ; *Bacteria/classification/genetics/metabolism/isolation & purification ; Shotgun Sequencing ; Metagenome ; Metabolic Networks and Pathways ; },
abstract = {The vaginal microbiome plays a critical role in reproductive health, yet most studies remain descriptive and functional features connecting microbial communities to host physiology remain poorly understood. In endometriosis, alterations in the vaginal microbiome and host glycosylation have been reported, yet ecological properties connecting these systems remain unclear. We investigated whether vaginal resistome functional redundancy (FR) represents a functional link between microbial community structure, microbial metabolic capacity, and host glycomic variation across endometriosis severity. Shotgun metagenomic sequencing was performed on high vaginal swabs from women with healthy controls, minimal/mild, and moderate/severe endometriosis (n = 40). While total antibiotic resistance genes (ARG) abundance remained relatively stable across disease groups, resistome FR increased significantly in moderate/severe endometriosis, reflecting a transition from low-diversity, Lactobacillus-dominated communities toward more taxonomically diverse, ARG-harbouring taxa. Increased redundancy was associated with higher microbial alpha diversity and enrichment of glycan-related metabolic pathways, particularly UDP-N-acetyl-D-glucosamine biosynthesis, a precursor pathway for glycan assembly. These microbial functional features correlated with host glycomic signatures in serum and urine. Mediation and path modelling analyses supported associations among resistome FR, predicted UDP-sugar pathway potential, host glycomic variation, and endocrine-related traits. Our findings identify resistome FR as a quantifiable ecological property of the vaginal microbiome that is associated with microbial metabolic capacity and host glycomic variation in endometriosis. Rather than acting as a direct driver, FR may serve as a marker of microbiome organisation that relates to host-associated phenotypes, providing a framework for understanding microbiome-glycome interactions in reproductive tract disease.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Female
Humans
*Endometriosis/microbiology/metabolism
*Vagina/microbiology
*Microbiota
*Glycomics
Metagenomics/methods
*Bacteria/classification/genetics/metabolism/isolation & purification
Shotgun Sequencing
Metagenome
Metabolic Networks and Pathways
RevDate: 2026-11-12
CmpDate: 2026-11-12
NemaCapsules: microfluidics-integrated biocells for investigating C. elegans host-microbiome interactions in spaceflight.
NPJ microgravity, 12(1):.
Spaceflight studies in humans show that microgravity alters the gut microbiome, posing a risk to health. Understanding how microbiomes and host genetics influence physiology is critical for long-duration missions. The low-cost spaceflight model C. elegans provides a suitable means to study host-microbiome interactions. However, whole-organism measurements are limited by lacking suitable flight-ready hardware that enables long-term nematode culture compatible with on-orbit imaging. These limitations were addressed through development of NemaCapsules- microfluidics-integrated biocells, for spaceflight assessment of C. elegans locomotion under defined microbiome conditions, with minimal astronaut workload. This provided a fully sealed, gas-permeable system with media reservoir, enabling passive, long-term nematode culturing. NemaCapsules were evaluated across four host strains and three model microbiomes. Multi-day worm viability studies demonstrated high resilience across diverse host strains and microbiomes except for daf-16 mutants. Crawling and swimming studies of worms with model microbiomes of C. elegans natural microbiota exhibited distinct locomotory activity from those exposed to E. coli OP50. Wild-type N2 generally exhibited the highest activity, followed by wild isolate strains; daf-16 mutants demonstrated lowest activity. Compared to crawling, swimming better revealed differences arising from host-microbiome interactions. NemaCapsules provide a modular solution for on-orbit, multi-functional readouts in C. elegans with minimal crew intervention.
Additional Links: PMID-43020865
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43020865,
year = {2026},
author = {Rahman, B and Saroyia, AP and Blackburn, D and Parish, AJ and Szewczyk, N and Driscoll, M and Samuel, BS and Vanapalli, SA},
title = {NemaCapsules: microfluidics-integrated biocells for investigating C. elegans host-microbiome interactions in spaceflight.},
journal = {NPJ microgravity},
volume = {12},
number = {1},
pages = {},
pmid = {43020865},
issn = {2373-8065},
support = {project# 80NSSC22K0250//NASA/ ; },
abstract = {Spaceflight studies in humans show that microgravity alters the gut microbiome, posing a risk to health. Understanding how microbiomes and host genetics influence physiology is critical for long-duration missions. The low-cost spaceflight model C. elegans provides a suitable means to study host-microbiome interactions. However, whole-organism measurements are limited by lacking suitable flight-ready hardware that enables long-term nematode culture compatible with on-orbit imaging. These limitations were addressed through development of NemaCapsules- microfluidics-integrated biocells, for spaceflight assessment of C. elegans locomotion under defined microbiome conditions, with minimal astronaut workload. This provided a fully sealed, gas-permeable system with media reservoir, enabling passive, long-term nematode culturing. NemaCapsules were evaluated across four host strains and three model microbiomes. Multi-day worm viability studies demonstrated high resilience across diverse host strains and microbiomes except for daf-16 mutants. Crawling and swimming studies of worms with model microbiomes of C. elegans natural microbiota exhibited distinct locomotory activity from those exposed to E. coli OP50. Wild-type N2 generally exhibited the highest activity, followed by wild isolate strains; daf-16 mutants demonstrated lowest activity. Compared to crawling, swimming better revealed differences arising from host-microbiome interactions. NemaCapsules provide a modular solution for on-orbit, multi-functional readouts in C. elegans with minimal crew intervention.},
}
RevDate: 2026-11-12
CmpDate: 2026-11-12
PHI: a Galaxy-based workflow for reproducible prophage-host interaction analysis and standardized viral-genomics reporting.
BMC bioinformatics, 27(1):.
BACKGROUND: Viruses that infect bacteria, known as bacteriophages or phages, are widespread in nature and play important roles in shaping microbial communities and ecosystem functions. Some phages can integrate into bacterial genomes as "prophages", where they may influence the biology of their host by carrying genes that affect metabolism, virulence, or environmental adaptation. Despite their importance, studying prophages and their interactions with bacterial hosts remains challenging because it typically requires combining many complex computational tools and can be resource-intensive.
RESULTS: In this study, we introduce the Prophage-Host Interaction Toolkit (PHI), a user-friendly and automated workflow available through the Galaxy platform. PHI brings together multiple established tools into a single, reproducible pipeline that identifies candidate prophages, evaluates their quality, predicts host relationships, and characterizes key functional genes. Importantly, all results are summarized in an interactive report that simplifies interpretation. Applied to the MBARC22 mock community of 22 bacterial reference genomes as a workflow demonstration, PHI identified 41 candidate prophages across 14 hosts, including high- and medium-quality viral genomes. Most host genomes exhibited > 99% completeness and < 1% contamination, while DefenseFinder identified between 3 and 24 antiviral defense systems per genome. To further evaluate applicability to metagenome-derived assemblies, PHI was also applied to three MAGs and their matched reference genomes. Differences in candidate prophage counts and sequence-level correspondence between MAGs and references demonstrated that prophage recovery can be strongly influenced by genome fragmentation and assembly continuity, even in otherwise high-quality MAGs.
CONCLUSIONS: By removing installation barriers and consolidating the outputs of multiple established tools, PHI lowers the barrier to advanced phage analysis, enabling both specialists and non-experts to explore phage-host interactions and their implications in areas such as microbiome research, biotechnology, and environmental science.
Additional Links: PMID-43021054
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021054,
year = {2026},
author = {Saraiva, JP and Borim Corrêa, F and Bernt, M and Ghanem, N and Nieto, E and Brizola Toscan, R and Wick, LY and Chatzinotas, A},
title = {PHI: a Galaxy-based workflow for reproducible prophage-host interaction analysis and standardized viral-genomics reporting.},
journal = {BMC bioinformatics},
volume = {27},
number = {1},
pages = {},
pmid = {43021054},
issn = {1471-2105},
mesh = {*Prophages/genetics/physiology ; *Genomics/methods ; *Genome, Viral ; *Software ; Workflow ; Genome, Bacterial ; *Host-Pathogen Interactions ; Computational Biology/methods ; Bacteria/virology/genetics ; },
abstract = {BACKGROUND: Viruses that infect bacteria, known as bacteriophages or phages, are widespread in nature and play important roles in shaping microbial communities and ecosystem functions. Some phages can integrate into bacterial genomes as "prophages", where they may influence the biology of their host by carrying genes that affect metabolism, virulence, or environmental adaptation. Despite their importance, studying prophages and their interactions with bacterial hosts remains challenging because it typically requires combining many complex computational tools and can be resource-intensive.
RESULTS: In this study, we introduce the Prophage-Host Interaction Toolkit (PHI), a user-friendly and automated workflow available through the Galaxy platform. PHI brings together multiple established tools into a single, reproducible pipeline that identifies candidate prophages, evaluates their quality, predicts host relationships, and characterizes key functional genes. Importantly, all results are summarized in an interactive report that simplifies interpretation. Applied to the MBARC22 mock community of 22 bacterial reference genomes as a workflow demonstration, PHI identified 41 candidate prophages across 14 hosts, including high- and medium-quality viral genomes. Most host genomes exhibited > 99% completeness and < 1% contamination, while DefenseFinder identified between 3 and 24 antiviral defense systems per genome. To further evaluate applicability to metagenome-derived assemblies, PHI was also applied to three MAGs and their matched reference genomes. Differences in candidate prophage counts and sequence-level correspondence between MAGs and references demonstrated that prophage recovery can be strongly influenced by genome fragmentation and assembly continuity, even in otherwise high-quality MAGs.
CONCLUSIONS: By removing installation barriers and consolidating the outputs of multiple established tools, PHI lowers the barrier to advanced phage analysis, enabling both specialists and non-experts to explore phage-host interactions and their implications in areas such as microbiome research, biotechnology, and environmental science.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Prophages/genetics/physiology
*Genomics/methods
*Genome, Viral
*Software
Workflow
Genome, Bacterial
*Host-Pathogen Interactions
Computational Biology/methods
Bacteria/virology/genetics
RevDate: 2026-11-12
CmpDate: 2026-11-12
Gut microbiota dysbiosis in Post-COVID condition symptoms: relation with chronic fatigue and functional limitation.
BMC microbiology, 26(1):.
BACKGROUND: Post-COVID Condition (PCC) is a heterogeneous clinical syndrome in which chronic fatigue and functional limitation are among the most disabling manifestations. Alterations in the gut microbiota have been proposed as contributors to PCC pathophysiology; however, it remains unclear whether distinct clinical PCC phenotypes are associated with specific microbiota signatures.
RESULTS: This cross-sectional pilot study analysed gut microbiota composition and predicted functional capacity in 50 participants from Aragón (Spain), including 30 patients with PCC and 20 healthy controls. PCC patients were stratified according to the presence of Myalgic Encephalomyelitis/Chronic Fatigue Syndrome (ME/CFS)-like symptoms and functional limitation assessed using the 6-Minute Walking Test. Alpha and beta diversity did not differ significantly between groups (Figs. 2 and 3). Differential abundance analysis identified taxon-level differences between groups. Escherichia/Shigella was enriched in PCC-CFS relative to controls, both overall and in the subgroup without functional limitation, representing the most consistent signal. Additional differences included higher Streptococcus and Veillonella in controls, and higher Flavonifractor and Hungatella in PCC-NCFS. After adjustment for digestive symptoms and vaccine dose, these associations attenuated and none remained significant, indicating that they may be partly driven by confounding.PICRUSt2 functional predictions suggested differences in pathways related to nucleotide and amino acid metabolism; however, none remained significant after correction for multiple testing and should therefore be considered exploratory.
CONCLUSIONS: These findings suggest the presence of phenotype-associated taxonomic differences within PCC despite the absence of global diversity alterations. The results should be considered exploratory and hypothesis-generating due to the limited subgroup sample sizes. Larger studies incorporating metagenomic and metabolomic approaches are needed to validate these observations and clarify their potential clinical relevance.
Additional Links: PMID-43021070
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021070,
year = {2026},
author = {Carpallo-Porcar, B and García-Martínez, L and Jiménez-Sánchez, C and Calvo, S and Roig, FJ},
title = {Gut microbiota dysbiosis in Post-COVID condition symptoms: relation with chronic fatigue and functional limitation.},
journal = {BMC microbiology},
volume = {26},
number = {1},
pages = {},
pmid = {43021070},
issn = {1471-2180},
support = {B61_23D//Departamento de Ciencia, Universidad y Sociedad del Conocimiento, from the Gobierno de Aragón (Spain)/ ; T71_23D//Departamento de Ciencia, Universidad y Sociedad del Conocimiento, from the Gobierno de Aragón (Spain)/ ; },
mesh = {Humans ; *Dysbiosis/microbiology ; Cross-Sectional Studies ; *Fatigue Syndrome, Chronic/microbiology ; Female ; *Gastrointestinal Microbiome ; Middle Aged ; Male ; *COVID-19/complications/microbiology ; *Bacteria/classification/genetics/isolation & purification ; Pilot Projects ; Post-Acute COVID-19 Syndrome ; Adult ; Spain ; SARS-CoV-2 ; Feces/microbiology ; },
abstract = {BACKGROUND: Post-COVID Condition (PCC) is a heterogeneous clinical syndrome in which chronic fatigue and functional limitation are among the most disabling manifestations. Alterations in the gut microbiota have been proposed as contributors to PCC pathophysiology; however, it remains unclear whether distinct clinical PCC phenotypes are associated with specific microbiota signatures.
RESULTS: This cross-sectional pilot study analysed gut microbiota composition and predicted functional capacity in 50 participants from Aragón (Spain), including 30 patients with PCC and 20 healthy controls. PCC patients were stratified according to the presence of Myalgic Encephalomyelitis/Chronic Fatigue Syndrome (ME/CFS)-like symptoms and functional limitation assessed using the 6-Minute Walking Test. Alpha and beta diversity did not differ significantly between groups (Figs. 2 and 3). Differential abundance analysis identified taxon-level differences between groups. Escherichia/Shigella was enriched in PCC-CFS relative to controls, both overall and in the subgroup without functional limitation, representing the most consistent signal. Additional differences included higher Streptococcus and Veillonella in controls, and higher Flavonifractor and Hungatella in PCC-NCFS. After adjustment for digestive symptoms and vaccine dose, these associations attenuated and none remained significant, indicating that they may be partly driven by confounding.PICRUSt2 functional predictions suggested differences in pathways related to nucleotide and amino acid metabolism; however, none remained significant after correction for multiple testing and should therefore be considered exploratory.
CONCLUSIONS: These findings suggest the presence of phenotype-associated taxonomic differences within PCC despite the absence of global diversity alterations. The results should be considered exploratory and hypothesis-generating due to the limited subgroup sample sizes. Larger studies incorporating metagenomic and metabolomic approaches are needed to validate these observations and clarify their potential clinical relevance.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
*Dysbiosis/microbiology
Cross-Sectional Studies
*Fatigue Syndrome, Chronic/microbiology
Female
*Gastrointestinal Microbiome
Middle Aged
Male
*COVID-19/complications/microbiology
*Bacteria/classification/genetics/isolation & purification
Pilot Projects
Post-Acute COVID-19 Syndrome
Adult
Spain
SARS-CoV-2
Feces/microbiology
RevDate: 2026-11-12
CmpDate: 2026-11-13
Nocardia farcinica brain abscess in an immunocompromised patient receiving tofacitinib: a case report.
BMC infectious diseases, 26(1):.
BACKGROUND: Nocardia farcinica (N. farcinica) brain abscess is a rare but potentially life-threatening opportunistic infection that predominantly occurs in immunocompromised individuals. Patients receiving immunosuppressive therapies may be at increased risk of opportunistic infections; however, the clinical recognition and diagnosis of N.farcinica brain abscess remain challenging.
CASE PRESENTATION: A 55-year-old woman presented with a one-month history of headache and right-sided limb numbness. Brain magnetic resonance imaging (MRI) revealed multiple intracranial space-occupying lesions. Brain biopsy excluded primary or metastatic malignancy and suggested an infectious etiology. The patient had been receiving long-term tofacitinib therapy for two years, with prednisolone and azathioprine discontinued approximately two years before the onset of infection. Next-generation sequencing (NGS) combined with circulating tumor DNA (ctDNA) analysis of brain tissue identified N. farcinica as the causative pathogen. Tofacitinib was discontinued, and the patient was treated with intravenous imipenem-cilastatin (0.5 g every 8 h) and amikacin (0.4 g once daily), combined with oral trimethoprim-sulfamethoxazole (0.48 g every 12 h). After 2.5 months of antimicrobial therapy, the patient showed marked clinical and radiological improvement. Treatment was continued for a total of six months, with no evidence of recurrence during follow-up.
CONCLUSION: This case highlights the importance of considering N. farcinica brain abscess in immunocompromised patients presenting with intracranial lesions. Tissue-based metagenomic sequencing combined with ctDNA analysis may facilitate rapid and accurate identification of N. farcinica, enabling the timely initiation of targeted antimicrobial therapy.
CLINICAL TRIAL NUMBER: Not applicable. This report retrospectively describes the clinical course and management of a patient and did not involve any experimental interventions on a live subject.
Additional Links: PMID-43021133
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021133,
year = {2026},
author = {Huang, S and Bei, Y and Yuan, L and Su, T and Gao, W and Zhou, S and Nan, H},
title = {Nocardia farcinica brain abscess in an immunocompromised patient receiving tofacitinib: a case report.},
journal = {BMC infectious diseases},
volume = {26},
number = {1},
pages = {},
pmid = {43021133},
issn = {1471-2334},
support = {D202319019427//The Scientific Research Project of Hunan Provincial Health Commission/ ; kq2208144//The Natural Science Foundation Project of Changsha/ ; Grant No. HZ2026ZCYX002//Xuanwu Hospital Talent Convergence Program/ ; },
mesh = {Humans ; Female ; *Brain Abscess/microbiology/drug therapy/diagnostic imaging/diagnosis ; Middle Aged ; *Immunocompromised Host ; *Nocardia Infections/drug therapy/microbiology/diagnosis/diagnostic imaging ; *Nocardia/isolation & purification/genetics/drug effects ; *Pyrimidines/therapeutic use/adverse effects ; *Piperidines/therapeutic use/adverse effects ; Trimethoprim, Sulfamethoxazole Drug Combination/therapeutic use ; Magnetic Resonance Imaging ; Anti-Bacterial Agents/therapeutic use ; Cilastatin, Imipenem Drug Combination/therapeutic use ; Brain/diagnostic imaging/pathology/microbiology ; Opportunistic Infections/drug therapy/microbiology ; Cilastatin/therapeutic use ; Imipenem/therapeutic use ; },
abstract = {BACKGROUND: Nocardia farcinica (N. farcinica) brain abscess is a rare but potentially life-threatening opportunistic infection that predominantly occurs in immunocompromised individuals. Patients receiving immunosuppressive therapies may be at increased risk of opportunistic infections; however, the clinical recognition and diagnosis of N.farcinica brain abscess remain challenging.
CASE PRESENTATION: A 55-year-old woman presented with a one-month history of headache and right-sided limb numbness. Brain magnetic resonance imaging (MRI) revealed multiple intracranial space-occupying lesions. Brain biopsy excluded primary or metastatic malignancy and suggested an infectious etiology. The patient had been receiving long-term tofacitinib therapy for two years, with prednisolone and azathioprine discontinued approximately two years before the onset of infection. Next-generation sequencing (NGS) combined with circulating tumor DNA (ctDNA) analysis of brain tissue identified N. farcinica as the causative pathogen. Tofacitinib was discontinued, and the patient was treated with intravenous imipenem-cilastatin (0.5 g every 8 h) and amikacin (0.4 g once daily), combined with oral trimethoprim-sulfamethoxazole (0.48 g every 12 h). After 2.5 months of antimicrobial therapy, the patient showed marked clinical and radiological improvement. Treatment was continued for a total of six months, with no evidence of recurrence during follow-up.
CONCLUSION: This case highlights the importance of considering N. farcinica brain abscess in immunocompromised patients presenting with intracranial lesions. Tissue-based metagenomic sequencing combined with ctDNA analysis may facilitate rapid and accurate identification of N. farcinica, enabling the timely initiation of targeted antimicrobial therapy.
CLINICAL TRIAL NUMBER: Not applicable. This report retrospectively describes the clinical course and management of a patient and did not involve any experimental interventions on a live subject.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
Female
*Brain Abscess/microbiology/drug therapy/diagnostic imaging/diagnosis
Middle Aged
*Immunocompromised Host
*Nocardia Infections/drug therapy/microbiology/diagnosis/diagnostic imaging
*Nocardia/isolation & purification/genetics/drug effects
*Pyrimidines/therapeutic use/adverse effects
*Piperidines/therapeutic use/adverse effects
Trimethoprim, Sulfamethoxazole Drug Combination/therapeutic use
Magnetic Resonance Imaging
Anti-Bacterial Agents/therapeutic use
Cilastatin, Imipenem Drug Combination/therapeutic use
Brain/diagnostic imaging/pathology/microbiology
Opportunistic Infections/drug therapy/microbiology
Cilastatin/therapeutic use
Imipenem/therapeutic use
RevDate: 2026-11-13
CmpDate: 2026-11-13
Gut microbiota-driven hippuric acid hydrolysis promotes pancreatic tumorigenesis by suppressing NK cell activity.
Journal of translational medicine, 24(1):.
BACKGROUND: Pancreatic ductal adenocarcinoma (PDAC) remains one of the most lethal malignancies, characterized by a lack of effective early detection, profoundly immunosuppressive microenvironment, and poor responsiveness to current therapies. Emerging evidence suggests that gut microbiota and their metabolites play important roles in pancreatic tumorigenesis. Glycoproteins shed from pancreatic precancerous lesions may influence the composition of the gut microbiota and their metabolites.
METHODS: We established a pancreatic precancerous lesion mouse model and performed integrated fecal metagenomic sequencing and non-targeted metabolomic profiling to characterize microbiota composition and metabolite alterations. Further we use plasma samples from patients, in vitro cytotoxicity assays and in vivo orthotopic pancreatic tumor models, fecal microbiota transplantation and pancreatic homogenate gavage to assess the effect of microbiota and metabolite alterations. And we conduct RNA sequencing, qPCR, ELISA, flow cytometry, molecular docking, and surface plasmon resonance to further explore its mechanisms.
RESULTS: We identified hippuric acid as a key microbiota-derived metabolite that was significantly reduced during pancreatic tumorigenesis, which was associated with suppressed anti-tumor immunity and enhanced PDAC progression. This reduction was associated with enrichment of microbiota harboring hippurate hydrolase and correlated with poor prognosis in PDAC patients. Mechanistically, hippuric acid activated IL-21R-STAT3-TBX21 signaling axis, promoting the expression of cytotoxic effector molecules, including granzyme B and perforin, thus enhancing NK cell cytotoxicity. Molecular docking and surface plasmon resonance analyses further suggested a direct interaction between hippuric acid and IL-21R.
CONCLUSIONS: Collectively, our study uncovers a previously unrecognized positive feedback regulatory loop involving the PDAC-gut microbiota-hippuric acid-NK cell-PDAC axis and highlights the critical role of gut microbiota and their metabolites in pancreatic tumorigenesis.
Additional Links: PMID-43021363
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021363,
year = {2026},
author = {Zhang, X and Qin, C and Zhao, Y and Li, Z and Xie, Y and Yan, Y and Li, T and Huang, L and Shi, H and Lin, C and Wang, W},
title = {Gut microbiota-driven hippuric acid hydrolysis promotes pancreatic tumorigenesis by suppressing NK cell activity.},
journal = {Journal of translational medicine},
volume = {24},
number = {1},
pages = {},
pmid = {43021363},
issn = {1479-5876},
support = {No. 82573412//National Natural Science Foundation of China/ ; No. 82503800//National Natural Science Foundation of China/ ; 2025-PUMCH-C-018//National High Level Hospital Clinical Research Funding/ ; 2025-PUMCH-A-073//National High Level Hospital Clinical Research Funding/ ; 2025-PUMCH-A-075//National High Level Hospital Clinical Research Funding/ ; 2024-2-4017//Capital's Funds for Health Improvement and Research/ ; ZK12101//National Multidisciplinary Cooperative Diagnosis and Treatment Capacity Building Project for Major Diseases/ ; No.UGG04598//Peking Union Medical College Hospital Talent Cultivation Program (Category B)/ ; No. 2024-I2M-ZD-001//CAMS Innovation Fund for Medical Sciences/ ; No. 2024bkjg015,No.2025bkjg005//Peking Union Medical College Hospital Undergraduate Educational Reform Project/ ; No. 2024dcxm100, No.2026dcxm275//Student Research Training Project, Chinese Academy of Medical Sciences & Peking Union Medical College/ ; Number GZC20240146//Postdoctoral Fellowship Program of CPSF under Grant/ ; 3332025001//Fundamental Research Funds for the Central Universities, Peking Union Medical College/ ; },
mesh = {*Killer Cells, Natural/immunology ; Animals ; Humans ; *Pancreatic Neoplasms/pathology/microbiology/immunology ; *Hippurates/metabolism ; *Carcinogenesis/pathology ; *Gastrointestinal Microbiome ; Hydrolysis ; Cell Line, Tumor ; Carcinoma, Pancreatic Ductal/pathology/microbiology ; Mice, Inbred C57BL ; Male ; Signal Transduction ; },
abstract = {BACKGROUND: Pancreatic ductal adenocarcinoma (PDAC) remains one of the most lethal malignancies, characterized by a lack of effective early detection, profoundly immunosuppressive microenvironment, and poor responsiveness to current therapies. Emerging evidence suggests that gut microbiota and their metabolites play important roles in pancreatic tumorigenesis. Glycoproteins shed from pancreatic precancerous lesions may influence the composition of the gut microbiota and their metabolites.
METHODS: We established a pancreatic precancerous lesion mouse model and performed integrated fecal metagenomic sequencing and non-targeted metabolomic profiling to characterize microbiota composition and metabolite alterations. Further we use plasma samples from patients, in vitro cytotoxicity assays and in vivo orthotopic pancreatic tumor models, fecal microbiota transplantation and pancreatic homogenate gavage to assess the effect of microbiota and metabolite alterations. And we conduct RNA sequencing, qPCR, ELISA, flow cytometry, molecular docking, and surface plasmon resonance to further explore its mechanisms.
RESULTS: We identified hippuric acid as a key microbiota-derived metabolite that was significantly reduced during pancreatic tumorigenesis, which was associated with suppressed anti-tumor immunity and enhanced PDAC progression. This reduction was associated with enrichment of microbiota harboring hippurate hydrolase and correlated with poor prognosis in PDAC patients. Mechanistically, hippuric acid activated IL-21R-STAT3-TBX21 signaling axis, promoting the expression of cytotoxic effector molecules, including granzyme B and perforin, thus enhancing NK cell cytotoxicity. Molecular docking and surface plasmon resonance analyses further suggested a direct interaction between hippuric acid and IL-21R.
CONCLUSIONS: Collectively, our study uncovers a previously unrecognized positive feedback regulatory loop involving the PDAC-gut microbiota-hippuric acid-NK cell-PDAC axis and highlights the critical role of gut microbiota and their metabolites in pancreatic tumorigenesis.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Killer Cells, Natural/immunology
Animals
Humans
*Pancreatic Neoplasms/pathology/microbiology/immunology
*Hippurates/metabolism
*Carcinogenesis/pathology
*Gastrointestinal Microbiome
Hydrolysis
Cell Line, Tumor
Carcinoma, Pancreatic Ductal/pathology/microbiology
Mice, Inbred C57BL
Male
Signal Transduction
RevDate: 2026-11-13
Syndromic diagnosis and precision medicine in severe infections.
Revista espanola de quimioterapia : publicacion oficial de la Sociedad Espanola de Quimioterapia pii:v39.s01.03.2026 [Epub ahead of print].
The implementation of syndromic PCR panels has transformed the management of severe infections, including bloodstream, lower respiratory tract, and central nervous system infections, by enabling rapid pathogen identification and earlier optimization of antimicrobial therapy. As regulatory, technical, and interpretative challenges are overcome, unbiased shotgun metagenomic next-generation sequencing is expected to complement these platforms and become integrated into routine clinical practice, offering comprehensive, hypothesis-free detection of known and emerging pathogens in a single assay.
Additional Links: PMID-43021616
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021616,
year = {2026},
author = {Clari, MÁ and Carbonell, N and Navarro, D},
title = {Syndromic diagnosis and precision medicine in severe infections.},
journal = {Revista espanola de quimioterapia : publicacion oficial de la Sociedad Espanola de Quimioterapia},
volume = {},
number = {},
pages = {},
doi = {10.37201/req/v39.s01.03.2026},
pmid = {43021616},
issn = {1988-9518},
abstract = {The implementation of syndromic PCR panels has transformed the management of severe infections, including bloodstream, lower respiratory tract, and central nervous system infections, by enabling rapid pathogen identification and earlier optimization of antimicrobial therapy. As regulatory, technical, and interpretative challenges are overcome, unbiased shotgun metagenomic next-generation sequencing is expected to complement these platforms and become integrated into routine clinical practice, offering comprehensive, hypothesis-free detection of known and emerging pathogens in a single assay.},
}
RevDate: 2026-11-13
Molecular diagnostics in immunocompromised patients: promises, pitfalls, and practical strategies.
Revista espanola de quimioterapia : publicacion oficial de la Sociedad Espanola de Quimioterapia pii:v39.s01.04.2026 [Epub ahead of print].
Infections remain a leading cause of morbidity and mortality in immunocompromised patients, frequently presenting with atypical clinical manifestations and a broad spectrum of potential pathogens. Advances in molecular diagnostics, particularly multiplex PCR panels and metagenomic next-generation sequencing, have substantially improved pathogen detection rates and reduced time to diagnosis. Nevertheless, their routine clinical implementation remains challenging. Key limitations include: the inability to reliably distinguish colonization from active infection, high costs, and the risk of over-interpretation in the absence of clinical correlation. This mini-review provides an etiological framework for infections in immunocompromised hosts, critically examines the indications and limitations of current molecular diagnostic platforms, and proposes integrated multimodal strategies to optimize clinical decision-making and antimicrobial stewardship.
Additional Links: PMID-43021617
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021617,
year = {2026},
author = {Rua, M and Pina-Sánchez, M and Candel, FJ and Del Pozo, JL},
title = {Molecular diagnostics in immunocompromised patients: promises, pitfalls, and practical strategies.},
journal = {Revista espanola de quimioterapia : publicacion oficial de la Sociedad Espanola de Quimioterapia},
volume = {},
number = {},
pages = {},
doi = {10.37201/req/v39.s01.04.2026},
pmid = {43021617},
issn = {1988-9518},
abstract = {Infections remain a leading cause of morbidity and mortality in immunocompromised patients, frequently presenting with atypical clinical manifestations and a broad spectrum of potential pathogens. Advances in molecular diagnostics, particularly multiplex PCR panels and metagenomic next-generation sequencing, have substantially improved pathogen detection rates and reduced time to diagnosis. Nevertheless, their routine clinical implementation remains challenging. Key limitations include: the inability to reliably distinguish colonization from active infection, high costs, and the risk of over-interpretation in the absence of clinical correlation. This mini-review provides an etiological framework for infections in immunocompromised hosts, critically examines the indications and limitations of current molecular diagnostic platforms, and proposes integrated multimodal strategies to optimize clinical decision-making and antimicrobial stewardship.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Diagnostic performance of metagenomic next-generation sequencing in Mycobacterium and non-Mycobacterium infection patients: a retrospective study at a large comprehensive medical center.
Frontiers in cellular and infection microbiology, 16:1935536.
BACKGROUND: Metagenomic next-generation sequencing (mNGS) has become a potentially valuable complementary test, if not an essential one, in some challenging clinical cases. This study comprehensively compared mNGS with other methods, including TB-IgG, TB-IGRA, TB-DNA, Xpert, culture, and AFS for the diagnosis of and differentiation between TB infection from NTM infection using multiple sample types, including BALF, CSF, blood, pus, tissue, hydrothorax, and ascites.
METHODS: From 2021 to 2024, the clinical data, including imaging, laboratory results, and medical records, of 425 patients with suspected TB infection were retrospectively collected and classified as PTB [306 (72%)], EPTB [67 (15.76%)], and non-TB [52 (12.24%)], according to the final clinical diagnosis.
RESULTS: Among the three groups, significant differences were observed in age, sex, cough, smoking history, hsCRP, IFN-γ, and hypertension (P < 0.05). According to the final clinical diagnosis, the sensitivity and specificity of mNGS in all samples were 90.1% (95% CI: 0.866-0.927) and 75.0% (95% CI: 0.618-0.848), respectively. The sensitivity of mNGS was higher than that of TB-IGRA (79.1%), Xpert (72.4%), culture (52.3%), TB-DNA (45.6%), TB-IgG (36.4%), and AFS (10.7%). For distinguishing TB from non-TB, the sensitivity of mNGS was approximately 37.8% higher than that of standard test culture (90.1% vs 36.4%; P < 0.001). Further analysis also showed that MTB read counts from mNGS were highly correlated with MTB burden measured by other laboratory tests (P < 0.05). Analysis of the ROC curve of mNGS reads showed that the AUC for MTB diagnosis was 0.9073, with a sensitivity of 73.19% and specificity of 96.15% when the recommended cutoff value of 4 reads (Youden's index, 0.6934) was applied.
CONCLUSIONS: For patients with suspected TB or NTM infection, mNGS may serve as a helpful adjunctive tool, particularly when routine diagnostic tests are unrevealing or indeterminate. It has the potential to assist in distinguishing MTB from NTM and to provide additional evidence to guide appropriate treatment choices, though its results should always be interpreted alongside clinical findings and conventional microbiological testing.
Additional Links: PMID-43021990
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43021990,
year = {2026},
author = {Zhou, X and Peng, S and Tan, R and Wang, J and Hu, H and Zhang, K and Liu, A},
title = {Diagnostic performance of metagenomic next-generation sequencing in Mycobacterium and non-Mycobacterium infection patients: a retrospective study at a large comprehensive medical center.},
journal = {Frontiers in cellular and infection microbiology},
volume = {16},
number = {},
pages = {1935536},
pmid = {43021990},
issn = {2235-2988},
mesh = {Humans ; Retrospective Studies ; *High-Throughput Nucleotide Sequencing/methods ; *Metagenomics/methods ; Sensitivity and Specificity ; Female ; Male ; Middle Aged ; Adult ; Aged ; *Tuberculosis/diagnosis/microbiology ; *Mycobacterium Infections, Nontuberculous/diagnosis/microbiology ; Diagnosis, Differential ; *Mycobacterium/genetics ; },
abstract = {BACKGROUND: Metagenomic next-generation sequencing (mNGS) has become a potentially valuable complementary test, if not an essential one, in some challenging clinical cases. This study comprehensively compared mNGS with other methods, including TB-IgG, TB-IGRA, TB-DNA, Xpert, culture, and AFS for the diagnosis of and differentiation between TB infection from NTM infection using multiple sample types, including BALF, CSF, blood, pus, tissue, hydrothorax, and ascites.
METHODS: From 2021 to 2024, the clinical data, including imaging, laboratory results, and medical records, of 425 patients with suspected TB infection were retrospectively collected and classified as PTB [306 (72%)], EPTB [67 (15.76%)], and non-TB [52 (12.24%)], according to the final clinical diagnosis.
RESULTS: Among the three groups, significant differences were observed in age, sex, cough, smoking history, hsCRP, IFN-γ, and hypertension (P < 0.05). According to the final clinical diagnosis, the sensitivity and specificity of mNGS in all samples were 90.1% (95% CI: 0.866-0.927) and 75.0% (95% CI: 0.618-0.848), respectively. The sensitivity of mNGS was higher than that of TB-IGRA (79.1%), Xpert (72.4%), culture (52.3%), TB-DNA (45.6%), TB-IgG (36.4%), and AFS (10.7%). For distinguishing TB from non-TB, the sensitivity of mNGS was approximately 37.8% higher than that of standard test culture (90.1% vs 36.4%; P < 0.001). Further analysis also showed that MTB read counts from mNGS were highly correlated with MTB burden measured by other laboratory tests (P < 0.05). Analysis of the ROC curve of mNGS reads showed that the AUC for MTB diagnosis was 0.9073, with a sensitivity of 73.19% and specificity of 96.15% when the recommended cutoff value of 4 reads (Youden's index, 0.6934) was applied.
CONCLUSIONS: For patients with suspected TB or NTM infection, mNGS may serve as a helpful adjunctive tool, particularly when routine diagnostic tests are unrevealing or indeterminate. It has the potential to assist in distinguishing MTB from NTM and to provide additional evidence to guide appropriate treatment choices, though its results should always be interpreted alongside clinical findings and conventional microbiological testing.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
Retrospective Studies
*High-Throughput Nucleotide Sequencing/methods
*Metagenomics/methods
Sensitivity and Specificity
Female
Male
Middle Aged
Adult
Aged
*Tuberculosis/diagnosis/microbiology
*Mycobacterium Infections, Nontuberculous/diagnosis/microbiology
Diagnosis, Differential
*Mycobacterium/genetics
RevDate: 2026-11-13
CmpDate: 2026-11-13
From maternal microbial transmission to neonatal gut microbiota restoration: the potential and challenges of maternal fecal microbiota transplantation.
Frontiers in microbiology, 17:1972665.
Early-life gut microbiota establishment is a critical developmental process shaped by maternal microbial transmission and ecological succession. Microorganisms from the maternal gut, vagina, breast milk, skin, and oral cavity contribute to neonatal microbial acquisition, while strain-resolved studies increasingly support the maternal gut as an important reservoir of strains capable of persistent colonization in the infant intestine. This transmission can be disrupted by cesarean delivery, prematurity, perinatal antibiotic exposure, and neonatal intensive care, resulting in delayed acquisition of maternal commensals, altered microbial succession, and increased colonization by opportunistic microorganisms. Current restoration strategies, including probiotics, synbiotics, and vaginal microbiota seeding, can modify early microbial communities but do not fully reproduce the diversity, maternal specificity, and persistence of naturally transmitted maternal gut strains. Maternal fecal microbiota transplantation (FMT) has therefore emerged as a potential strategy to restore disrupted maternal gut-to-infant microbial transmission. Initial proof-of-concept and strain-resolved studies suggest that carefully screened maternal FMT can increase mother-to-infant strain sharing, promote recovery from cesarean-associated microbiota perturbations, and reduce colonization by potential pathogens. However, evidence for durable engraftment, functional benefits, clinical efficacy, and long-term safety remains limited. Future studies should prioritize rigorous donor screening, standardized intervention protocols, appropriate recipient selection, and longitudinal integration of strain-resolved metagenomics, metabolomics, immune profiling, and clinical outcomes. Maternal FMT remains investigational but provides a promising framework for precision early-life microbiota restoration.
Additional Links: PMID-43022205
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43022205,
year = {2026},
author = {Hu, X and Ma, G and Han, J and Zhu, L and He, Y and Zhu, D},
title = {From maternal microbial transmission to neonatal gut microbiota restoration: the potential and challenges of maternal fecal microbiota transplantation.},
journal = {Frontiers in microbiology},
volume = {17},
number = {},
pages = {1972665},
pmid = {43022205},
issn = {1664-302X},
abstract = {Early-life gut microbiota establishment is a critical developmental process shaped by maternal microbial transmission and ecological succession. Microorganisms from the maternal gut, vagina, breast milk, skin, and oral cavity contribute to neonatal microbial acquisition, while strain-resolved studies increasingly support the maternal gut as an important reservoir of strains capable of persistent colonization in the infant intestine. This transmission can be disrupted by cesarean delivery, prematurity, perinatal antibiotic exposure, and neonatal intensive care, resulting in delayed acquisition of maternal commensals, altered microbial succession, and increased colonization by opportunistic microorganisms. Current restoration strategies, including probiotics, synbiotics, and vaginal microbiota seeding, can modify early microbial communities but do not fully reproduce the diversity, maternal specificity, and persistence of naturally transmitted maternal gut strains. Maternal fecal microbiota transplantation (FMT) has therefore emerged as a potential strategy to restore disrupted maternal gut-to-infant microbial transmission. Initial proof-of-concept and strain-resolved studies suggest that carefully screened maternal FMT can increase mother-to-infant strain sharing, promote recovery from cesarean-associated microbiota perturbations, and reduce colonization by potential pathogens. However, evidence for durable engraftment, functional benefits, clinical efficacy, and long-term safety remains limited. Future studies should prioritize rigorous donor screening, standardized intervention protocols, appropriate recipient selection, and longitudinal integration of strain-resolved metagenomics, metabolomics, immune profiling, and clinical outcomes. Maternal FMT remains investigational but provides a promising framework for precision early-life microbiota restoration.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Recurrent pulmonary infection, a vicious cycle of multifactorial interactions, and prevention/treatment strategies.
Frontiers in immunology, 17:1945273.
Recurrent pulmonary infection (RPI) refers to the frequent and repeated episodes of lower respiratory tract infections. At its core, it represents a vicious cycle arising from the disruption of the homeostatic balance among pathogens, host immunity, and pulmonary architecture. RPI accounts for a considerable proportion of pediatric pneumonia cases, and the cumulative incidence among elderly patients hospitalized for pneumonia is also markedly elevated, imposing a substantial disease burden. The etiologies are notably diverse, beyond common pathogens such as bacteria and viruses, the ubiquity of asymptomatic colonization, underlying comorbidities, immune deficiencies, medication-related factors, and aspiration all constitute critical predisposing factors. This review summarizes the pathogenic characteristics of RPI, host-related factors, and the mechanistic roles of structural lung damage in infection recurrence. It further focuses on the current advances and limitations of precision diagnostic approaches (metagenomic sequencing), immunomodulatory and microbiome-targeted interventions, and vaccination strategies. Existing evidence suggests that the clinical benefits of these interventions in RPI remain to be validated by more high-quality studies. The aim of this article is to provide a reference for the clinical management of RPI and to inform future research directions.
Additional Links: PMID-43022214
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43022214,
year = {2026},
author = {Shen, C and Li, Y and Xiang, Z and Zhao, M and Wang, J and Wang, X},
title = {Recurrent pulmonary infection, a vicious cycle of multifactorial interactions, and prevention/treatment strategies.},
journal = {Frontiers in immunology},
volume = {17},
number = {},
pages = {1945273},
pmid = {43022214},
issn = {1664-3224},
mesh = {Humans ; *Respiratory Tract Infections/prevention & control/therapy/immunology/diagnosis ; Recurrence ; Host-Pathogen Interactions/immunology ; Animals ; *Reinfection/prevention & control ; Risk Factors ; Disease Susceptibility ; Vaccination ; },
abstract = {Recurrent pulmonary infection (RPI) refers to the frequent and repeated episodes of lower respiratory tract infections. At its core, it represents a vicious cycle arising from the disruption of the homeostatic balance among pathogens, host immunity, and pulmonary architecture. RPI accounts for a considerable proportion of pediatric pneumonia cases, and the cumulative incidence among elderly patients hospitalized for pneumonia is also markedly elevated, imposing a substantial disease burden. The etiologies are notably diverse, beyond common pathogens such as bacteria and viruses, the ubiquity of asymptomatic colonization, underlying comorbidities, immune deficiencies, medication-related factors, and aspiration all constitute critical predisposing factors. This review summarizes the pathogenic characteristics of RPI, host-related factors, and the mechanistic roles of structural lung damage in infection recurrence. It further focuses on the current advances and limitations of precision diagnostic approaches (metagenomic sequencing), immunomodulatory and microbiome-targeted interventions, and vaccination strategies. Existing evidence suggests that the clinical benefits of these interventions in RPI remain to be validated by more high-quality studies. The aim of this article is to provide a reference for the clinical management of RPI and to inform future research directions.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
Humans
*Respiratory Tract Infections/prevention & control/therapy/immunology/diagnosis
Recurrence
Host-Pathogen Interactions/immunology
Animals
*Reinfection/prevention & control
Risk Factors
Disease Susceptibility
Vaccination
RevDate: 2026-11-13
CmpDate: 2026-11-13
Decoding multispecies oral biofilms: molecular architecture, host modulation and systems-guided precision anti-biofilm strategies.
Frontiers in cellular and infection microbiology, 16:2002174.
Oral biofilms are spatially ordered microbial communities whose pathogenic behavior is shaped by more than taxonomic composition. Their development reflects the combined effects of adhesin-receptor recognition, interspecies coaggregation, matrix formation, metabolite exchange, local physicochemical gradients and host-derived selection. Here, we synthesize molecular and ecological evidence on the bacterial cell surface-matrix interface and consider how these interactions contribute to cariogenic, periodontal and peri-implant biofilms. Particular attention is given to glucosyltransferase-dependent polysaccharides, extracellular DNA, amyloidogenic proteins, sortase-anchored surface factors and outer membrane vesicles, together with metabolic interactions involving lactate, arginine, hydrogen peroxide and heme. We further evaluate functional-genomic and systems-level approaches (including transposon sequencing, CRISPR interference, metagenomics, metatranscriptomics, metaproteomics, metabolomics, spatial imaging, genome-scale metabolic modeling and machine learning) while distinguishing association and prediction from experimentally demonstrated mechanism. Emerging interventions are assessed according to their biological target and evidence level, including adhesion blockade, specifically targeted antimicrobials, matrix synthesis inhibition and degradation, quorum interference, probiotics, catalytic nanoparticles, antimicrobial photodynamic therapy and antibiofilm dental materials. Across these approaches, reduction in biofilm biomass is not equivalent to ecological restoration or clinical benefit. Key translational barriers include model fidelity, donor-to-donor variation, spatial heterogeneity, treatment durability, endpoint selection and the limited external validation of many computational and preclinical findings. Future precision strategies should therefore integrate microbial identity, matrix architecture, spatial metabolism and host context, with experimental perturbation used to establish causal vulnerabilities before therapeutic translation.
Additional Links: PMID-43022273
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43022273,
year = {2026},
author = {Almusallam, AA and Allemailem, KS},
title = {Decoding multispecies oral biofilms: molecular architecture, host modulation and systems-guided precision anti-biofilm strategies.},
journal = {Frontiers in cellular and infection microbiology},
volume = {16},
number = {},
pages = {2002174},
pmid = {43022273},
issn = {2235-2988},
mesh = {*Biofilms/drug effects/growth & development ; Humans ; *Mouth/microbiology ; Bacteria/genetics/drug effects ; Microbiota ; },
abstract = {Oral biofilms are spatially ordered microbial communities whose pathogenic behavior is shaped by more than taxonomic composition. Their development reflects the combined effects of adhesin-receptor recognition, interspecies coaggregation, matrix formation, metabolite exchange, local physicochemical gradients and host-derived selection. Here, we synthesize molecular and ecological evidence on the bacterial cell surface-matrix interface and consider how these interactions contribute to cariogenic, periodontal and peri-implant biofilms. Particular attention is given to glucosyltransferase-dependent polysaccharides, extracellular DNA, amyloidogenic proteins, sortase-anchored surface factors and outer membrane vesicles, together with metabolic interactions involving lactate, arginine, hydrogen peroxide and heme. We further evaluate functional-genomic and systems-level approaches (including transposon sequencing, CRISPR interference, metagenomics, metatranscriptomics, metaproteomics, metabolomics, spatial imaging, genome-scale metabolic modeling and machine learning) while distinguishing association and prediction from experimentally demonstrated mechanism. Emerging interventions are assessed according to their biological target and evidence level, including adhesion blockade, specifically targeted antimicrobials, matrix synthesis inhibition and degradation, quorum interference, probiotics, catalytic nanoparticles, antimicrobial photodynamic therapy and antibiofilm dental materials. Across these approaches, reduction in biofilm biomass is not equivalent to ecological restoration or clinical benefit. Key translational barriers include model fidelity, donor-to-donor variation, spatial heterogeneity, treatment durability, endpoint selection and the limited external validation of many computational and preclinical findings. Future precision strategies should therefore integrate microbial identity, matrix architecture, spatial metabolism and host context, with experimental perturbation used to establish causal vulnerabilities before therapeutic translation.},
}
MeSH Terms:
show MeSH Terms
hide MeSH Terms
*Biofilms/drug effects/growth & development
Humans
*Mouth/microbiology
Bacteria/genetics/drug effects
Microbiota
RevDate: 2026-11-13
CmpDate: 2026-11-13
Detection and genomic characterization of an avian orthoavulavirus 1 from wild birds in Mongolia in 2022 using metagenomic analysis.
Clinical and experimental vaccine research, 15(4):346-352.
PURPOSE: Mongolia encompasses the breeding areas of various migratory wild birds along multiple flyways, making it important for the study of virus circulation in wild bird populations. While previous surveillance has primarily focused on avian influenza, metagenomic next-generation sequencing offers an opportunity to detect and characterize a broader range of viral pathogens. This study aimed to characterize viral communities and potential avian pathogens in wild bird samples from Mongolia and to investigate the genetic and phylogenetic characteristics of recovered viral genomes.
MATERIALS AND METHODS: Egg-propagated samples derived from wild bird feces, which showed hemagglutination-positive but influenza-negative results, were subjected to metagenomic next-generation sequencing. Sequence reads were taxonomically classified using Kraken2, and samples with substantial viral reads were further analyzed. The recovered viral genome was characterized by phylogenetic and phylogenetic incongruence analyses.
RESULTS: Metagenomic analysis identified abundant bacteriophage reads from seven genera, Asteriusvirus, Saphexavirus, Krischvirus, Mooglevirus, Tequatrovirus, Kagunavirus, and Felixounavirus, as well as Avian orthoavulavirus 1. The complete avian orthoavulavirus 1 genome was recovered and classified as class II genotype I.2. The isolate clustered with recent wild bird strains from Ukraine and Russia, and its fusion protein cleavage-site motif was consistent with a lentogenic pathotype. No clear evidence of phylogenetic incongruence was identified.
CONCLUSION: This study highlights the value of metagenomic surveillance of wild birds for early detection and genomic characterization of avian pathogens, with potential relevance to disease preparedness and subsequent vaccine-related evaluation.
Additional Links: PMID-43022326
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43022326,
year = {2026},
author = {Kim, DW and Kim, JY and Lee, DW and Lee, YJ and Tseren-Ochir, EO and Kwon, JH},
title = {Detection and genomic characterization of an avian orthoavulavirus 1 from wild birds in Mongolia in 2022 using metagenomic analysis.},
journal = {Clinical and experimental vaccine research},
volume = {15},
number = {4},
pages = {346-352},
pmid = {43022326},
issn = {2287-3651},
abstract = {PURPOSE: Mongolia encompasses the breeding areas of various migratory wild birds along multiple flyways, making it important for the study of virus circulation in wild bird populations. While previous surveillance has primarily focused on avian influenza, metagenomic next-generation sequencing offers an opportunity to detect and characterize a broader range of viral pathogens. This study aimed to characterize viral communities and potential avian pathogens in wild bird samples from Mongolia and to investigate the genetic and phylogenetic characteristics of recovered viral genomes.
MATERIALS AND METHODS: Egg-propagated samples derived from wild bird feces, which showed hemagglutination-positive but influenza-negative results, were subjected to metagenomic next-generation sequencing. Sequence reads were taxonomically classified using Kraken2, and samples with substantial viral reads were further analyzed. The recovered viral genome was characterized by phylogenetic and phylogenetic incongruence analyses.
RESULTS: Metagenomic analysis identified abundant bacteriophage reads from seven genera, Asteriusvirus, Saphexavirus, Krischvirus, Mooglevirus, Tequatrovirus, Kagunavirus, and Felixounavirus, as well as Avian orthoavulavirus 1. The complete avian orthoavulavirus 1 genome was recovered and classified as class II genotype I.2. The isolate clustered with recent wild bird strains from Ukraine and Russia, and its fusion protein cleavage-site motif was consistent with a lentogenic pathotype. No clear evidence of phylogenetic incongruence was identified.
CONCLUSION: This study highlights the value of metagenomic surveillance of wild birds for early detection and genomic characterization of avian pathogens, with potential relevance to disease preparedness and subsequent vaccine-related evaluation.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Shotgun Metagenomics Versus 16S rDNA Sanger Sequencing for the Diagnosis of Blood Culture-Negative Endocarditis: A Draw?.
Open forum infectious diseases, 13(11):ofag720.
Although valuable for diagnosing blood culture-negative infective endocarditis, 16S rDNA sequencing and metagenomics on valve/vegetation have never been compared using identical DNA extracts. We compared methods on surgical samples from definite, possible, and rejected infective endocarditis involving culture-negative native valves. Positivity rates were similar, with taxonomic resolution being higher for metagenomics.
Additional Links: PMID-43022586
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43022586,
year = {2026},
author = {Ben Brahim, S and Royer, G and Jacquier, H and Lepeule, R and Goehringer, F and Cheval, J and Rodriguez, C and Alauzet, C and Woerther, PL},
title = {Shotgun Metagenomics Versus 16S rDNA Sanger Sequencing for the Diagnosis of Blood Culture-Negative Endocarditis: A Draw?.},
journal = {Open forum infectious diseases},
volume = {13},
number = {11},
pages = {ofag720},
pmid = {43022586},
issn = {2328-8957},
abstract = {Although valuable for diagnosing blood culture-negative infective endocarditis, 16S rDNA sequencing and metagenomics on valve/vegetation have never been compared using identical DNA extracts. We compared methods on surgical samples from definite, possible, and rejected infective endocarditis involving culture-negative native valves. Positivity rates were similar, with taxonomic resolution being higher for metagenomics.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Oral-predominant microbial and functional dysbiosis in craniofacial microsomia: species-level profiling with compartment-matched statistical testing.
Journal of oral microbiology, 18(1):2754580.
BACKGROUND: Craniofacial microsomia (CFM) is the second most common congenital craniofacial anomaly. Oral dysbiosis has been described at genus level, but whether it extends beyond the mouth, or relates to the defining alveolar bone deficiency, is unknown.
Oral and gut microbiomes of 45 children with CFM and 18 matched controls were profiled at species level by 2bRAD-M, alongside shotgun metagenomes and blood, faecal and oral metabolomes. Alveolar bone loss was quantified on computed tomography against each patient's contralateral side. Centred-log-ratio methods with Benjamini-Hochberg false-discovery-rate (FDR) control were used throughout. Compartment specificity was tested formally by a group-by-compartment interaction model in the 58 participants with paired oral and gut profiles, by sample-size-matched subsampling, and by minimum-detectable-effect calculation.
RESULTS: Oral alpha diversity was higher in CFM than in controls (Shannon P = 0.014; all three alpha-diversity indices P < 0.05). Four species-level taxa were depleted after false-discovery-rate (FDR) correction (q < 0.05), and 21 interpretable metagenomic pathways differed between groups among 38 FDR-significant pathways. These annotations were enriched for carbohydrate transport, glycan processing and cell-surface or biofilm-related functions, but they are not specific markers of a cariogenic or pathogenic community. No FDR-robust gut or systemic signature was detected. At the species level, the oral and gut analyses were power-matched (minimum detectable |r| = 0.74 at the FDR-equivalent threshold), and the group-by-compartment interaction for Shannon diversity was significant (P = 0.026). This comparison supports an oral-predominant taxonomic pattern but does not establish biological absence of a gut effect. The functional comparison was not power-matched. In the patient-only post hoc analyses, the oral functional pattern was not associated with tooth-brushing frequency, sugary-drink intake or caries burden. Affected-side alveolar bone density was lower than contralateral density (paired P < 0.001) and was not associated with anatomical severity grade.
CONCLUSIONS: CFM was associated with an oral-predominant taxonomic and functional microbiome pattern. The matched-power comparison is informative for taxonomic analyses but does not exclude a biologically meaningful gut effect, and the functional comparison remains underpowered. These observations motivate prospective investigation of local, biofilm-directed interventions and of quantitative alveolar bone assessment as a follow-up endpoint.
Additional Links: PMID-43022730
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43022730,
year = {2026},
author = {Li, Z and Zang, T and Zhang, Z and Liu, W and Yin, L and Zhao, S and Liu, B and Ma, L and Tang, X},
title = {Oral-predominant microbial and functional dysbiosis in craniofacial microsomia: species-level profiling with compartment-matched statistical testing.},
journal = {Journal of oral microbiology},
volume = {18},
number = {1},
pages = {2754580},
pmid = {43022730},
issn = {2000-2297},
abstract = {BACKGROUND: Craniofacial microsomia (CFM) is the second most common congenital craniofacial anomaly. Oral dysbiosis has been described at genus level, but whether it extends beyond the mouth, or relates to the defining alveolar bone deficiency, is unknown.
Oral and gut microbiomes of 45 children with CFM and 18 matched controls were profiled at species level by 2bRAD-M, alongside shotgun metagenomes and blood, faecal and oral metabolomes. Alveolar bone loss was quantified on computed tomography against each patient's contralateral side. Centred-log-ratio methods with Benjamini-Hochberg false-discovery-rate (FDR) control were used throughout. Compartment specificity was tested formally by a group-by-compartment interaction model in the 58 participants with paired oral and gut profiles, by sample-size-matched subsampling, and by minimum-detectable-effect calculation.
RESULTS: Oral alpha diversity was higher in CFM than in controls (Shannon P = 0.014; all three alpha-diversity indices P < 0.05). Four species-level taxa were depleted after false-discovery-rate (FDR) correction (q < 0.05), and 21 interpretable metagenomic pathways differed between groups among 38 FDR-significant pathways. These annotations were enriched for carbohydrate transport, glycan processing and cell-surface or biofilm-related functions, but they are not specific markers of a cariogenic or pathogenic community. No FDR-robust gut or systemic signature was detected. At the species level, the oral and gut analyses were power-matched (minimum detectable |r| = 0.74 at the FDR-equivalent threshold), and the group-by-compartment interaction for Shannon diversity was significant (P = 0.026). This comparison supports an oral-predominant taxonomic pattern but does not establish biological absence of a gut effect. The functional comparison was not power-matched. In the patient-only post hoc analyses, the oral functional pattern was not associated with tooth-brushing frequency, sugary-drink intake or caries burden. Affected-side alveolar bone density was lower than contralateral density (paired P < 0.001) and was not associated with anatomical severity grade.
CONCLUSIONS: CFM was associated with an oral-predominant taxonomic and functional microbiome pattern. The matched-power comparison is informative for taxonomic analyses but does not exclude a biologically meaningful gut effect, and the functional comparison remains underpowered. These observations motivate prospective investigation of local, biofilm-directed interventions and of quantitative alveolar bone assessment as a follow-up endpoint.},
}
RevDate: 2026-11-13
CmpDate: 2026-11-13
Plasma metagenomic cell-free DNA sequencing identifies clinically unsuspected cases of Toxoplasma gondii infection in immunocompromised patients: a single-centre experience.
Access microbiology, 8(11):.
Toxoplasmosis can be challenging to diagnose, particularly in immunocompromised patients with nonspecific clinical presentations. We present a retrospective, single-centre case series of eight Toxoplasma gondii infections identified by plasma metagenomic cell-free DNA (cfDNA) sequencing in allogeneic haematopoietic stem cell transplant recipients. Nested within a larger cohort evaluating cfDNA utilization, this case series highlights a positive clinical impact of cfDNA testing in 5 out of 8 (62.5%) patients, including earlier diagnosis and changes in clinical management. These findings illustrate the potential utility of plasma cfDNA sequencing as a complement to conventional diagnostics across diverse clinical presentations of toxoplasmosis.
Additional Links: PMID-43023095
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43023095,
year = {2026},
author = {Goren, LR and Lehman, AC and Drozdov, D and Thielen, BK},
title = {Plasma metagenomic cell-free DNA sequencing identifies clinically unsuspected cases of Toxoplasma gondii infection in immunocompromised patients: a single-centre experience.},
journal = {Access microbiology},
volume = {8},
number = {11},
pages = {},
pmid = {43023095},
issn = {2516-8290},
abstract = {Toxoplasmosis can be challenging to diagnose, particularly in immunocompromised patients with nonspecific clinical presentations. We present a retrospective, single-centre case series of eight Toxoplasma gondii infections identified by plasma metagenomic cell-free DNA (cfDNA) sequencing in allogeneic haematopoietic stem cell transplant recipients. Nested within a larger cohort evaluating cfDNA utilization, this case series highlights a positive clinical impact of cfDNA testing in 5 out of 8 (62.5%) patients, including earlier diagnosis and changes in clinical management. These findings illustrate the potential utility of plasma cfDNA sequencing as a complement to conventional diagnostics across diverse clinical presentations of toxoplasmosis.},
}
RevDate: 2026-11-13
Predominance of viral core functions over auxiliary metabolism among carbohydrate-active enzymes annotated in the global soil virosphere.
Applied and environmental microbiology [Epub ahead of print].
UNLABELLED: Soil viruses are increasingly recognized as components of microbial communities with the potential to alter ecosystem functioning, yet the frequency and functional distribution of virus-encoded metabolic genes in soils remain poorly understood. Here, the Global Soil Virus Atlas gene catalog, comprising 1,432,147 viral genes from 1,223 samples across 12 ecosystem types, was searched against KEGG, Pfam, and CAZy for functions related to carbon cycling, nitrogen cycling, and antimicrobial resistance. Annotations associated with host entry or cell-wall lysis were separated from putative auxiliary metabolic functions. Of 5,020 initial gene-level functional annotations across the three categories, 3,322 (66.2%) were classified as peptidoglycanases. Their exclusion left 1,698 genes (0.14% of the gene catalog restricted to contigs ≥10 kb), and carbon-cycling functions represented 96.7% of the enzyme-resolved annotations. Chitinase-associated genes formed the largest carbon category, but 78.0% belonged to GH19, a family containing both chitinases and phage endolysins. Among GH19 genes on contigs encoding detectable holins or spanins, 84.2% occurred within three genes of these lysis components, a distribution comparable to that of canonical phage lysozymes. Across six matched metagenomic studies, estimated viral contributions to the targeted functions ranged from 0.00% to 8.45% (mean: 2.27%). Two of the three estimates exceeding 2% were GH19-associated chitinase estimates from rhizosphere soils and were therefore treated as upper estimates. Thus, the putative soil viral metabolic inventory was sparse and concentrated in carbon-degradation functions, whereas much of the initial carbohydrate-active enzyme signal reflected core viral lysis functions rather than auxiliary metabolism.
IMPORTANCE: Soil microorganisms drive decomposition and nutrient cycling, and the viruses that infect them may alter these processes by carrying genes that affect host metabolism. However, genes annotated as carbohydrate-degradation enzymes may instead help viruses enter cells or break host cell walls when new virus particles are released. In a worldwide catalog of soil viruses, approximately two-thirds of the initial functional assignments represented these infection or release activities. After their removal, putative viral auxiliary metabolic genes were uncommon and concentrated mainly in carbon degradation. Genes potentially involved in chitin degradation were most frequent, but many occurred near viral genes used to break host cells; their estimated contribution should therefore be viewed as an upper limit. Distinguishing core viral functions from host metabolic modification is essential to avoid overestimating viral contributions to soil processes and to identify the strongest candidates for experimental testing.
Additional Links: PMID-43023715
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43023715,
year = {2026},
author = {Merges, D},
title = {Predominance of viral core functions over auxiliary metabolism among carbohydrate-active enzymes annotated in the global soil virosphere.},
journal = {Applied and environmental microbiology},
volume = {},
number = {},
pages = {e0181326},
doi = {10.1128/aem.01813-26},
pmid = {43023715},
issn = {1098-5336},
abstract = {UNLABELLED: Soil viruses are increasingly recognized as components of microbial communities with the potential to alter ecosystem functioning, yet the frequency and functional distribution of virus-encoded metabolic genes in soils remain poorly understood. Here, the Global Soil Virus Atlas gene catalog, comprising 1,432,147 viral genes from 1,223 samples across 12 ecosystem types, was searched against KEGG, Pfam, and CAZy for functions related to carbon cycling, nitrogen cycling, and antimicrobial resistance. Annotations associated with host entry or cell-wall lysis were separated from putative auxiliary metabolic functions. Of 5,020 initial gene-level functional annotations across the three categories, 3,322 (66.2%) were classified as peptidoglycanases. Their exclusion left 1,698 genes (0.14% of the gene catalog restricted to contigs ≥10 kb), and carbon-cycling functions represented 96.7% of the enzyme-resolved annotations. Chitinase-associated genes formed the largest carbon category, but 78.0% belonged to GH19, a family containing both chitinases and phage endolysins. Among GH19 genes on contigs encoding detectable holins or spanins, 84.2% occurred within three genes of these lysis components, a distribution comparable to that of canonical phage lysozymes. Across six matched metagenomic studies, estimated viral contributions to the targeted functions ranged from 0.00% to 8.45% (mean: 2.27%). Two of the three estimates exceeding 2% were GH19-associated chitinase estimates from rhizosphere soils and were therefore treated as upper estimates. Thus, the putative soil viral metabolic inventory was sparse and concentrated in carbon-degradation functions, whereas much of the initial carbohydrate-active enzyme signal reflected core viral lysis functions rather than auxiliary metabolism.
IMPORTANCE: Soil microorganisms drive decomposition and nutrient cycling, and the viruses that infect them may alter these processes by carrying genes that affect host metabolism. However, genes annotated as carbohydrate-degradation enzymes may instead help viruses enter cells or break host cell walls when new virus particles are released. In a worldwide catalog of soil viruses, approximately two-thirds of the initial functional assignments represented these infection or release activities. After their removal, putative viral auxiliary metabolic genes were uncommon and concentrated mainly in carbon degradation. Genes potentially involved in chitin degradation were most frequent, but many occurred near viral genes used to break host cells; their estimated contribution should therefore be viewed as an upper limit. Distinguishing core viral functions from host metabolic modification is essential to avoid overestimating viral contributions to soil processes and to identify the strongest candidates for experimental testing.},
}
RevDate: 2026-11-13
Characterizing gut microbial structural variation reveals strain heterogeneity and metabolic dysfunction in colorectal cancer.
mSystems [Epub ahead of print].
Previous studies investigating gastrointestinal (GI) diseases have primarily focused on the composition, gene expression, and specific strains of gut microbiota. However, the impact of genetic variations, especially structural variations (SVs) in microbial genomes, on the development of GI tumors has been largely overlooked. In this study, we analyzed 1,027 fecal metagenomes from seven data sets and identified 8,278 coverage-defined SVs across 51 bacterial species in the gut microbiomes. We found that SVs were prevalent in the gut microbiota of studied cohorts. Surprisingly, the genetic makeup of SV-based clusters differed significantly between healthy individuals and those with colorectal cancer (CRC), indicating substantial intra-specific diversity within these populations. By integrating taxonomic-SV markers, we achieved optimal accuracy in early CRC detection, and the SVs also displayed regional heterogeneity. Microbial genetic variation provides additional insights into taxonomy and functionality, allowing us to have a more precise understanding of the microbiota associated with CRC. Notably, our study uncovered potential links between microbial SVs and metabolic disorders of CRC, including SV regions that encode glycoside hydrolases associated with disordered carbohydrate metabolism in CRC. We integrated a framework called MicroSVsProfiler to identify and classify SVs in individuals diagnosed with CRC, as well as in healthy controls. Our findings underscore the importance of considering SVs in microbiome studies, especially in the absence of strain-specific differences in abundance. Future research should explore the deeper layer of the microbiota's structural variantome for a better understanding of the occurrence and characteristics of CRC across different stages.IMPORTANCEMost microbiome studies of colorectal cancer focus on which bacterial species are more or less abundant, potentially overlooking genetic differences among strains of the same species. By analyzing 1,027 fecal metagenomes from seven independent cohorts, we show that large-scale changes in bacterial genomes, known as structural variations, provide disease-related information that is not captured by species abundance alone. These strain-level SV signatures distinguished healthy individuals from patients with colorectal cancer, improved early-cancer classification when combined with taxonomic markers, and varied across geographic populations. We also found that cancer-associated structural variations were linked to altered microbial metabolic functions, particularly carbohydrate metabolism and glycoside hydrolases. Together, these findings establish microbial structural variation as a complementary dimension of the gut microbiome and provide a framework for developing more precise, population-aware biomarkers and for investigating how strain-level genetic changes may contribute to metabolic disturbances associated with colorectal cancer.
Additional Links: PMID-43023736
Publisher:
PubMed:
Citation:
show bibtex listing
hide bibtex listing
@article {pmid43023736,
year = {2026},
author = {Cui, Y and Qiu, Q and Chen, Y and Ding, T},
title = {Characterizing gut microbial structural variation reveals strain heterogeneity and metabolic dysfunction in colorectal cancer.},
journal = {mSystems},
volume = {},
number = {},
pages = {e0110926},
doi = {10.1128/msystems.01109-26},
pmid = {43023736},
issn = {2379-5077},
abstract = {Previous studies investigating gastrointestinal (GI) diseases have primarily focused on the composition, gene expression, and specific strains of gut microbiota. However, the impact of genetic variations, especially structural variations (SVs) in microbial genomes, on the development of GI tumors has been largely overlooked. In this study, we analyzed 1,027 fecal metagenomes from seven data sets and identified 8,278 coverage-defined SVs across 51 bacterial species in the gut microbiomes. We found that SVs were prevalent in the gut microbiota of studied cohorts. Surprisingly, the genetic makeup of SV-based clusters differed significantly between healthy individuals and those with colorectal cancer (CRC), indicating substantial intra-specific diversity within these populations. By integrating taxonomic-SV markers, we achieved optimal accuracy in early CRC detection, and the SVs also displayed regional heterogeneity. Microbial genetic variation provides additional insights into taxonomy and functionality, allowing us to have a more precise understanding of the microbiota associated with CRC. Notably, our study uncovered potential links between microbial SVs and metabolic disorders of CRC, including SV regions that encode glycoside hydrolases associated with disordered carbohydrate metabolism in CRC. We integrated a framework called MicroSVsProfiler to identify and classify SVs in individuals diagnosed with CRC, as well as in healthy controls. Our findings underscore the importance of considering SVs in microbiome studies, especially in the absence of strain-specific differences in abundance. Future research should explore the deeper layer of the microbiota's structural variantome for a better understanding of the occurrence and characteristics of CRC across different stages.IMPORTANCEMost microbiome studies of colorectal cancer focus on which bacterial species are more or less abundant, potentially overlooking genetic differences among strains of the same species. By analyzing 1,027 fecal metagenomes from seven independent cohorts, we show that large-scale changes in bacterial genomes, known as structural variations, provide disease-related information that is not captured by species abundance alone. These strain-level SV signatures distinguished healthy individuals from patients with colorectal cancer, improved early-cancer classification when combined with taxonomic markers, and varied across geographic populations. We also found that cancer-associated structural variations were linked to altered microbial metabolic functions, particularly carbohydrate metabolism and glycoside hydrolases. Together, these findings establish microbial structural variation as a complementary dimension of the gut microbiome and provide a framework for developing more precise, population-aware biomarkers and for investigating how strain-level genetic changes may contribute to metabolic disturbances associated with colorectal cancer.},
}
▼ ▼ LOAD NEXT 100 CITATIONS
RJR Experience and Expertise
Researcher
Robbins holds BS, MS, and PhD degrees in the life sciences. He served as a tenured faculty member in the Zoology and Biological Science departments at Michigan State University. He is currently exploring the intersection between genomics, microbial ecology, and biodiversity — an area that promises to transform our understanding of the biosphere.
Educator
Robbins has extensive experience in college-level education: At MSU he taught introductory biology, genetics, and population genetics. At JHU, he was an instructor for a special course on biological database design. At FHCRC, he team-taught a graduate-level course on the history of genetics. At Bellevue College he taught medical informatics.
Administrator
Robbins has been involved in science administration at both the federal and the institutional levels. At NSF he was a program officer for database activities in the life sciences, at DOE he was a program officer for information infrastructure in the human genome project. At the Fred Hutchinson Cancer Research Center, he served as a vice president for fifteen years.
Technologist
Robbins has been involved with information technology since writing his first Fortran program as a college student. At NSF he was the first program officer for database activities in the life sciences. At JHU he held an appointment in the CS department and served as director of the informatics core for the Genome Data Base. At the FHCRC he was VP for Information Technology.
Publisher
While still at Michigan State, Robbins started his first publishing venture, founding a small company that addressed the short-run publishing needs of instructors in very large undergraduate classes. For more than 20 years, Robbins has been operating The Electronic Scholarly Publishing Project, a web site dedicated to the digital publishing of critical works in science, especially classical genetics.
Speaker
Robbins is well-known for his speaking abilities and is often called upon to provide keynote or plenary addresses at international meetings. For example, in July, 2012, he gave a well-received keynote address at the Global Biodiversity Informatics Congress, sponsored by GBIF and held in Copenhagen. The slides from that talk can be seen HERE.
Facilitator
Robbins is a skilled meeting facilitator. He prefers a participatory approach, with part of the meeting involving dynamic breakout groups, created by the participants in real time: (1) individuals propose breakout groups; (2) everyone signs up for one (or more) groups; (3) the groups with the most interested parties then meet, with reports from each group presented and discussed in a subsequent plenary session.
Designer
Robbins has been engaged with photography and design since the 1960s, when he worked for a professional photography laboratory. He now prefers digital photography and tools for their precision and reproducibility. He designed his first web site more than 20 years ago and he personally designed and implemented this web site. He engages in graphic design as a hobby.
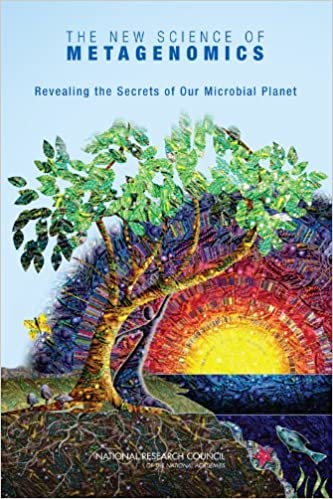
Support this website:
Order from Amazon
We will earn a commission.
Although we can't usually see them, microbes are essential for every part of human life—indeed all life on Earth. The emerging field of metagenomics offers a new way of exploring the microbial world that will transform modern microbiology and lead to practical applications in medicine, agriculture, alternative energy, environmental remediation, and many others areas. Metagenomics allows researchers to look at the genomes of all of the microbes in an environment at once, providing a "meta" view of the whole microbial community and the complex interactions within it. It's a quantum leap beyond traditional research techniques that rely on studying—one at a time—the few microbes that can be grown in the laboratory. At the request of the National Science Foundation, five Institutes of the National Institutes of Health, and the Department of Energy, the National Research Council organized a committee to address the current state of metagenomics and identify obstacles current researchers are facing in order to determine how to best support the field and encourage its success. The New Science of Metagenomics recommends the establishment of a "Global Metagenomics Initiative" comprising a small number of large-scale metagenomics projects as well as many medium- and small-scale projects to advance the technology and develop the standard practices needed to advance the field. The report also addresses database needs, methodological challenges, and the importance of interdisciplinary collaboration in supporting this new field.
RJR Picks from Around the Web (updated 11 MAY 2018 )
Old Science
Weird Science
Treating Disease with Fecal Transplantation
Fossils of miniature humans (hobbits) discovered in Indonesia
Paleontology
Dinosaur tail, complete with feathers, found preserved in amber.
Astronomy
Mysterious fast radio burst (FRB) detected in the distant universe.
Big Data & Informatics
Big Data: Buzzword or Big Deal?
Hacking the genome: Identifying anonymized human subjects using publicly available data.